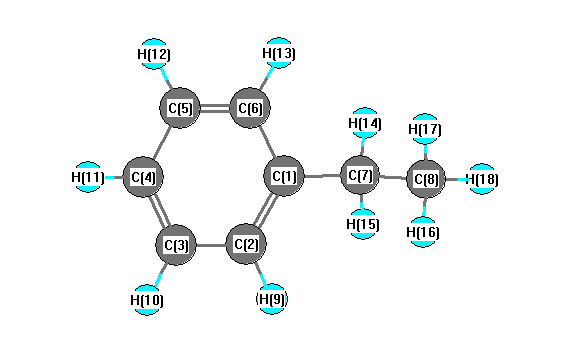
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3219 |
3079 |
19.96 |
|
|
|
| 2 |
A' |
3197 |
3058 |
6.10 |
|
|
|
| 3 |
A' |
3181 |
3043 |
10.59 |
|
|
|
| 4 |
A' |
3134 |
2999 |
41.85 |
|
|
|
| 5 |
A' |
3057 |
2924 |
34.00 |
|
|
|
| 6 |
A' |
3054 |
2922 |
29.70 |
|
|
|
| 7 |
A' |
1677 |
1605 |
4.76 |
|
|
|
| 8 |
A' |
1549 |
1482 |
12.22 |
|
|
|
| 9 |
A' |
1532 |
1466 |
2.77 |
|
|
|
| 10 |
A' |
1512 |
1447 |
1.07 |
|
|
|
| 11 |
A' |
1428 |
1366 |
1.85 |
|
|
|
| 12 |
A' |
1368 |
1308 |
3.29 |
|
|
|
| 13 |
A' |
1238 |
1185 |
0.22 |
|
|
|
| 14 |
A' |
1213 |
1161 |
0.17 |
|
|
|
| 15 |
A' |
1094 |
1047 |
9.12 |
|
|
|
| 16 |
A' |
1064 |
1018 |
2.76 |
|
|
|
| 17 |
A' |
1017 |
973 |
0.20 |
|
|
|
| 18 |
A' |
994 |
951 |
0.16 |
|
|
|
| 19 |
A' |
989 |
946 |
2.35 |
|
|
|
| 20 |
A' |
920 |
881 |
1.56 |
|
|
|
| 21 |
A' |
788 |
754 |
5.51 |
|
|
|
| 22 |
A' |
768 |
735 |
24.72 |
|
|
|
| 23 |
A' |
715 |
684 |
29.79 |
|
|
|
| 24 |
A' |
564 |
540 |
5.89 |
|
|
|
| 25 |
A' |
500 |
479 |
3.93 |
|
|
|
| 26 |
A' |
310 |
296 |
0.27 |
|
|
|
| 27 |
A' |
135 |
129 |
0.61 |
|
|
|
| 28 |
A" |
3206 |
3067 |
40.85 |
|
|
|
| 29 |
A" |
3182 |
3045 |
7.16 |
|
|
|
| 30 |
A" |
3140 |
3004 |
30.13 |
|
|
|
| 31 |
A" |
3094 |
2960 |
9.74 |
|
|
|
| 32 |
A" |
1655 |
1584 |
0.77 |
|
|
|
| 33 |
A" |
1520 |
1454 |
8.92 |
|
|
|
| 34 |
A" |
1502 |
1437 |
5.61 |
|
|
|
| 35 |
A" |
1381 |
1322 |
0.23 |
|
|
|
| 36 |
A" |
1366 |
1307 |
0.36 |
|
|
|
| 37 |
A" |
1283 |
1228 |
0.07 |
|
|
|
| 38 |
A" |
1191 |
1140 |
0.05 |
|
|
|
| 39 |
A" |
1131 |
1082 |
3.11 |
|
|
|
| 40 |
A" |
1066 |
1020 |
2.36 |
|
|
|
| 41 |
A" |
967 |
925 |
0.00 |
|
|
|
| 42 |
A" |
859 |
822 |
0.01 |
|
|
|
| 43 |
A" |
794 |
760 |
0.95 |
|
|
|
| 44 |
A" |
633 |
606 |
0.04 |
|
|
|
| 45 |
A" |
413 |
395 |
0.00 |
|
|
|
| 46 |
A" |
354 |
339 |
0.06 |
|
|
|
| 47 |
A" |
221 |
212 |
0.01 |
|
|
|
| 48 |
A" |
40 |
38 |
0.01 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 34607.4 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 33108.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.153 |
|
|
|
| 2 |
C |
-0.206 |
|
|
|
| 3 |
C |
-0.161 |
|
|
|
| 4 |
C |
-0.166 |
|
|
|
| 5 |
C |
-0.161 |
|
|
|
| 6 |
C |
-0.206 |
|
|
|
| 7 |
C |
-0.394 |
|
|
|
| 8 |
C |
-0.500 |
|
|
|
| 9 |
H |
0.154 |
|
|
|
| 10 |
H |
0.160 |
|
|
|
| 11 |
H |
0.160 |
|
|
|
| 12 |
H |
0.160 |
|
|
|
| 13 |
H |
0.154 |
|
|
|
| 14 |
H |
0.170 |
|
|
|
| 15 |
H |
0.170 |
|
|
|
| 16 |
H |
0.173 |
|
|
|
| 17 |
H |
0.173 |
|
|
|
| 18 |
H |
0.167 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.058 |
0.301 |
0.000 |
0.306 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-51.643 |
-0.044 |
0.000 |
| y |
-0.044 |
-44.005 |
0.000 |
| z |
0.000 |
0.000 |
-44.079 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-7.601 |
-0.044 |
0.000 |
| y |
-0.044 |
3.856 |
0.000 |
| z |
0.000 |
0.000 |
3.745 |
|
| Polar |
|---|
| 3z2-r2 | 7.490 |
|---|
| x2-y2 | -7.638 |
|---|
| xy | -0.044 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.311 |
0.666 |
0.000 |
| y |
0.666 |
15.585 |
0.000 |
| z |
0.000 |
0.000 |
12.803 |
<r2> (average value of r
2) Å
2
| <r2> |
290.821 |
| (<r2>)1/2 |
17.053 |