return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H5CH2CH3 (Ethylbenzene)

using model chemistry: QCISD/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at QCISD/6-31G*
 hartrees
Energy at 0K-309.855394
Energy at 298.15K-309.865685
HF Energy-308.772739
Nuclear repulsion energy339.398060
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3218 3064 19.44      
2 A' 3195 3043 5.42      
3 A' 3179 3028 10.29      
4 A' 3139 2989 42.42      
5 A' 3063 2917 23.96      
6 A' 3060 2914 32.40      
7 A' 1702 1621 4.09      
8 A' 1569 1494 12.29      
9 A' 1557 1482 2.61      
10 A' 1541 1467 0.39      
11 A' 1460 1390 1.92      
12 A' 1400 1334 3.50      
13 A' 1257 1197 0.15      
14 A' 1232 1174 0.14      
15 A' 1115 1062 6.90      
16 A' 1073 1021 1.71      
17 A' 1025 976 0.28      
18 A' 1005 957 1.95      
19 A' 914 871 0.13      
20 A' 881 839 0.60      
21 A' 793 755 0.99      
22 A' 746 711 58.12      
23 A' 629 599 8.75      
24 A' 547 521 1.83      
25 A' 495 472 5.05      
26 A' 311 296 0.25      
27 A' 137 130 0.63      
28 A" 3204 3052 38.89      
29 A" 3180 3029 7.13      
30 A" 3143 2994 30.66      
31 A" 3099 2951 10.50      
32 A" 1677 1597 0.83      
33 A" 1545 1472 7.26      
34 A" 1519 1447 7.39      
35 A" 1381 1315 0.38      
36 A" 1337 1273 0.09      
37 A" 1295 1234 0.08      
38 A" 1204 1146 0.06      
39 A" 1144 1089 2.69      
40 A" 1080 1029 1.50      
41 A" 919 875 0.00      
42 A" 849 809 0.02      
43 A" 804 766 0.60      
44 A" 639 609 0.03      
45 A" 405 386 0.00      
46 A" 363 346 0.07      
47 A" 227 217 0.02      
48 A" 36 34 0.00      

Unscaled Zero Point Vibrational Energy (zpe) 34645.5 cm-1
Scaled (by 0.9523) Zero Point Vibrational Energy (zpe) 32992.9 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD/6-31G*
ABC
0.15052 0.04876 0.04044

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD/6-31G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -0.240 0.496 0.000
C2 -0.238 -0.221 1.204
C3 -0.238 -1.618 1.207
C4 -0.237 -2.321 0.000
C5 -0.238 -1.618 -1.207
C6 -0.238 -0.221 -1.204
C7 -0.188 2.009 0.000
C8 1.251 2.541 0.000
H9 -0.243 0.321 2.151
H10 -0.242 -2.158 2.153
H11 -0.240 -3.410 0.000
H12 -0.242 -2.158 -2.153
H13 -0.243 0.321 -2.151
H14 -0.723 2.392 -0.881
H15 -0.723 2.392 0.881
H16 1.794 2.189 0.886
H17 1.794 2.189 -0.886
H18 1.267 3.638 0.000

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 C7 C8 H9 H10 H11 H12 H13 H14 H15 H16 H17 H18
C11.40142.43432.81762.43431.40141.51332.53102.15783.41753.90663.41752.15782.14542.14542.79132.79133.4852
C21.40141.39682.42092.78642.40792.53443.36101.09092.15673.40913.87583.39823.37782.67733.16863.78274.3141
C32.43431.39681.39712.41402.78643.82234.57932.15621.08942.16113.40343.87724.54684.05224.32744.79645.5992
C42.81762.42091.39711.39712.42094.33025.08493.40702.15961.08912.15963.40704.81964.81965.02565.02566.1467
C52.43432.78642.41401.39711.39683.82234.57933.87723.40342.16111.08942.15624.05224.54684.79644.32745.5992
C61.40142.40792.78642.42091.39682.53443.36103.39823.87583.40912.15671.09092.67733.37783.78273.16864.3141
C71.51332.53443.82234.33023.82232.53441.53502.73434.69015.41934.69012.73431.09921.09922.17912.17912.1853
C82.53103.36104.57935.08494.57933.36101.53503.43335.37986.13535.37983.43332.16662.16661.09721.09721.0976
H92.15781.09092.15623.40703.87723.39822.73433.43332.47864.30684.96674.30143.70262.47633.03984.10684.2324
H103.41752.15671.08942.15963.40343.87584.69015.37982.47862.49134.30674.96675.48974.74874.96445.68156.3646
H113.90663.40912.16111.08912.16113.40915.41936.13534.30682.49132.49134.30685.88885.88886.02326.02327.2081
H123.41753.87583.40342.15961.08942.15674.69015.37984.96674.30672.49132.47864.74875.48975.68154.96446.3646
H132.15783.39823.87723.40702.15621.09092.73433.43334.30144.96674.30682.47862.47633.70264.10683.03984.2324
H142.14543.37784.54684.81964.05222.67731.09922.16663.70265.48975.88884.74872.47631.76163.08182.52482.5077
H152.14542.67734.05224.81964.54683.37781.09922.16662.47634.74875.88885.48973.70261.76162.52483.08182.5077
H162.79133.16864.32745.02564.79643.78272.17911.09723.03984.96446.02325.68154.10683.08182.52481.77261.7784
H172.79133.78274.79645.02564.32743.16862.17911.09724.10685.68156.02324.96443.03982.52483.08181.77261.7784
H183.48524.31415.59926.14675.59924.31412.18531.09764.23246.36467.20816.36464.23242.50772.50771.77841.7784

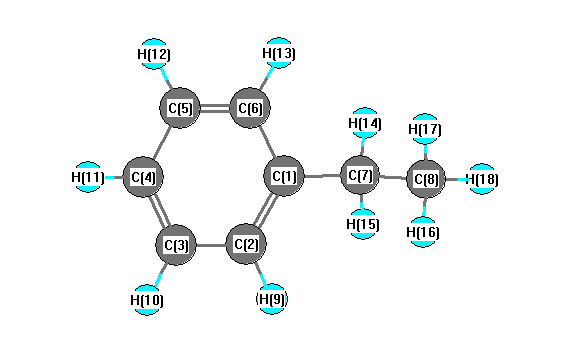
picture of Ethylbenzene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 120.908 C1 C2 H9 119.424
C1 C6 C5 120.908 C1 C6 H13 119.424
C1 C7 C8 112.258 C1 C7 H14 109.393
C1 C7 H15 109.393 C2 C1 C6 118.433
C2 C1 C7 120.759 C2 C3 C4 120.112
C2 C3 H10 119.825 C3 C2 H9 119.666
C3 C4 C5 119.526 C3 C4 H11 120.237
C4 C3 H10 120.062 C4 C5 C6 120.112
C4 C5 H12 120.062 C5 C4 H11 120.237
C5 C6 H13 119.666 C6 C1 C7 120.759
C6 C5 H12 119.825 C7 C8 H16 110.664
C7 C8 H17 110.664 C7 C8 H18 111.131
C8 C7 H14 109.559 C8 C7 H15 109.559
H14 C7 H15 106.513 H16 C8 H17 107.766
H16 C8 H18 108.251 H17 C8 H18 108.251
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability