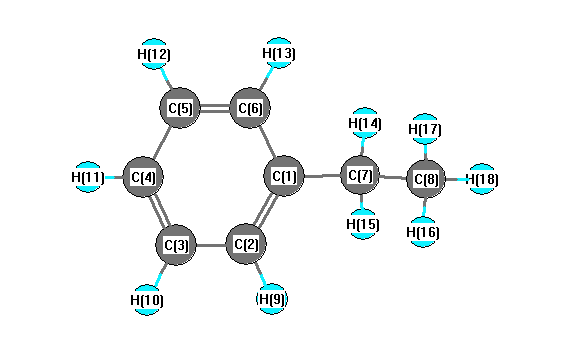
Vibrational Frequencies calculated at HSEh1PBE/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3235 |
3077 |
18.94 |
|
|
|
| 2 |
A' |
3213 |
3056 |
4.56 |
|
|
|
| 3 |
A' |
3197 |
3040 |
10.76 |
|
|
|
| 4 |
A' |
3151 |
2996 |
40.23 |
|
|
|
| 5 |
A' |
3069 |
2919 |
33.63 |
|
|
|
| 6 |
A' |
3068 |
2917 |
26.83 |
|
|
|
| 7 |
A' |
1690 |
1608 |
4.87 |
|
|
|
| 8 |
A' |
1556 |
1480 |
12.62 |
|
|
|
| 9 |
A' |
1534 |
1459 |
2.87 |
|
|
|
| 10 |
A' |
1513 |
1439 |
1.18 |
|
|
|
| 11 |
A' |
1430 |
1360 |
2.16 |
|
|
|
| 12 |
A' |
1373 |
1306 |
3.11 |
|
|
|
| 13 |
A' |
1246 |
1185 |
0.20 |
|
|
|
| 14 |
A' |
1215 |
1156 |
0.15 |
|
|
|
| 15 |
A' |
1101 |
1047 |
9.23 |
|
|
|
| 16 |
A' |
1070 |
1017 |
2.53 |
|
|
|
| 17 |
A' |
1022 |
972 |
0.18 |
|
|
|
| 18 |
A' |
1000 |
951 |
0.15 |
|
|
|
| 19 |
A' |
996 |
947 |
2.70 |
|
|
|
| 20 |
A' |
925 |
880 |
1.55 |
|
|
|
| 21 |
A' |
793 |
754 |
4.75 |
|
|
|
| 22 |
A' |
773 |
735 |
25.49 |
|
|
|
| 23 |
A' |
718 |
683 |
30.16 |
|
|
|
| 24 |
A' |
567 |
539 |
6.03 |
|
|
|
| 25 |
A' |
503 |
479 |
4.00 |
|
|
|
| 26 |
A' |
311 |
296 |
0.28 |
|
|
|
| 27 |
A' |
136 |
129 |
0.62 |
|
|
|
| 28 |
A" |
3222 |
3064 |
36.63 |
|
|
|
| 29 |
A" |
3198 |
3041 |
7.45 |
|
|
|
| 30 |
A" |
3154 |
3000 |
28.79 |
|
|
|
| 31 |
A" |
3109 |
2957 |
9.03 |
|
|
|
| 32 |
A" |
1668 |
1587 |
0.77 |
|
|
|
| 33 |
A" |
1522 |
1448 |
9.41 |
|
|
|
| 34 |
A" |
1509 |
1435 |
5.84 |
|
|
|
| 35 |
A" |
1390 |
1322 |
0.25 |
|
|
|
| 36 |
A" |
1368 |
1301 |
0.36 |
|
|
|
| 37 |
A" |
1287 |
1224 |
0.07 |
|
|
|
| 38 |
A" |
1193 |
1134 |
0.04 |
|
|
|
| 39 |
A" |
1134 |
1079 |
3.30 |
|
|
|
| 40 |
A" |
1070 |
1018 |
2.30 |
|
|
|
| 41 |
A" |
972 |
925 |
0.01 |
|
|
|
| 42 |
A" |
864 |
822 |
0.01 |
|
|
|
| 43 |
A" |
796 |
757 |
0.91 |
|
|
|
| 44 |
A" |
635 |
604 |
0.04 |
|
|
|
| 45 |
A" |
416 |
395 |
0.00 |
|
|
|
| 46 |
A" |
357 |
339 |
0.06 |
|
|
|
| 47 |
A" |
225 |
214 |
0.01 |
|
|
|
| 48 |
A" |
37 |
35 |
0.01 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 34766.4 cm
-1
Scaled (by 0.951) Zero Point Vibrational Energy (zpe) 33062.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.143 |
|
|
|
| 2 |
C |
-0.205 |
|
|
|
| 3 |
C |
-0.164 |
|
|
|
| 4 |
C |
-0.169 |
|
|
|
| 5 |
C |
-0.164 |
|
|
|
| 6 |
C |
-0.205 |
|
|
|
| 7 |
C |
-0.397 |
|
|
|
| 8 |
C |
-0.504 |
|
|
|
| 9 |
H |
0.157 |
|
|
|
| 10 |
H |
0.163 |
|
|
|
| 11 |
H |
0.163 |
|
|
|
| 12 |
H |
0.163 |
|
|
|
| 13 |
H |
0.157 |
|
|
|
| 14 |
H |
0.173 |
|
|
|
| 15 |
H |
0.173 |
|
|
|
| 16 |
H |
0.175 |
|
|
|
| 17 |
H |
0.175 |
|
|
|
| 18 |
H |
0.169 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.058 |
0.289 |
0.000 |
0.294 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-51.791 |
-0.043 |
0.000 |
| y |
-0.043 |
-44.097 |
0.000 |
| z |
0.000 |
0.000 |
-44.163 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-7.661 |
-0.043 |
0.000 |
| y |
-0.043 |
3.880 |
0.000 |
| z |
0.000 |
0.000 |
3.781 |
|
| Polar |
|---|
| 3z2-r2 | 7.562 |
|---|
| x2-y2 | -7.694 |
|---|
| xy | -0.043 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.299 |
0.665 |
0.000 |
| y |
0.665 |
15.497 |
0.000 |
| z |
0.000 |
0.000 |
12.745 |
<r2> (average value of r
2) Å
2
| <r2> |
289.644 |
| (<r2>)1/2 |
17.019 |