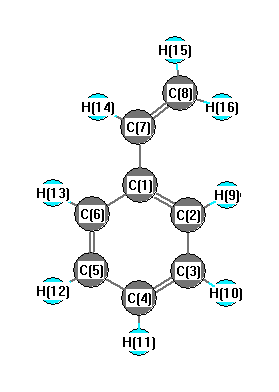
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3262 |
3121 |
14.24 |
|
|
|
| 2 |
A' |
3222 |
3082 |
15.96 |
|
|
|
| 3 |
A' |
3213 |
3074 |
35.69 |
|
|
|
| 4 |
A' |
3204 |
3066 |
14.69 |
|
|
|
| 5 |
A' |
3194 |
3056 |
0.26 |
|
|
|
| 6 |
A' |
3188 |
3050 |
8.71 |
|
|
|
| 7 |
A' |
3182 |
3044 |
3.88 |
|
|
|
| 8 |
A' |
3160 |
3023 |
16.64 |
|
|
|
| 9 |
A' |
1721 |
1646 |
8.83 |
|
|
|
| 10 |
A' |
1675 |
1602 |
1.20 |
|
|
|
| 11 |
A' |
1647 |
1576 |
2.59 |
|
|
|
| 12 |
A' |
1549 |
1482 |
10.07 |
|
|
|
| 13 |
A' |
1500 |
1435 |
4.84 |
|
|
|
| 14 |
A' |
1469 |
1406 |
5.31 |
|
|
|
| 15 |
A' |
1383 |
1323 |
0.75 |
|
|
|
| 16 |
A' |
1371 |
1311 |
2.75 |
|
|
|
| 17 |
A' |
1334 |
1276 |
2.01 |
|
|
|
| 18 |
A' |
1242 |
1188 |
2.40 |
|
|
|
| 19 |
A' |
1217 |
1164 |
0.08 |
|
|
|
| 20 |
A' |
1193 |
1141 |
0.13 |
|
|
|
| 21 |
A' |
1124 |
1075 |
4.55 |
|
|
|
| 22 |
A' |
1069 |
1023 |
0.16 |
|
|
|
| 23 |
A' |
1047 |
1001 |
6.43 |
|
|
|
| 24 |
A' |
1015 |
971 |
0.03 |
|
|
|
| 25 |
A' |
792 |
757 |
0.24 |
|
|
|
| 26 |
A' |
632 |
604 |
0.03 |
|
|
|
| 27 |
A' |
558 |
534 |
4.74 |
|
|
|
| 28 |
A' |
448 |
428 |
0.10 |
|
|
|
| 29 |
A' |
235 |
225 |
0.44 |
|
|
|
| 30 |
A" |
1035 |
990 |
15.81 |
|
|
|
| 31 |
A" |
995 |
952 |
0.14 |
|
|
|
| 32 |
A" |
968 |
926 |
0.02 |
|
|
|
| 33 |
A" |
926 |
886 |
5.18 |
|
|
|
| 34 |
A" |
921 |
882 |
40.66 |
|
|
|
| 35 |
A" |
854 |
817 |
0.40 |
|
|
|
| 36 |
A" |
799 |
765 |
33.70 |
|
|
|
| 37 |
A" |
712 |
681 |
34.57 |
|
|
|
| 38 |
A" |
655 |
626 |
0.02 |
|
|
|
| 39 |
A" |
446 |
426 |
5.83 |
|
|
|
| 40 |
A" |
411 |
393 |
0.30 |
|
|
|
| 41 |
A" |
206 |
197 |
1.67 |
|
|
|
| 42 |
A" |
26 |
25 |
0.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 29398.3 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 28125.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.143 |
|
|
|
| 2 |
C |
-0.202 |
|
|
|
| 3 |
C |
-0.165 |
|
|
|
| 4 |
C |
-0.162 |
|
|
|
| 5 |
C |
-0.162 |
|
|
|
| 6 |
C |
-0.220 |
|
|
|
| 7 |
C |
-0.140 |
|
|
|
| 8 |
C |
-0.399 |
|
|
|
| 9 |
H |
0.160 |
|
|
|
| 10 |
H |
0.164 |
|
|
|
| 11 |
H |
0.163 |
|
|
|
| 12 |
H |
0.164 |
|
|
|
| 13 |
H |
0.161 |
|
|
|
| 14 |
H |
0.158 |
|
|
|
| 15 |
H |
0.170 |
|
|
|
| 16 |
H |
0.166 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.078 |
-0.164 |
0.000 |
0.182 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-41.257 |
0.133 |
0.000 |
| y |
0.133 |
-41.410 |
0.000 |
| z |
0.000 |
0.000 |
-51.646 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
5.271 |
0.133 |
0.000 |
| y |
0.133 |
5.041 |
0.000 |
| z |
0.000 |
0.000 |
-10.312 |
|
| Polar |
|---|
| 3z2-r2 | -20.625 |
|---|
| x2-y2 | 0.153 |
|---|
| xy | 0.133 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
14.363 |
-2.714 |
0.000 |
| y |
-2.714 |
17.588 |
-0.001 |
| z |
0.000 |
-0.001 |
4.108 |
<r2> (average value of r
2) Å
2
| <r2> |
276.672 |
| (<r2>)1/2 |
16.633 |