return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H5CHCH2 (Styrene)

using model chemistry: B2PLYP/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at B2PLYP/6-31G*
 hartrees
Energy at 0K-309.283813
Energy at 298.15K-309.291485
HF Energy-308.947022
Nuclear repulsion energy319.661972
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3283 3116 14.75      
2 A 3238 3073 15.38      
3 A 3229 3065 35.13      
4 A 3220 3057 14.85      
5 A 3211 3048 0.46      
6 A 3203 3040 10.06      
7 A 3201 3039 2.50      
8 A 3180 3019 14.91      
9 A 1717 1630 6.82      
10 A 1672 1587 1.57      
11 A 1646 1562 2.66      
12 A 1556 1477 10.10      
13 A 1510 1433 3.14      
14 A 1484 1409 5.95      
15 A 1388 1318 0.29      
16 A 1382 1311 2.54      
17 A 1346 1277 2.20      
18 A 1248 1184 1.88      
19 A 1228 1166 0.06      
20 A 1204 1143 0.12      
21 A 1129 1071 3.49      
22 A 1073 1018 0.27      
23 A 1054 1000 5.15      
24 A 1033 981 16.39      
25 A 1021 969 0.07      
26 A 956 908 0.23      
27 A 943 895 0.15      
28 A 920 874 39.82      
29 A 903 857 0.09      
30 A 850 806 0.40      
31 A 794 754 0.30      
32 A 774 735 44.10      
33 A 702 666 19.87      
34 A 636 604 0.03      
35 A 634 602 0.07      
36 A 563 534 4.52      
37 A 456 433 2.23      
38 A 437 415 2.91      
39 A 409 388 0.21      
40 A 238 226 0.48      
41 A 204 194 1.50      
42 A 32 30 0.03      

Unscaled Zero Point Vibrational Energy (zpe) 29451.5 cm-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 27955.4 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at B2PLYP/6-31G*
ABC
0.17253 0.05135 0.03963

See section I.F.4 to change rotational constant units
Geometric Data calculated at B2PLYP/6-31G*

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.514 -0.224 -0.065
C2 0.013 1.089 -0.092
C3 -1.358 1.330 -0.030
C4 -2.260 0.263 0.052
C5 -1.776 -1.047 0.067
C6 -0.402 -1.286 0.004
C7 1.958 -0.522 -0.112
C8 2.957 0.333 0.148
H9 0.700 1.926 -0.177
H10 -1.726 2.351 -0.055
H11 -3.328 0.453 0.097
H12 -2.467 -1.883 0.125
H13 -0.032 -2.308 0.017
H14 2.204 -1.551 -0.369
H15 3.993 0.019 0.086
H16 2.785 1.365 0.438

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 C7 C8 H9 H10 H11 H12 H13 H14 H15 H16
C11.40572.43302.81852.43731.40411.47522.51502.16103.41323.90433.41672.15592.17023.49122.8171
C21.40571.39372.42282.79132.41302.52553.04901.08602.14933.40643.87733.39933.44214.12552.8353
C32.43301.39371.39912.41502.78473.79894.43252.14821.08612.15973.40233.87194.59385.51104.1696
C42.81852.42281.39911.39622.41874.29325.21823.40302.15801.08582.15743.40174.83666.25805.1781
C52.43732.79132.41501.39621.39613.77534.93133.87713.40072.15811.08612.15274.03595.86745.1732
C61.40412.41302.78472.41871.39612.48353.73193.40063.87083.40442.15261.08722.64634.58594.1682
C71.47522.52553.79894.29323.77532.48351.34032.75294.67235.37904.63552.67731.08912.11512.1322
C82.51503.04904.43255.21824.93133.73191.34032.78165.10386.28645.85933.99112.09391.08471.0858
H92.16101.08602.14823.40303.87713.40062.75292.78162.46654.29784.96314.30133.79323.81482.2447
H103.41322.14931.08612.15803.40073.87084.67235.10382.46652.48834.30254.95805.54746.17864.6438
H113.90433.40642.15971.08582.15813.40445.37906.28644.29782.48832.49004.29995.90217.33426.1900
H123.41673.87733.40232.15741.08612.15264.63555.85934.96314.30252.49002.47354.70866.73446.1832
H132.15593.39933.87193.40172.15271.08722.67733.99114.30134.95804.29992.47352.39234.65024.6483
H142.17023.44214.59384.83664.03592.64631.08912.09393.79325.54745.90214.70862.39232.42333.0811
H153.49124.12555.51106.25805.86744.58592.11511.08473.81486.17867.33426.73444.65022.42331.8433
H162.81712.83534.16965.17815.17324.16822.13221.08582.24474.64386.19006.18324.64833.08111.8433

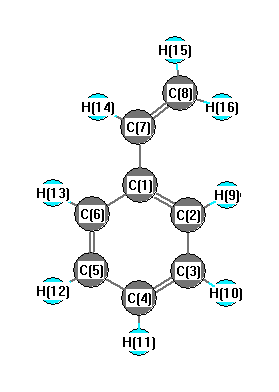
picture of Styrene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 120.712 C1 C2 H9 119.735
C1 C6 C5 121.006 C1 C6 H13 119.307
C1 C7 C8 126.508 C1 C7 H14 114.795
C2 C1 C6 118.356 C2 C1 C7 122.459
C2 C3 C4 120.345 C2 C3 H10 119.642
C3 C2 H9 119.546 C3 C4 C5 119.526
C3 C4 H11 120.187 C4 C3 H10 120.010
C4 C5 C6 120.047 C4 C5 H12 120.196
C5 C4 H11 120.285 C5 C6 H13 119.686
C6 C1 C7 119.186 C6 C5 H12 119.757
C7 C8 H15 121.078 C7 C8 H16 122.664
C8 C7 H14 118.690 H15 C8 H16 116.256
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G* Charges (e)
Number Element Mulliken CHELPG AIM ESP
1 C 0.110      
2 C -0.195      
3 C -0.159      
4 C -0.161      
5 C -0.158      
6 C -0.209      
7 C -0.130      
8 C -0.375      
9 H 0.160      
10 H 0.161      
11 H 0.161      
12 H 0.161      
13 H 0.160      
14 H 0.155      
15 H 0.162      
16 H 0.158      


Electric dipole moments


Electric Quadrupole moment
Quadrupole components in D Å


Polarizabilities
Components of the polarizability tensor.
Units are Å3 (Angstrom cubed)
Change units.
  x y z
x 18.391 0.671 0.016
y 0.671 12.770 0.138
z 0.016 0.138 4.091


<r2> (average value of r2) Å2
<r2> 277.061
(<r2>)1/2 16.645