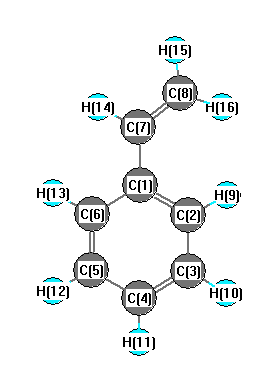
Vibrational Frequencies calculated at HSEh1PBE/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3277 |
3117 |
12.94 |
|
|
|
| 2 |
A |
3238 |
3080 |
15.64 |
|
|
|
| 3 |
A |
3228 |
3070 |
33.73 |
|
|
|
| 4 |
A |
3220 |
3062 |
10.76 |
|
|
|
| 5 |
A |
3210 |
3053 |
0.61 |
|
|
|
| 6 |
A |
3203 |
3046 |
8.66 |
|
|
|
| 7 |
A |
3195 |
3038 |
3.17 |
|
|
|
| 8 |
A |
3175 |
3019 |
16.24 |
|
|
|
| 9 |
A |
1733 |
1648 |
8.51 |
|
|
|
| 10 |
A |
1687 |
1604 |
1.38 |
|
|
|
| 11 |
A |
1659 |
1578 |
2.70 |
|
|
|
| 12 |
A |
1555 |
1479 |
10.66 |
|
|
|
| 13 |
A |
1506 |
1432 |
5.53 |
|
|
|
| 14 |
A |
1473 |
1401 |
5.46 |
|
|
|
| 15 |
A |
1391 |
1323 |
0.84 |
|
|
|
| 16 |
A |
1373 |
1306 |
2.65 |
|
|
|
| 17 |
A |
1338 |
1272 |
2.08 |
|
|
|
| 18 |
A |
1250 |
1189 |
2.44 |
|
|
|
| 19 |
A |
1220 |
1160 |
0.06 |
|
|
|
| 20 |
A |
1194 |
1136 |
0.12 |
|
|
|
| 21 |
A |
1129 |
1073 |
4.66 |
|
|
|
| 22 |
A |
1074 |
1021 |
0.25 |
|
|
|
| 23 |
A |
1049 |
998 |
6.50 |
|
|
|
| 24 |
A |
1040 |
989 |
16.04 |
|
|
|
| 25 |
A |
1020 |
970 |
0.05 |
|
|
|
| 26 |
A |
1000 |
951 |
0.17 |
|
|
|
| 27 |
A |
973 |
925 |
0.02 |
|
|
|
| 28 |
A |
931 |
885 |
5.99 |
|
|
|
| 29 |
A |
929 |
883 |
39.70 |
|
|
|
| 30 |
A |
858 |
816 |
0.39 |
|
|
|
| 31 |
A |
803 |
763 |
33.86 |
|
|
|
| 32 |
A |
796 |
757 |
0.26 |
|
|
|
| 33 |
A |
715 |
680 |
34.88 |
|
|
|
| 34 |
A |
658 |
626 |
0.02 |
|
|
|
| 35 |
A |
633 |
602 |
0.03 |
|
|
|
| 36 |
A |
560 |
532 |
4.76 |
|
|
|
| 37 |
A |
450 |
428 |
0.09 |
|
|
|
| 38 |
A |
447 |
425 |
5.93 |
|
|
|
| 39 |
A |
412 |
392 |
0.34 |
|
|
|
| 40 |
A |
236 |
225 |
0.43 |
|
|
|
| 41 |
A |
207 |
197 |
1.76 |
|
|
|
| 42 |
A |
30 |
29 |
0.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 29535.6 cm
-1
Scaled (by 0.951) Zero Point Vibrational Energy (zpe) 28088.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.132 |
|
|
|
| 2 |
C |
-0.202 |
|
|
|
| 3 |
C |
-0.168 |
|
|
|
| 4 |
C |
-0.165 |
|
|
|
| 5 |
C |
-0.165 |
|
|
|
| 6 |
C |
-0.220 |
|
|
|
| 7 |
C |
-0.141 |
|
|
|
| 8 |
C |
-0.400 |
|
|
|
| 9 |
H |
0.163 |
|
|
|
| 10 |
H |
0.167 |
|
|
|
| 11 |
H |
0.167 |
|
|
|
| 12 |
H |
0.167 |
|
|
|
| 13 |
H |
0.165 |
|
|
|
| 14 |
H |
0.161 |
|
|
|
| 15 |
H |
0.172 |
|
|
|
| 16 |
H |
0.168 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.178 |
-0.006 |
-0.000 |
0.178 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-41.534 |
0.035 |
0.001 |
| y |
0.035 |
-41.262 |
0.001 |
| z |
0.001 |
0.001 |
-51.780 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
4.986 |
0.035 |
0.001 |
| y |
0.035 |
5.395 |
0.001 |
| z |
0.001 |
0.001 |
-10.382 |
|
| Polar |
|---|
| 3z2-r2 | -20.763 |
|---|
| x2-y2 | -0.272 |
|---|
| xy | 0.035 |
|---|
| xz | 0.001 |
|---|
| yz | 0.001 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
18.885 |
-0.715 |
0.000 |
| y |
-0.715 |
12.847 |
0.000 |
| z |
0.000 |
0.000 |
4.104 |
<r2> (average value of r
2) Å
2
| <r2> |
275.858 |
| (<r2>)1/2 |
16.609 |