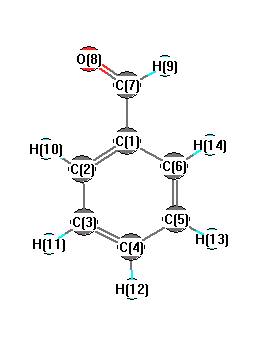
Vibrational Frequencies calculated at B2PLYP/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3245 |
3080 |
7.90 |
171.89 |
0.12 |
0.22 |
| 2 |
A' |
3237 |
3073 |
16.07 |
115.78 |
0.13 |
0.23 |
| 3 |
A' |
3228 |
3064 |
15.80 |
81.73 |
0.75 |
0.86 |
| 4 |
A' |
3216 |
3053 |
1.33 |
78.96 |
0.71 |
0.83 |
| 5 |
A' |
3203 |
3041 |
5.77 |
48.19 |
0.43 |
0.60 |
| 6 |
A' |
2950 |
2800 |
129.74 |
120.19 |
0.29 |
0.45 |
| 7 |
A' |
1780 |
1690 |
194.38 |
75.59 |
0.34 |
0.50 |
| 8 |
A' |
1669 |
1585 |
23.88 |
76.10 |
0.53 |
0.69 |
| 9 |
A' |
1652 |
1568 |
12.82 |
11.56 |
0.53 |
0.70 |
| 10 |
A' |
1549 |
1470 |
0.44 |
1.20 |
0.27 |
0.42 |
| 11 |
A' |
1510 |
1434 |
12.51 |
1.61 |
0.22 |
0.36 |
| 12 |
A' |
1448 |
1374 |
7.71 |
4.24 |
0.62 |
0.77 |
| 13 |
A' |
1394 |
1323 |
4.72 |
0.97 |
0.73 |
0.84 |
| 14 |
A' |
1357 |
1288 |
16.52 |
1.60 |
0.51 |
0.68 |
| 15 |
A' |
1246 |
1183 |
55.86 |
22.85 |
0.28 |
0.43 |
| 16 |
A' |
1210 |
1149 |
18.75 |
7.23 |
0.17 |
0.29 |
| 17 |
A' |
1205 |
1144 |
3.96 |
9.14 |
0.66 |
0.79 |
| 18 |
A' |
1118 |
1061 |
3.78 |
0.62 |
0.32 |
0.48 |
| 19 |
A' |
1057 |
1004 |
1.15 |
9.31 |
0.11 |
0.20 |
| 20 |
A' |
1022 |
970 |
0.51 |
24.18 |
0.10 |
0.19 |
| 21 |
A' |
847 |
804 |
30.68 |
10.05 |
0.16 |
0.28 |
| 22 |
A' |
663 |
630 |
22.47 |
3.07 |
0.37 |
0.54 |
| 23 |
A' |
631 |
599 |
0.51 |
5.23 |
0.75 |
0.86 |
| 24 |
A' |
444 |
422 |
0.18 |
5.37 |
0.35 |
0.52 |
| 25 |
A' |
222 |
211 |
7.77 |
0.62 |
0.56 |
0.72 |
| 26 |
A" |
1028 |
976 |
0.11 |
7.33 |
0.75 |
0.86 |
| 27 |
A" |
974 |
924 |
0.04 |
0.15 |
0.75 |
0.86 |
| 28 |
A" |
963 |
914 |
0.19 |
0.01 |
0.75 |
0.86 |
| 29 |
A" |
926 |
879 |
1.39 |
1.34 |
0.75 |
0.86 |
| 30 |
A" |
864 |
821 |
0.05 |
5.23 |
0.75 |
0.86 |
| 31 |
A" |
753 |
715 |
60.41 |
1.53 |
0.75 |
0.86 |
| 32 |
A" |
670 |
636 |
5.86 |
0.08 |
0.75 |
0.86 |
| 33 |
A" |
460 |
436 |
4.00 |
0.39 |
0.75 |
0.86 |
| 34 |
A" |
415 |
394 |
0.12 |
0.03 |
0.75 |
0.86 |
| 35 |
A" |
237 |
225 |
6.89 |
1.35 |
0.75 |
0.86 |
| 36 |
A" |
122 |
116 |
4.35 |
2.30 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 24258.0 cm
-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 23025.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.044 |
|
|
|
| 2 |
C |
-0.163 |
|
|
|
| 3 |
C |
-0.150 |
|
|
|
| 4 |
C |
-0.138 |
|
|
|
| 5 |
C |
-0.151 |
|
|
|
| 6 |
C |
-0.183 |
|
|
|
| 7 |
C |
0.201 |
|
|
|
| 8 |
O |
-0.401 |
|
|
|
| 9 |
H |
0.117 |
|
|
|
| 10 |
H |
0.184 |
|
|
|
| 11 |
H |
0.161 |
|
|
|
| 12 |
H |
0.160 |
|
|
|
| 13 |
H |
0.159 |
|
|
|
| 14 |
H |
0.160 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
2.226 |
-2.343 |
0.000 |
3.232 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-44.211 |
5.949 |
-0.002 |
| y |
5.949 |
-46.252 |
0.003 |
| z |
-0.002 |
0.003 |
-48.186 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.008 |
5.949 |
-0.002 |
| y |
5.949 |
-0.054 |
0.003 |
| z |
-0.002 |
0.003 |
-2.954 |
|
| Polar |
|---|
| 3z2-r2 | -5.908 |
|---|
| x2-y2 | 2.041 |
|---|
| xy | 5.949 |
|---|
| xz | -0.002 |
|---|
| yz | 0.003 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
12.404 |
-1.111 |
0.000 |
| y |
-1.111 |
14.630 |
-0.000 |
| z |
0.000 |
-0.000 |
3.822 |
<r2> (average value of r
2) Å
2
| <r2> |
261.867 |
| (<r2>)1/2 |
16.182 |