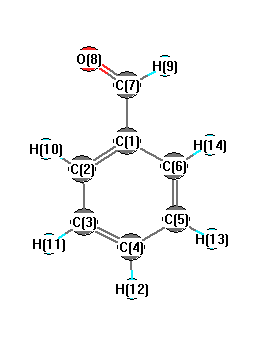
Vibrational Frequencies calculated at HSEh1PBE/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3242 |
3083 |
8.76 |
195.65 |
0.12 |
0.21 |
| 2 |
A' |
3235 |
3077 |
14.10 |
97.65 |
0.14 |
0.24 |
| 3 |
A' |
3226 |
3067 |
13.55 |
83.34 |
0.75 |
0.86 |
| 4 |
A' |
3214 |
3056 |
1.30 |
78.59 |
0.71 |
0.83 |
| 5 |
A' |
3201 |
3044 |
5.15 |
47.51 |
0.42 |
0.59 |
| 6 |
A' |
2935 |
2791 |
131.61 |
120.83 |
0.30 |
0.46 |
| 7 |
A' |
1835 |
1745 |
238.24 |
80.16 |
0.34 |
0.51 |
| 8 |
A' |
1684 |
1601 |
28.37 |
79.32 |
0.52 |
0.69 |
| 9 |
A' |
1667 |
1586 |
10.83 |
8.79 |
0.55 |
0.71 |
| 10 |
A' |
1548 |
1472 |
0.82 |
1.07 |
0.27 |
0.42 |
| 11 |
A' |
1510 |
1436 |
13.15 |
1.11 |
0.26 |
0.42 |
| 12 |
A' |
1442 |
1372 |
9.54 |
4.13 |
0.70 |
0.83 |
| 13 |
A' |
1399 |
1330 |
6.04 |
0.93 |
0.70 |
0.82 |
| 14 |
A' |
1348 |
1282 |
16.00 |
1.63 |
0.49 |
0.66 |
| 15 |
A' |
1249 |
1188 |
61.01 |
23.25 |
0.26 |
0.41 |
| 16 |
A' |
1201 |
1142 |
16.83 |
6.51 |
0.20 |
0.33 |
| 17 |
A' |
1196 |
1137 |
2.74 |
8.59 |
0.69 |
0.82 |
| 18 |
A' |
1116 |
1062 |
5.32 |
0.73 |
0.25 |
0.40 |
| 19 |
A' |
1061 |
1009 |
1.67 |
12.52 |
0.11 |
0.20 |
| 20 |
A' |
1023 |
973 |
0.20 |
20.99 |
0.10 |
0.18 |
| 21 |
A' |
851 |
809 |
30.51 |
9.57 |
0.15 |
0.27 |
| 22 |
A' |
664 |
632 |
23.60 |
2.84 |
0.38 |
0.55 |
| 23 |
A' |
629 |
598 |
0.51 |
5.35 |
0.75 |
0.86 |
| 24 |
A' |
445 |
423 |
0.17 |
5.47 |
0.36 |
0.53 |
| 25 |
A' |
218 |
207 |
7.91 |
0.58 |
0.55 |
0.71 |
| 26 |
A" |
1042 |
991 |
0.33 |
7.15 |
0.75 |
0.86 |
| 27 |
A" |
1015 |
966 |
0.01 |
0.14 |
0.75 |
0.86 |
| 28 |
A" |
991 |
943 |
0.11 |
0.01 |
0.75 |
0.86 |
| 29 |
A" |
945 |
899 |
1.73 |
0.81 |
0.75 |
0.86 |
| 30 |
A" |
872 |
830 |
0.04 |
4.66 |
0.75 |
0.86 |
| 31 |
A" |
766 |
728 |
48.41 |
1.53 |
0.75 |
0.86 |
| 32 |
A" |
707 |
672 |
21.79 |
0.17 |
0.75 |
0.86 |
| 33 |
A" |
465 |
442 |
4.59 |
0.43 |
0.75 |
0.86 |
| 34 |
A" |
419 |
398 |
0.13 |
0.03 |
0.75 |
0.86 |
| 35 |
A" |
240 |
229 |
7.23 |
1.23 |
0.75 |
0.86 |
| 36 |
A" |
125 |
119 |
4.07 |
2.37 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 24361.9 cm
-1
Scaled (by 0.951) Zero Point Vibrational Energy (zpe) 23168.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.041 |
|
|
|
| 2 |
C |
-0.174 |
|
|
|
| 3 |
C |
-0.169 |
|
|
|
| 4 |
C |
-0.153 |
|
|
|
| 5 |
C |
-0.170 |
|
|
|
| 6 |
C |
-0.199 |
|
|
|
| 7 |
C |
0.195 |
|
|
|
| 8 |
O |
-0.408 |
|
|
|
| 9 |
H |
0.128 |
|
|
|
| 10 |
H |
0.200 |
|
|
|
| 11 |
H |
0.178 |
|
|
|
| 12 |
H |
0.177 |
|
|
|
| 13 |
H |
0.177 |
|
|
|
| 14 |
H |
0.177 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
2.233 |
-2.423 |
0.000 |
3.295 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-43.491 |
5.946 |
0.000 |
| y |
5.946 |
-45.665 |
0.000 |
| z |
0.000 |
0.000 |
-48.097 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.390 |
5.946 |
0.000 |
| y |
5.946 |
0.129 |
0.000 |
| z |
0.000 |
0.000 |
-3.519 |
|
| Polar |
|---|
| 3z2-r2 | -7.037 |
|---|
| x2-y2 | 2.174 |
|---|
| xy | 5.946 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
12.419 |
-1.112 |
0.000 |
| y |
-1.112 |
14.750 |
0.000 |
| z |
0.000 |
0.000 |
3.882 |
<r2> (average value of r
2) Å
2
| <r2> |
260.375 |
| (<r2>)1/2 |
16.136 |