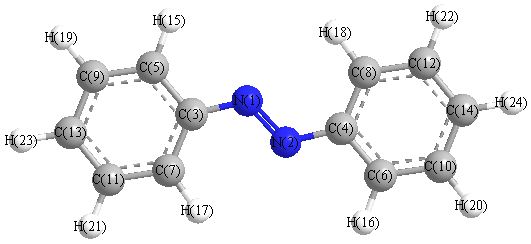
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3241 |
3100 |
0.00 |
|
|
|
| 2 |
Ag |
3227 |
3087 |
0.00 |
|
|
|
| 3 |
Ag |
3217 |
3078 |
0.00 |
|
|
|
| 4 |
Ag |
3206 |
3068 |
0.00 |
|
|
|
| 5 |
Ag |
3196 |
3058 |
0.00 |
|
|
|
| 6 |
Ag |
1673 |
1600 |
0.00 |
|
|
|
| 7 |
Ag |
1657 |
1585 |
0.00 |
|
|
|
| 8 |
Ag |
1578 |
1509 |
0.00 |
|
|
|
| 9 |
Ag |
1523 |
1457 |
0.00 |
|
|
|
| 10 |
Ag |
1498 |
1433 |
0.00 |
|
|
|
| 11 |
Ag |
1393 |
1332 |
0.00 |
|
|
|
| 12 |
Ag |
1346 |
1288 |
0.00 |
|
|
|
| 13 |
Ag |
1223 |
1170 |
0.00 |
|
|
|
| 14 |
Ag |
1191 |
1139 |
0.00 |
|
|
|
| 15 |
Ag |
1177 |
1126 |
0.00 |
|
|
|
| 16 |
Ag |
1106 |
1058 |
0.00 |
|
|
|
| 17 |
Ag |
1052 |
1007 |
0.00 |
|
|
|
| 18 |
Ag |
1018 |
974 |
0.00 |
|
|
|
| 19 |
Ag |
944 |
903 |
0.00 |
|
|
|
| 20 |
Ag |
682 |
653 |
0.00 |
|
|
|
| 21 |
Ag |
623 |
596 |
0.00 |
|
|
|
| 22 |
Ag |
307 |
294 |
0.00 |
|
|
|
| 23 |
Ag |
224 |
214 |
0.00 |
|
|
|
| 24 |
Au |
1007 |
964 |
1.04 |
|
|
|
| 25 |
Au |
987 |
944 |
0.05 |
|
|
|
| 26 |
Au |
953 |
912 |
9.45 |
|
|
|
| 27 |
Au |
863 |
825 |
0.01 |
|
|
|
| 28 |
Au |
800 |
765 |
64.74 |
|
|
|
| 29 |
Au |
705 |
674 |
67.36 |
|
|
|
| 30 |
Au |
560 |
536 |
9.45 |
|
|
|
| 31 |
Au |
416 |
398 |
0.01 |
|
|
|
| 32 |
Au |
308 |
294 |
0.73 |
|
|
|
| 33 |
Au |
64 |
61 |
1.33 |
|
|
|
| 34 |
Au |
26 |
24 |
0.04 |
|
|
|
| 35 |
Bg |
1007 |
963 |
0.00 |
|
|
|
| 36 |
Bg |
987 |
944 |
0.00 |
|
|
|
| 37 |
Bg |
948 |
907 |
0.00 |
|
|
|
| 38 |
Bg |
864 |
826 |
0.00 |
|
|
|
| 39 |
Bg |
780 |
746 |
0.00 |
|
|
|
| 40 |
Bg |
698 |
668 |
0.00 |
|
|
|
| 41 |
Bg |
489 |
468 |
0.00 |
|
|
|
| 42 |
Bg |
420 |
402 |
0.00 |
|
|
|
| 43 |
Bg |
257 |
246 |
0.00 |
|
|
|
| 44 |
Bg |
105 |
100 |
0.00 |
|
|
|
| 45 |
Bu |
3241 |
3100 |
10.19 |
|
|
|
| 46 |
Bu |
3227 |
3087 |
38.72 |
|
|
|
| 47 |
Bu |
3217 |
3078 |
49.92 |
|
|
|
| 48 |
Bu |
3206 |
3068 |
23.00 |
|
|
|
| 49 |
Bu |
3196 |
3057 |
2.65 |
|
|
|
| 50 |
Bu |
1668 |
1596 |
3.80 |
|
|
|
| 51 |
Bu |
1651 |
1579 |
4.90 |
|
|
|
| 52 |
Bu |
1536 |
1469 |
10.73 |
|
|
|
| 53 |
Bu |
1501 |
1436 |
13.29 |
|
|
|
| 54 |
Bu |
1389 |
1329 |
8.30 |
|
|
|
| 55 |
Bu |
1339 |
1281 |
1.70 |
|
|
|
| 56 |
Bu |
1275 |
1220 |
27.49 |
|
|
|
| 57 |
Bu |
1191 |
1139 |
0.15 |
|
|
|
| 58 |
Bu |
1184 |
1133 |
31.97 |
|
|
|
| 59 |
Bu |
1110 |
1062 |
13.66 |
|
|
|
| 60 |
Bu |
1052 |
1007 |
11.44 |
|
|
|
| 61 |
Bu |
1017 |
973 |
3.64 |
|
|
|
| 62 |
Bu |
844 |
807 |
0.65 |
|
|
|
| 63 |
Bu |
630 |
602 |
0.68 |
|
|
|
| 64 |
Bu |
544 |
521 |
6.01 |
|
|
|
| 65 |
Bu |
531 |
508 |
25.42 |
|
|
|
| 66 |
Bu |
85 |
81 |
1.82 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 42085.0 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 40262.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.311 |
|
|
|
| 2 |
N |
-0.311 |
|
|
|
| 3 |
C |
0.257 |
|
|
|
| 4 |
C |
0.257 |
|
|
|
| 5 |
C |
-0.167 |
|
|
|
| 6 |
C |
-0.167 |
|
|
|
| 7 |
C |
-0.146 |
|
|
|
| 8 |
C |
-0.146 |
|
|
|
| 9 |
C |
-0.174 |
|
|
|
| 10 |
C |
-0.174 |
|
|
|
| 11 |
C |
-0.181 |
|
|
|
| 12 |
C |
-0.181 |
|
|
|
| 13 |
C |
-0.150 |
|
|
|
| 14 |
C |
-0.150 |
|
|
|
| 15 |
H |
0.177 |
|
|
|
| 16 |
H |
0.177 |
|
|
|
| 17 |
H |
0.190 |
|
|
|
| 18 |
H |
0.190 |
|
|
|
| 19 |
H |
0.169 |
|
|
|
| 20 |
H |
0.169 |
|
|
|
| 21 |
H |
0.170 |
|
|
|
| 22 |
H |
0.170 |
|
|
|
| 23 |
H |
0.168 |
|
|
|
| 24 |
H |
0.168 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-64.705 |
5.067 |
0.000 |
| y |
5.067 |
-67.266 |
0.000 |
| z |
0.000 |
0.000 |
-84.969 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
11.412 |
5.067 |
0.000 |
| y |
5.067 |
7.571 |
0.000 |
| z |
0.000 |
0.000 |
-18.983 |
|
| Polar |
|---|
| 3z2-r2 | -37.967 |
|---|
| x2-y2 | 2.561 |
|---|
| xy | 5.067 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
34.024 |
11.000 |
0.000 |
| y |
11.000 |
29.620 |
0.000 |
| z |
0.000 |
0.000 |
6.717 |
<r2> (average value of r
2) Å
2
| <r2> |
1092.733 |
| (<r2>)1/2 |
33.057 |