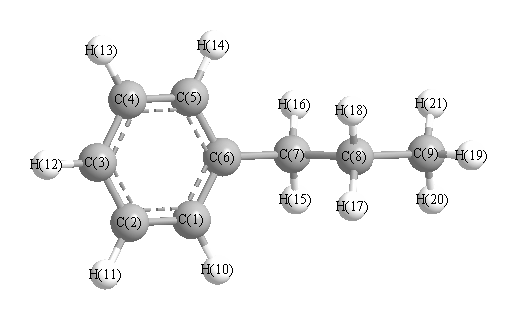
Vibrational Frequencies calculated at PBEPBEultrafine/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3138 |
3086 |
21.72 |
|
|
|
| 2 |
A' |
3116 |
3064 |
4.92 |
|
|
|
| 3 |
A' |
3100 |
3049 |
11.59 |
|
|
|
| 4 |
A' |
3058 |
3008 |
33.05 |
|
|
|
| 5 |
A' |
2979 |
2930 |
62.86 |
|
|
|
| 6 |
A' |
2975 |
2926 |
19.46 |
|
|
|
| 7 |
A' |
2962 |
2914 |
10.16 |
|
|
|
| 8 |
A' |
1622 |
1595 |
6.03 |
|
|
|
| 9 |
A' |
1500 |
1476 |
10.93 |
|
|
|
| 10 |
A' |
1491 |
1466 |
5.48 |
|
|
|
| 11 |
A' |
1475 |
1450 |
0.20 |
|
|
|
| 12 |
A' |
1463 |
1439 |
0.33 |
|
|
|
| 13 |
A' |
1388 |
1365 |
1.11 |
|
|
|
| 14 |
A' |
1348 |
1325 |
2.17 |
|
|
|
| 15 |
A' |
1279 |
1258 |
2.15 |
|
|
|
| 16 |
A' |
1205 |
1185 |
1.52 |
|
|
|
| 17 |
A' |
1179 |
1159 |
0.12 |
|
|
|
| 18 |
A' |
1088 |
1071 |
9.47 |
|
|
|
| 19 |
A' |
1039 |
1022 |
1.56 |
|
|
|
| 20 |
A' |
1034 |
1016 |
1.43 |
|
|
|
| 21 |
A' |
989 |
972 |
0.29 |
|
|
|
| 22 |
A' |
951 |
935 |
0.03 |
|
|
|
| 23 |
A' |
895 |
881 |
0.18 |
|
|
|
| 24 |
A' |
877 |
863 |
2.17 |
|
|
|
| 25 |
A' |
806 |
793 |
0.25 |
|
|
|
| 26 |
A' |
738 |
726 |
25.56 |
|
|
|
| 27 |
A' |
691 |
680 |
26.45 |
|
|
|
| 28 |
A' |
579 |
569 |
3.78 |
|
|
|
| 29 |
A' |
492 |
484 |
5.80 |
|
|
|
| 30 |
A' |
303 |
298 |
0.05 |
|
|
|
| 31 |
A' |
270 |
266 |
0.56 |
|
|
|
| 32 |
A' |
99 |
98 |
0.24 |
|
|
|
| 33 |
A" |
3125 |
3073 |
40.04 |
|
|
|
| 34 |
A" |
3101 |
3050 |
7.65 |
|
|
|
| 35 |
A" |
3051 |
3000 |
48.53 |
|
|
|
| 36 |
A" |
3019 |
2970 |
18.03 |
|
|
|
| 37 |
A" |
2997 |
2948 |
0.98 |
|
|
|
| 38 |
A" |
1601 |
1575 |
0.78 |
|
|
|
| 39 |
A" |
1481 |
1456 |
6.70 |
|
|
|
| 40 |
A" |
1456 |
1432 |
5.16 |
|
|
|
| 41 |
A" |
1369 |
1346 |
0.16 |
|
|
|
| 42 |
A" |
1323 |
1301 |
0.53 |
|
|
|
| 43 |
A" |
1292 |
1271 |
0.16 |
|
|
|
| 44 |
A" |
1223 |
1203 |
0.26 |
|
|
|
| 45 |
A" |
1158 |
1139 |
0.04 |
|
|
|
| 46 |
A" |
1103 |
1085 |
4.49 |
|
|
|
| 47 |
A" |
1049 |
1032 |
0.76 |
|
|
|
| 48 |
A" |
922 |
906 |
0.01 |
|
|
|
| 49 |
A" |
858 |
844 |
0.99 |
|
|
|
| 50 |
A" |
826 |
813 |
0.03 |
|
|
|
| 51 |
A" |
732 |
720 |
1.86 |
|
|
|
| 52 |
A" |
616 |
606 |
0.02 |
|
|
|
| 53 |
A" |
400 |
394 |
0.00 |
|
|
|
| 54 |
A" |
336 |
330 |
0.04 |
|
|
|
| 55 |
A" |
242 |
238 |
0.01 |
|
|
|
| 56 |
A" |
88 |
87 |
0.01 |
|
|
|
| 57 |
A" |
35 |
34 |
0.02 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 39764.4 cm
-1
Scaled (by 0.9835) Zero Point Vibrational Energy (zpe) 39108.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.184 |
|
|
|
| 2 |
C |
-0.138 |
|
|
|
| 3 |
C |
-0.139 |
|
|
|
| 4 |
C |
-0.138 |
|
|
|
| 5 |
C |
-0.184 |
|
|
|
| 6 |
C |
0.176 |
|
|
|
| 7 |
C |
-0.380 |
|
|
|
| 8 |
C |
-0.274 |
|
|
|
| 9 |
C |
-0.482 |
|
|
|
| 10 |
H |
0.128 |
|
|
|
| 11 |
H |
0.135 |
|
|
|
| 12 |
H |
0.134 |
|
|
|
| 13 |
H |
0.135 |
|
|
|
| 14 |
H |
0.128 |
|
|
|
| 15 |
H |
0.153 |
|
|
|
| 16 |
H |
0.153 |
|
|
|
| 17 |
H |
0.154 |
|
|
|
| 18 |
H |
0.154 |
|
|
|
| 19 |
H |
0.160 |
|
|
|
| 20 |
H |
0.156 |
|
|
|
| 21 |
H |
0.156 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.132 |
0.333 |
0.000 |
0.358 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-58.053 |
-0.145 |
0.000 |
| y |
-0.145 |
-51.526 |
0.000 |
| z |
0.000 |
0.000 |
-51.016 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-6.782 |
-0.145 |
0.000 |
| y |
-0.145 |
3.009 |
0.000 |
| z |
0.000 |
0.000 |
3.773 |
|
| Polar |
|---|
| 3z2-r2 | 7.547 |
|---|
| x2-y2 | -6.527 |
|---|
| xy | -0.145 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.944 |
-1.136 |
0.000 |
| y |
-1.136 |
18.733 |
0.000 |
| z |
0.000 |
0.000 |
14.394 |
<r2> (average value of r
2) Å
2
| <r2> |
442.837 |
| (<r2>)1/2 |
21.044 |