return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H7 (cyclopentenyl radical)

using model chemistry: B2PLYP/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 2A2
Energy calculated at B2PLYP/6-31G*
 hartrees
Energy at 0K-194.434621
Energy at 298.15K-194.441843
HF Energy-194.232055
Nuclear repulsion energy163.346823
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A1 3273 3107 8.05      
2 A1 3218 3055 13.83      
3 A1 3061 2905 38.04      
4 A1 1544 1466 2.02      
5 A1 1525 1448 0.73      
6 A1 1316 1249 0.07      
7 A1 1129 1071 2.64      
8 A1 1057 1004 0.00      
9 A1 930 883 2.46      
10 A1 819 778 0.24      
11 A2 3070 2914 0.00      
12 A2 1255 1192 0.00      
13 A2 1019 967 0.00      
14 A2 605 574 0.00      
15 A2 110 105 0.00      
16 B1 3090 2933 53.12      
17 B1 1167 1108 2.13      
18 B1 958 909 4.46      
19 B1 829 787 1.01      
20 B1 608 577 49.41      
21 B1 420 399 9.05      
22 B2 3264 3098 11.51      
23 B2 3049 2894 36.64      
24 B2 1520 1443 0.29      
25 B2 1445 1372 3.70      
26 B2 1342 1273 1.20      
27 B2 1332 1264 5.63      
28 B2 1101 1045 0.91      
29 B2 930 883 12.66      
30 B2 730 693 0.33      

Unscaled Zero Point Vibrational Energy (zpe) 22858.2 cm-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 21697.0 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at B2PLYP/6-31G*
ABC
0.26156 0.25479 0.13548

See section I.F.4 to change rotational constant units
Geometric Data calculated at B2PLYP/6-31G*

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.000 1.267
C2 0.000 1.146 0.505
C3 0.000 -1.146 0.505
C4 0.000 0.773 -0.963
C5 0.000 -0.773 -0.963
H6 0.000 0.000 2.354
H7 0.000 2.206 0.725
H8 0.000 -2.206 0.725
H9 0.876 1.186 -1.480
H10 -0.876 1.186 -1.480
H11 0.876 -1.186 -1.480
H12 -0.876 -1.186 -1.480

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 H6 H7 H8 H9 H10 H11 H12
C11.37641.37642.36062.36061.08642.27142.27143.11793.11793.11793.1179
C21.37642.29231.51492.41672.17491.08223.35912.17032.17033.18553.1855
C31.37642.29232.41671.51492.17493.35911.08223.18553.18552.17032.1703
C42.36061.51492.41671.54663.40572.21413.42421.09771.09772.20762.2076
C52.36062.41671.51491.54663.40573.42422.21412.20762.20761.09771.0977
H61.08642.17492.17493.40573.40572.74182.74184.10744.10744.10744.1074
H72.27141.08223.35912.21413.42422.74184.41152.58262.58264.13934.1393
H82.27143.35911.08223.42422.21412.74184.41154.13934.13932.58262.5826
H93.11792.17033.18551.09772.20764.10742.58264.13931.75202.37202.9489
H103.11792.17033.18551.09772.20764.10742.58264.13931.75202.94892.3720
H113.11793.18552.17032.20761.09774.10744.13932.58262.37202.94891.7520
H123.11793.18552.17032.20761.09774.10744.13932.58262.94892.37201.7520

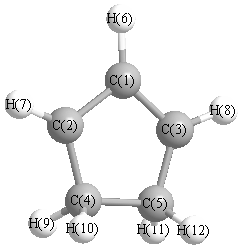
picture of cyclopentenyl radical state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C4 109.372 C1 C2 H7 134.653
C1 C3 C5 109.372 C1 C3 H8 134.653
C2 C1 C3 112.760 C2 C1 H6 123.620
C2 C4 C5 104.248 C2 C4 H9 111.338
C2 C4 H10 111.338 C3 C1 H6 123.620
C3 C5 C4 104.248 C3 C5 H11 111.338
C3 C5 H12 111.338 C4 C2 H7 115.975
C4 C5 H11 112.086 C4 C5 H12 112.086
C5 C3 H8 115.975 C5 C4 H9 112.086
C5 C4 H10 112.086 H9 C4 H10 105.888
H11 C5 H12 105.888
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G* Charges (e)
Number Element Mulliken CHELPG AIM ESP
1 C -0.116      
2 C -0.165      
3 C -0.165      
4 C -0.324      
5 C -0.324      
6 H 0.153      
7 H 0.159      
8 H 0.159      
9 H 0.155      
10 H 0.155      
11 H 0.155      
12 H 0.155      


Electric dipole moments


Electric Quadrupole moment
Quadrupole components in D Å


Polarizabilities
Components of the polarizability tensor.
Units are Å3 (Angstrom cubed)
Change units.
  x y z
x 4.449 0.000 0.000
y 0.000 8.875 0.000
z 0.000 0.000 7.697


<r2> (average value of r2) Å2
<r2> 99.513
(<r2>)1/2 9.976