return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H4Cl2 (1,4-dichlorobenzene)

using model chemistry: CID/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes D2H 1AG
Energy calculated at CID/6-31G*
 hartrees
Energy at 0K-1149.329369
Energy at 298.15K-1149.333780
HF Energy-1148.501733
Nuclear repulsion energy451.316879
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CID/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 Ag 3361 3105 0.00      
2 Ag 1758 1624 0.00      
3 Ag 1275 1177 0.00      
4 Ag 1193 1102 0.00      
5 Ag 806 744 0.00      
6 Ag 350 323 0.00      
7 Au 1013 936 0.00      
8 Au 438 405 0.00      
9 B1g 889 821 0.00      
10 B1u 3345 3090 1.50      
11 B1u 1618 1494 114.00      
12 B1u 1193 1102 91.91      
13 B1u 1079 997 55.20      
14 B1u 586 542 27.71      
15 B2g 974 900 0.00      
16 B2g 673 622 0.00      
17 B2g 317 292 0.00      
18 B2u 3359 3103 2.05      
19 B2u 1518 1403 8.53      
20 B2u 1304 1205 0.56      
21 B2u 1193 1102 3.07      
22 B2u 234 216 1.19      
23 B3g 3346 3091 0.00      
24 B3g 1739 1606 0.00      
25 B3g 1397 1291 0.00      
26 B3g 673 622 0.00      
27 B3g 377 348 0.00      
28 B3u 879 812 63.99      
29 B3u 521 482 15.79      
30 B3u 109 101 0.74      

Unscaled Zero Point Vibrational Energy (zpe) 18758.2 cm-1
Scaled (by 0.9237) Zero Point Vibrational Energy (zpe) 17326.9 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at CID/6-31G*
ABC
0.19184 0.02244 0.02009

See section I.F.4 to change rotational constant units
Geometric Data calculated at CID/6-31G*

Point Group is D2h

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.000 1.375
C2 0.000 0.000 -1.375
C3 0.000 1.203 0.692
C4 0.000 -1.203 0.692
C5 0.000 -1.203 -0.692
C6 0.000 1.203 -0.692
Cl7 0.000 0.000 3.110
Cl8 0.000 0.000 -3.110
H9 0.000 2.134 1.236
H10 0.000 -2.134 1.236
H11 0.000 -2.134 -1.236
H12 0.000 2.134 -1.236

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 Cl7 Cl8 H9 H10 H11 H12
C12.75081.38391.38392.39242.39241.73504.48582.13852.13853.37223.3722
C22.75082.39242.39241.38391.38394.48581.73503.37223.37222.13852.1385
C31.38392.39242.40692.77671.38442.70113.98851.07763.38133.85422.1407
C41.38392.39242.40691.38442.77672.70113.98853.38131.07762.14073.8542
C52.39241.38392.77671.38442.40693.98852.70113.85422.14071.07763.3813
C62.39241.38391.38442.77672.40693.98852.70112.14073.85423.38131.0776
Cl71.73504.48582.70112.70113.98853.98856.22072.84042.84044.84174.8417
Cl84.48581.73503.98853.98852.70112.70116.22074.84174.84172.84042.8404
H92.13853.37221.07763.38133.85422.14072.84044.84174.26784.93182.4714
H102.13853.37223.38131.07762.14073.85422.84044.84174.26782.47144.9318
H113.37222.13853.85422.14071.07763.38134.84172.84044.93182.47144.2678
H123.37222.13852.14073.85423.38131.07764.84172.84042.47144.93184.2678

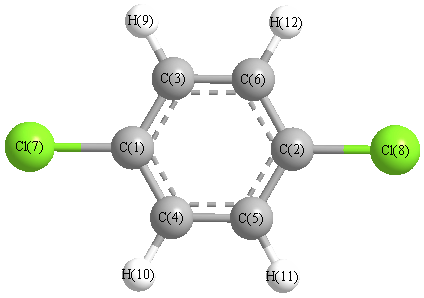
picture of 1,4-dichlorobenzene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C3 C6 119.583 C1 C3 H9 120.127
C1 C4 C5 119.583 C1 C4 H10 120.127
C2 C5 C4 119.583 C2 C5 H11 120.127
C2 C6 C3 119.583 C2 C6 H12 120.127
C3 C1 C4 120.833 C3 C1 Cl7 119.583
C3 C6 H12 120.290 C4 C1 Cl7 119.583
C4 C5 H11 120.290 C5 C2 C6 120.833
C5 C2 Cl8 119.583 C5 C4 H10 120.290
C6 C2 Cl8 119.583 C6 C3 H9 120.290
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability