return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H4O2 (parabenzoquinone)

using model chemistry: B2PLYP/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes D2H 1Ag
Energy calculated at B2PLYP/6-31G*
 hartrees
Energy at 0K-381.074838
Energy at 298.15K-381.079225
HF Energy-380.706125
Nuclear repulsion energy321.914823
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 Ag 3250 3085 0.00      
2 Ag 1738 1650 0.00      
3 Ag 1678 1593 0.00      
4 Ag 1183 1123 0.00      
5 Ag 784 744 0.00      
6 Ag 455 432 0.00      
7 Au 1000 949 0.00      
8 Au 333 316 0.00      
9 B1g 760 722 0.00      
10 B1u 3232 3068 1.96      
11 B1u 1735 1647 269.85      
12 B1u 1404 1333 5.15      
13 B1u 958 909 16.10      
14 B1u 761 722 0.48      
15 B2g 1003 952 0.00      
16 B2g 765 726 0.00      
17 B2g 227 216 0.00      
18 B2u 3248 3083 6.04      
19 B2u 1657 1572 17.67      
20 B2u 1346 1277 79.08      
21 B2u 1100 1044 46.52      
22 B2u 411 390 26.07      
23 B3g 3232 3068 0.00      
24 B3g 1423 1350 0.00      
25 B3g 1257 1193 0.00      
26 B3g 605 574 0.00      
27 B3g 453 430 0.00      
28 B3u 900 854 80.77      
29 B3u 512 486 2.02      
30 B3u 96 91 12.92      

Unscaled Zero Point Vibrational Energy (zpe) 18751.4 cm-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 17798.8 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at B2PLYP/6-31G*
ABC
0.17505 0.05499 0.04184

See section I.F.4 to change rotational constant units
Geometric Data calculated at B2PLYP/6-31G*

Point Group is D2h

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.000 1.443
C2 0.000 0.000 -1.443
C3 0.000 1.268 0.673
C4 0.000 -1.268 0.673
C5 0.000 -1.268 -0.673
C6 0.000 1.268 -0.673
O7 0.000 0.000 2.672
O8 0.000 0.000 -2.672
H9 0.000 2.181 1.259
H10 0.000 -2.181 1.259
H11 0.000 -2.181 -1.259
H12 0.000 2.181 -1.259

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 O7 O8 H9 H10 H11 H12
C12.88631.48351.48352.46642.46641.22934.11572.18852.18853.47233.4723
C22.88632.46642.46641.48351.48354.11571.22933.47233.47232.18852.1885
C31.48352.46642.53532.87011.34522.36783.57721.08513.49783.95242.1364
C41.48352.46642.53531.34522.87012.36783.57723.49781.08512.13643.9524
C52.46641.48352.87011.34522.53533.57722.36783.95242.13641.08513.4978
C62.46641.48351.34522.87012.53533.57722.36782.13643.95243.49781.0851
O71.22934.11572.36782.36783.57723.57725.34502.59882.59884.49574.4957
O84.11571.22933.57723.57722.36782.36785.34504.49574.49572.59882.5988
H92.18853.47231.08513.49783.95242.13642.59884.49574.36145.03602.5178
H102.18853.47233.49781.08512.13643.95242.59884.49574.36142.51785.0360
H113.47232.18853.95242.13641.08513.49784.49572.59885.03602.51784.3614
H123.47232.18852.13643.95243.49781.08514.49572.59882.51785.03604.3614

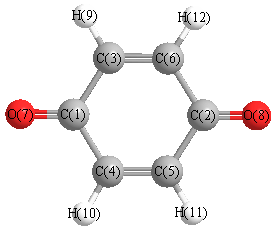
picture of parabenzoquinone state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C3 C6 121.295 C1 C3 H9 115.997
C1 C4 C5 121.295 C1 C4 H10 115.997
C2 C5 C4 121.295 C2 C5 H11 115.997
C2 C6 C3 121.295 C2 C6 H12 115.997
C3 C1 C4 117.411 C3 C1 O7 121.295
C3 C6 H12 122.708 C4 C1 O7 121.295
C4 C5 H11 122.708 C5 C2 C6 117.411
C5 C2 O8 121.295 C5 C4 H10 122.708
C6 C2 O8 121.295 C6 C3 H9 122.708
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G* Charges (e)
Number Element Mulliken CHELPG AIM ESP
1 C 0.465      
2 C 0.465      
3 C -0.192      
4 C -0.192      
5 C -0.192      
6 C -0.192      
7 O -0.495      
8 O -0.495      
9 H 0.207      
10 H 0.207      
11 H 0.207      
12 H 0.207      


Electric dipole moments


Electric Quadrupole moment
Quadrupole components in D Å


Polarizabilities
Components of the polarizability tensor.
Units are Å3 (Angstrom cubed)
Change units.
  x y z
x 3.633 0.000 0.000
y 0.000 8.760 0.000
z 0.000 0.000 15.190


<r2> (average value of r2) Å2
<r2> 244.036
(<r2>)1/2 15.622