Jump to
S1C2
Energy calculated at CID/6-31G*
| | hartrees |
|---|
| Energy at 0K | -5218.294994 |
| Energy at 298.15K | -5218.305265 |
| HF Energy | -5217.847284 |
| Nuclear repulsion energy | 417.564794 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CID/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3238 |
2991 |
0.00 |
|
|
|
| 2 |
Ag |
1579 |
1459 |
0.00 |
|
|
|
| 3 |
Ag |
1380 |
1274 |
0.00 |
|
|
|
| 4 |
Ag |
1126 |
1041 |
0.00 |
|
|
|
| 5 |
Ag |
719 |
664 |
0.00 |
|
|
|
| 6 |
Ag |
201 |
185 |
0.00 |
|
|
|
| 7 |
Au |
3323 |
3070 |
0.47 |
|
|
|
| 8 |
Au |
1169 |
1080 |
3.20 |
|
|
|
| 9 |
Au |
788 |
728 |
3.82 |
|
|
|
| 10 |
Au |
108 |
100 |
5.23 |
|
|
|
| 11 |
Bg |
3303 |
3051 |
0.00 |
|
|
|
| 12 |
Bg |
1360 |
1257 |
0.00 |
|
|
|
| 13 |
Bg |
999 |
922 |
0.00 |
|
|
|
| 14 |
Bu |
3243 |
2995 |
6.70 |
|
|
|
| 15 |
Bu |
1572 |
1452 |
8.28 |
|
|
|
| 16 |
Bu |
1287 |
1189 |
65.30 |
|
|
|
| 17 |
Bu |
636 |
588 |
67.93 |
|
|
|
| 18 |
Bu |
192 |
177 |
6.96 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13111.9 cm
-1
Scaled (by 0.9237) Zero Point Vibrational Energy (zpe) 12111.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CID/6-31G*
Point Group is C2h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.493 |
0.571 |
0.000 |
| C2 |
-0.493 |
-0.571 |
0.000 |
| Br3 |
-0.493 |
2.263 |
0.000 |
| Br4 |
0.493 |
-2.263 |
0.000 |
| H5 |
1.113 |
0.572 |
0.888 |
| H6 |
1.113 |
0.572 |
-0.888 |
| H7 |
-1.113 |
-0.572 |
0.888 |
| H8 |
-1.113 |
-0.572 |
-0.888 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Br3 |
Br4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5089 | 1.9578 | 2.8343 | 1.0836 | 1.0836 | 2.1623 | 2.1623 |
C2 | 1.5089 | | 2.8343 | 1.9578 | 2.1623 | 2.1623 | 1.0836 | 1.0836 | Br3 | 1.9578 | 2.8343 | | 4.6320 | 2.4957 | 2.4957 | 3.0349 | 3.0349 | Br4 | 2.8343 | 1.9578 | 4.6320 | | 3.0349 | 3.0349 | 2.4957 | 2.4957 | H5 | 1.0836 | 2.1623 | 2.4957 | 3.0349 | | 1.7763 | 2.5035 | 3.0696 | H6 | 1.0836 | 2.1623 | 2.4957 | 3.0349 | 1.7763 | | 3.0696 | 2.5035 | H7 | 2.1623 | 1.0836 | 3.0349 | 2.4957 | 2.5035 | 3.0696 | | 1.7763 | H8 | 2.1623 | 1.0836 | 3.0349 | 2.4957 | 3.0696 | 2.5035 | 1.7763 | |
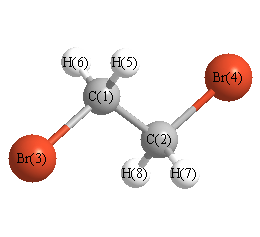 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
Br4 |
109.000 |
|
C1 |
C2 |
H7 |
111.999 |
| C1 |
C2 |
H8 |
111.999 |
|
C2 |
C1 |
Br3 |
109.000 |
| C2 |
C1 |
H5 |
111.999 |
|
C2 |
C1 |
H6 |
111.999 |
| Br3 |
C1 |
H5 |
106.728 |
|
Br3 |
C1 |
H6 |
106.728 |
| Br4 |
C2 |
H7 |
106.728 |
|
Br4 |
C2 |
H8 |
106.728 |
| H5 |
C1 |
H6 |
110.103 |
|
H7 |
C2 |
H8 |
110.103 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at CID/6-31G*
| | hartrees |
|---|
| Energy at 0K | -5218.292462 |
| Energy at 298.15K | |
| HF Energy | -5217.843920 |
| Nuclear repulsion energy | 455.239398 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CID/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3286 |
3035 |
0.04 |
|
|
|
| 2 |
A |
3221 |
2975 |
14.92 |
|
|
|
| 3 |
A |
1565 |
1445 |
0.06 |
|
|
|
| 4 |
A |
1394 |
1288 |
27.39 |
|
|
|
| 5 |
A |
1268 |
1171 |
1.71 |
|
|
|
| 6 |
A |
1092 |
1008 |
0.24 |
|
|
|
| 7 |
A |
966 |
892 |
6.88 |
|
|
|
| 8 |
A |
592 |
546 |
10.55 |
|
|
|
| 9 |
A |
239 |
221 |
1.12 |
|
|
|
| 10 |
A |
92 |
85 |
0.26 |
|
|
|
| 11 |
B |
3297 |
3046 |
1.14 |
|
|
|
| 12 |
B |
3211 |
2966 |
1.89 |
|
|
|
| 13 |
B |
1555 |
1437 |
10.80 |
|
|
|
| 14 |
B |
1364 |
1260 |
62.23 |
|
|
|
| 15 |
B |
1187 |
1097 |
2.42 |
|
|
|
| 16 |
B |
897 |
829 |
15.46 |
|
|
|
| 17 |
B |
627 |
580 |
12.76 |
|
|
|
| 18 |
B |
376 |
348 |
6.97 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13114.4 cm
-1
Scaled (by 0.9237) Zero Point Vibrational Energy (zpe) 12113.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CID/6-31G*
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.309 |
0.689 |
1.202 |
| C2 |
-0.309 |
-0.689 |
1.202 |
| Br3 |
-0.309 |
1.768 |
-0.298 |
| Br4 |
0.309 |
-1.768 |
-0.298 |
| H5 |
0.018 |
1.226 |
2.101 |
| H6 |
1.390 |
0.646 |
1.131 |
| H7 |
-0.018 |
-1.226 |
2.101 |
| H8 |
-1.390 |
-0.646 |
1.131 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Br3 |
Br4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5101 | 1.9483 | 2.8790 | 1.0867 | 1.0841 | 2.1407 | 2.1615 |
C2 | 1.5101 | | 2.8790 | 1.9483 | 2.1407 | 2.1615 | 1.0867 | 1.0841 | Br3 | 1.9483 | 2.8790 | | 3.5898 | 2.4812 | 2.4873 | 3.8478 | 3.0070 | Br4 | 2.8790 | 1.9483 | 3.5898 | | 3.8478 | 3.0070 | 2.4812 | 2.4873 | H5 | 1.0867 | 2.1407 | 2.4812 | 3.8478 | | 1.7767 | 2.4523 | 2.5351 | H6 | 1.0841 | 2.1615 | 2.4873 | 3.0070 | 1.7767 | | 2.5351 | 3.0650 | H7 | 2.1407 | 1.0867 | 3.8478 | 2.4812 | 2.4523 | 2.5351 | | 1.7767 | H8 | 2.1615 | 1.0841 | 3.0070 | 2.4873 | 2.5351 | 3.0650 | 1.7767 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
Br4 |
112.086 |
|
C1 |
C2 |
H7 |
109.979 |
| C1 |
C2 |
H8 |
111.811 |
|
C2 |
C1 |
Br3 |
112.086 |
| C2 |
C1 |
H5 |
109.979 |
|
C2 |
C1 |
H6 |
111.811 |
| Br3 |
C1 |
H5 |
106.176 |
|
Br3 |
C1 |
H6 |
106.720 |
| Br4 |
C2 |
H7 |
106.176 |
|
Br4 |
C2 |
H8 |
106.720 |
| H5 |
C1 |
H6 |
109.869 |
|
H7 |
C2 |
H8 |
109.869 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
