Jump to
S1C2
Energy calculated at QCISD/6-31G*
| | hartrees |
|---|
| Energy at 0K | -5218.375397 |
| Energy at 298.15K | -5218.385586 |
| HF Energy | -5217.846720 |
| Nuclear repulsion energy | 415.143712 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3151 |
3000 |
0.00 |
|
|
|
| 2 |
Ag |
1539 |
1466 |
0.00 |
|
|
|
| 3 |
Ag |
1333 |
1269 |
0.00 |
|
|
|
| 4 |
Ag |
1102 |
1049 |
0.00 |
|
|
|
| 5 |
Ag |
691 |
658 |
0.00 |
|
|
|
| 6 |
Ag |
194 |
185 |
0.00 |
|
|
|
| 7 |
Au |
3234 |
3079 |
1.10 |
|
|
|
| 8 |
Au |
1130 |
1076 |
3.17 |
|
|
|
| 9 |
Au |
770 |
734 |
3.67 |
|
|
|
| 10 |
Au |
105 |
100 |
4.95 |
|
|
|
| 11 |
Bg |
3214 |
3061 |
0.00 |
|
|
|
| 12 |
Bg |
1322 |
1259 |
0.00 |
|
|
|
| 13 |
Bg |
969 |
922 |
0.00 |
|
|
|
| 14 |
Bu |
3156 |
3005 |
6.83 |
|
|
|
| 15 |
Bu |
1532 |
1459 |
7.13 |
|
|
|
| 16 |
Bu |
1243 |
1184 |
58.45 |
|
|
|
| 17 |
Bu |
609 |
580 |
59.79 |
|
|
|
| 18 |
Bu |
186 |
178 |
6.78 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 12739.1 cm
-1
Scaled (by 0.9523) Zero Point Vibrational Energy (zpe) 12131.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD/6-31G*
Point Group is C2h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.497 |
0.571 |
0.000 |
| C2 |
-0.497 |
-0.571 |
0.000 |
| Br3 |
-0.497 |
2.275 |
0.000 |
| Br4 |
0.497 |
-2.275 |
0.000 |
| H5 |
1.120 |
0.572 |
0.895 |
| H6 |
1.120 |
0.572 |
-0.895 |
| H7 |
-1.120 |
-0.572 |
0.895 |
| H8 |
-1.120 |
-0.572 |
-0.895 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Br3 |
Br4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5133 | 1.9728 | 2.8460 | 1.0906 | 1.0906 | 2.1729 | 2.1729 |
C2 | 1.5133 | | 2.8460 | 1.9728 | 2.1729 | 2.1729 | 1.0906 | 1.0906 | Br3 | 1.9728 | 2.8460 | | 4.6576 | 2.5131 | 2.5131 | 3.0491 | 3.0491 | Br4 | 2.8460 | 1.9728 | 4.6576 | | 3.0491 | 3.0491 | 2.5131 | 2.5131 | H5 | 1.0906 | 2.1729 | 2.5131 | 3.0491 | | 1.7894 | 2.5160 | 3.0874 | H6 | 1.0906 | 2.1729 | 2.5131 | 3.0491 | 1.7894 | | 3.0874 | 2.5160 | H7 | 2.1729 | 1.0906 | 3.0491 | 2.5131 | 2.5160 | 3.0874 | | 1.7894 | H8 | 2.1729 | 1.0906 | 3.0491 | 2.5131 | 3.0874 | 2.5160 | 1.7894 | |
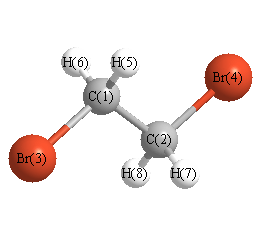 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
Br4 |
108.733 |
|
C1 |
C2 |
H7 |
112.110 |
| C1 |
C2 |
H8 |
112.110 |
|
C2 |
C1 |
Br3 |
108.733 |
| C2 |
C1 |
H5 |
112.110 |
|
C2 |
C1 |
H6 |
112.110 |
| Br3 |
C1 |
H5 |
106.667 |
|
Br3 |
C1 |
H6 |
106.667 |
| Br4 |
C2 |
H7 |
106.667 |
|
Br4 |
C2 |
H8 |
106.667 |
| H5 |
C1 |
H6 |
110.243 |
|
H7 |
C2 |
H8 |
110.243 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at QCISD/6-31G*
| | hartrees |
|---|
| Energy at 0K | -5218.373169 |
| Energy at 298.15K | |
| HF Energy | -5217.843318 |
| Nuclear repulsion energy | 453.528444 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3196 |
3044 |
0.12 |
|
|
|
| 2 |
A |
3132 |
2982 |
15.44 |
|
|
|
| 3 |
A |
1524 |
1451 |
0.07 |
|
|
|
| 4 |
A |
1350 |
1285 |
24.90 |
|
|
|
| 5 |
A |
1228 |
1169 |
1.91 |
|
|
|
| 6 |
A |
1064 |
1013 |
0.22 |
|
|
|
| 7 |
A |
940 |
895 |
5.99 |
|
|
|
| 8 |
A |
568 |
541 |
9.17 |
|
|
|
| 9 |
A |
234 |
222 |
1.06 |
|
|
|
| 10 |
A |
90 |
86 |
0.23 |
|
|
|
| 11 |
B |
3208 |
3055 |
1.91 |
|
|
|
| 12 |
B |
3124 |
2975 |
1.92 |
|
|
|
| 13 |
B |
1515 |
1443 |
10.00 |
|
|
|
| 14 |
B |
1321 |
1258 |
56.37 |
|
|
|
| 15 |
B |
1149 |
1094 |
2.37 |
|
|
|
| 16 |
B |
871 |
830 |
13.91 |
|
|
|
| 17 |
B |
603 |
575 |
10.64 |
|
|
|
| 18 |
B |
365 |
348 |
6.73 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 12740.6 cm
-1
Scaled (by 0.9523) Zero Point Vibrational Energy (zpe) 12132.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD/6-31G*
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.310 |
0.691 |
1.217 |
| C2 |
-0.310 |
-0.691 |
1.217 |
| Br3 |
-0.310 |
1.769 |
-0.302 |
| Br4 |
0.310 |
-1.769 |
-0.302 |
| H5 |
0.015 |
1.237 |
2.117 |
| H6 |
1.398 |
0.648 |
1.145 |
| H7 |
-0.015 |
-1.237 |
2.117 |
| H8 |
-1.398 |
-0.648 |
1.145 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Br3 |
Br4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5146 | 1.9628 | 2.8908 | 1.0939 | 1.0912 | 2.1524 | 2.1718 |
C2 | 1.5146 | | 2.8908 | 1.9628 | 2.1524 | 2.1718 | 1.0939 | 1.0912 | Br3 | 1.9628 | 2.8908 | | 3.5918 | 2.4983 | 2.5035 | 3.8697 | 3.0202 | Br4 | 2.8908 | 1.9628 | 3.5918 | | 3.8697 | 3.0202 | 2.4983 | 2.5035 | H5 | 1.0939 | 2.1524 | 2.4983 | 3.8697 | | 1.7901 | 2.4737 | 2.5486 | H6 | 1.0912 | 2.1718 | 2.5035 | 3.0202 | 1.7901 | | 2.5486 | 3.0823 | H7 | 2.1524 | 1.0939 | 3.8697 | 2.4983 | 2.4737 | 2.5486 | | 1.7901 | H8 | 2.1718 | 1.0912 | 3.0202 | 2.5035 | 2.5486 | 3.0823 | 1.7901 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
Br4 |
111.823 |
|
C1 |
C2 |
H7 |
110.168 |
| C1 |
C2 |
H8 |
111.884 |
|
C2 |
C1 |
Br3 |
111.823 |
| C2 |
C1 |
H5 |
110.168 |
|
C2 |
C1 |
H6 |
111.884 |
| Br3 |
C1 |
H5 |
106.127 |
|
Br3 |
C1 |
H6 |
106.607 |
| Br4 |
C2 |
H7 |
106.127 |
|
Br4 |
C2 |
H8 |
106.607 |
| H5 |
C1 |
H6 |
110.019 |
|
H7 |
C2 |
H8 |
110.019 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
