Jump to
S1C2
Energy calculated at B2PLYP/6-31G*
| | hartrees |
|---|
| Energy at 0K | -5220.967224 |
| Energy at 298.15K | |
| HF Energy | -5220.805701 |
| Nuclear repulsion energy | 415.293730 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3163 |
3003 |
0.00 |
|
|
|
| 2 |
Ag |
1536 |
1458 |
0.00 |
|
|
|
| 3 |
Ag |
1319 |
1252 |
0.00 |
|
|
|
| 4 |
Ag |
1088 |
1032 |
0.00 |
|
|
|
| 5 |
Ag |
674 |
640 |
0.00 |
|
|
|
| 6 |
Ag |
191 |
181 |
0.00 |
|
|
|
| 7 |
Au |
3250 |
3085 |
0.82 |
|
|
|
| 8 |
Au |
1122 |
1065 |
3.14 |
|
|
|
| 9 |
Au |
770 |
731 |
4.08 |
|
|
|
| 10 |
Au |
105 |
99 |
4.66 |
|
|
|
| 11 |
Bg |
3230 |
3065 |
0.00 |
|
|
|
| 12 |
Bg |
1319 |
1252 |
0.00 |
|
|
|
| 13 |
Bg |
958 |
909 |
0.00 |
|
|
|
| 14 |
Bu |
3170 |
3009 |
6.17 |
|
|
|
| 15 |
Bu |
1530 |
1452 |
6.32 |
|
|
|
| 16 |
Bu |
1237 |
1174 |
63.93 |
|
|
|
| 17 |
Bu |
597 |
567 |
72.28 |
|
|
|
| 18 |
Bu |
183 |
174 |
7.19 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 12719.6 cm
-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 12073.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP/6-31G*
Point Group is C2h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.496 |
0.569 |
0.000 |
| C2 |
-0.496 |
-0.569 |
0.000 |
| Br3 |
-0.496 |
2.275 |
0.000 |
| Br4 |
0.496 |
-2.275 |
0.000 |
| H5 |
1.117 |
0.573 |
0.892 |
| H6 |
1.117 |
0.573 |
-0.892 |
| H7 |
-1.117 |
-0.573 |
0.892 |
| H8 |
-1.117 |
-0.573 |
-0.892 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Br3 |
Br4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5097 | 1.9738 | 2.8439 | 1.0872 | 1.0872 | 2.1689 | 2.1689 |
C2 | 1.5097 | | 2.8439 | 1.9738 | 2.1689 | 2.1689 | 1.0872 | 1.0872 | Br3 | 1.9738 | 2.8439 | | 4.6570 | 2.5092 | 2.5092 | 3.0486 | 3.0486 | Br4 | 2.8439 | 1.9738 | 4.6570 | | 3.0486 | 3.0486 | 2.5092 | 2.5092 | H5 | 1.0872 | 2.1689 | 2.5092 | 3.0486 | | 1.7846 | 2.5116 | 3.0811 | H6 | 1.0872 | 2.1689 | 2.5092 | 3.0486 | 1.7846 | | 3.0811 | 2.5116 | H7 | 2.1689 | 1.0872 | 3.0486 | 2.5092 | 2.5116 | 3.0811 | | 1.7846 | H8 | 2.1689 | 1.0872 | 3.0486 | 2.5092 | 3.0811 | 2.5116 | 1.7846 | |
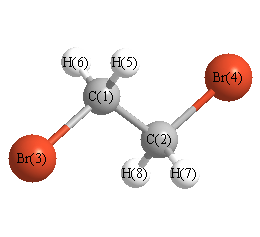 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
Br4 |
108.719 |
|
C1 |
C2 |
H7 |
112.244 |
| C1 |
C2 |
H8 |
112.244 |
|
C2 |
C1 |
Br3 |
108.719 |
| C2 |
C1 |
H5 |
112.244 |
|
C2 |
C1 |
H6 |
112.244 |
| Br3 |
C1 |
H5 |
106.492 |
|
Br3 |
C1 |
H6 |
106.492 |
| Br4 |
C2 |
H7 |
106.492 |
|
Br4 |
C2 |
H8 |
106.492 |
| H5 |
C1 |
H6 |
110.306 |
|
H7 |
C2 |
H8 |
110.306 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.300 |
|
|
|
| 2 |
C |
-0.300 |
|
|
|
| 3 |
Br |
-0.146 |
|
|
|
| 4 |
Br |
-0.146 |
|
|
|
| 5 |
H |
0.223 |
|
|
|
| 6 |
H |
0.223 |
|
|
|
| 7 |
H |
0.223 |
|
|
|
| 8 |
H |
0.223 |
|
|
|
Electric dipole moments
Electric Quadrupole moment
Quadrupole components in D Å
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.728 |
-1.879 |
0.000 |
| y |
-1.879 |
11.997 |
0.000 |
| z |
0.000 |
0.000 |
5.830 |
<r2> (average value of r
2) Å
2
| <r2> |
427.833 |
| (<r2>)1/2 |
20.684 |
Jump to
S1C1
Energy calculated at B2PLYP/6-31G*
| | hartrees |
|---|
| Energy at 0K | -5220.964564 |
| Energy at 298.15K | |
| HF Energy | -5220.802622 |
| Nuclear repulsion energy | 453.607582 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3210 |
3047 |
0.04 |
93.97 |
0.75 |
0.86 |
| 2 |
A |
3143 |
2983 |
15.05 |
189.44 |
0.02 |
0.04 |
| 3 |
A |
1518 |
1441 |
0.05 |
6.03 |
0.70 |
0.83 |
| 4 |
A |
1342 |
1274 |
27.01 |
3.80 |
0.60 |
0.75 |
| 5 |
A |
1216 |
1155 |
2.63 |
16.11 |
0.75 |
0.86 |
| 6 |
A |
1050 |
997 |
0.49 |
3.73 |
0.73 |
0.84 |
| 7 |
A |
930 |
883 |
6.89 |
6.86 |
0.47 |
0.64 |
| 8 |
A |
562 |
533 |
10.32 |
18.66 |
0.08 |
0.16 |
| 9 |
A |
230 |
218 |
1.09 |
1.03 |
0.29 |
0.45 |
| 10 |
A |
88 |
83 |
0.20 |
0.87 |
0.71 |
0.83 |
| 11 |
B |
3222 |
3058 |
1.59 |
27.96 |
0.75 |
0.86 |
| 12 |
B |
3135 |
2976 |
1.74 |
54.14 |
0.75 |
0.86 |
| 13 |
B |
1509 |
1432 |
9.26 |
17.51 |
0.75 |
0.86 |
| 14 |
B |
1310 |
1244 |
65.78 |
0.78 |
0.75 |
0.86 |
| 15 |
B |
1143 |
1085 |
2.29 |
5.59 |
0.75 |
0.86 |
| 16 |
B |
862 |
818 |
16.25 |
0.84 |
0.75 |
0.86 |
| 17 |
B |
596 |
566 |
12.99 |
10.12 |
0.75 |
0.86 |
| 18 |
B |
361 |
342 |
7.43 |
2.80 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 12712.7 cm
-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 12066.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP/6-31G*
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.308 |
0.690 |
1.217 |
| C2 |
-0.308 |
-0.690 |
1.217 |
| Br3 |
-0.308 |
1.770 |
-0.302 |
| Br4 |
0.308 |
-1.770 |
-0.302 |
| H5 |
0.008 |
1.235 |
2.113 |
| H6 |
1.393 |
0.653 |
1.148 |
| H7 |
-0.008 |
-1.235 |
2.113 |
| H8 |
-1.393 |
-0.653 |
1.148 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Br3 |
Br4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5109 | 1.9626 | 2.8911 | 1.0907 | 1.0878 | 2.1463 | 2.1675 |
C2 | 1.5109 | | 2.8911 | 1.9626 | 2.1463 | 2.1675 | 1.0907 | 1.0878 | Br3 | 1.9626 | 2.8911 | | 3.5929 | 2.4936 | 2.4984 | 3.8664 | 3.0248 | Br4 | 2.8911 | 1.9626 | 3.5929 | | 3.8664 | 3.0248 | 2.4936 | 2.4984 | H5 | 1.0907 | 2.1463 | 2.4936 | 3.8664 | | 1.7853 | 2.4693 | 2.5402 | H6 | 1.0878 | 2.1675 | 2.4984 | 3.0248 | 1.7853 | | 2.5402 | 3.0758 | H7 | 2.1463 | 1.0907 | 3.8664 | 2.4936 | 2.4693 | 2.5402 | | 1.7853 | H8 | 2.1675 | 1.0878 | 3.0248 | 2.4984 | 2.5402 | 3.0758 | 1.7853 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
Br4 |
112.023 |
|
C1 |
C2 |
H7 |
110.127 |
| C1 |
C2 |
H8 |
112.006 |
|
C2 |
C1 |
Br3 |
112.023 |
| C2 |
C1 |
H5 |
110.127 |
|
C2 |
C1 |
H6 |
112.006 |
| Br3 |
C1 |
H5 |
105.956 |
|
Br3 |
C1 |
H6 |
106.424 |
| Br4 |
C2 |
H7 |
105.956 |
|
Br4 |
C2 |
H8 |
106.424 |
| H5 |
C1 |
H6 |
110.076 |
|
H7 |
C2 |
H8 |
110.076 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.302 |
|
|
|
| 2 |
C |
-0.302 |
|
|
|
| 3 |
Br |
-0.120 |
|
|
|
| 4 |
Br |
-0.120 |
|
|
|
| 5 |
H |
0.205 |
|
|
|
| 6 |
H |
0.217 |
|
|
|
| 7 |
H |
0.205 |
|
|
|
| 8 |
H |
0.217 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
2.722 |
2.722 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-48.957 |
0.908 |
0.000 |
| y |
0.908 |
-53.504 |
0.000 |
| z |
0.000 |
0.000 |
-44.415 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.003 |
0.908 |
0.000 |
| y |
0.908 |
-6.818 |
0.000 |
| z |
0.000 |
0.000 |
6.816 |
|
| Polar |
|---|
| 3z2-r2 | 13.631 |
|---|
| x2-y2 | 4.547 |
|---|
| xy | 0.908 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.125 |
-0.755 |
0.000 |
| y |
-0.755 |
8.953 |
0.000 |
| z |
0.000 |
0.000 |
8.108 |
<r2> (average value of r
2) Å
2
| <r2> |
306.835 |
| (<r2>)1/2 |
17.517 |
