Jump to
S1C2
Energy calculated at PBEPBEultrafine/6-31G*
| | hartrees |
|---|
| Energy at 0K | -5221.112525 |
| Energy at 298.15K | |
| HF Energy | -5221.112525 |
| Nuclear repulsion energy | 414.457792 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBEPBEultrafine/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3059 |
3008 |
0.00 |
144.80 |
0.06 |
0.11 |
| 2 |
Ag |
1466 |
1441 |
0.00 |
15.39 |
0.72 |
0.84 |
| 3 |
Ag |
1247 |
1226 |
0.00 |
33.71 |
0.46 |
0.63 |
| 4 |
Ag |
1058 |
1041 |
0.00 |
13.70 |
0.66 |
0.80 |
| 5 |
Ag |
642 |
631 |
0.00 |
104.86 |
0.28 |
0.44 |
| 6 |
Ag |
185 |
182 |
0.00 |
4.25 |
0.31 |
0.47 |
| 7 |
Au |
3145 |
3093 |
1.03 |
0.00 |
0.00 |
0.00 |
| 8 |
Au |
1066 |
1049 |
3.29 |
0.00 |
0.00 |
0.00 |
| 9 |
Au |
741 |
729 |
5.15 |
0.00 |
0.00 |
0.00 |
| 10 |
Au |
102 |
100 |
4.06 |
0.00 |
0.00 |
0.00 |
| 11 |
Bg |
3122 |
3071 |
0.00 |
91.85 |
0.75 |
0.86 |
| 12 |
Bg |
1259 |
1238 |
0.00 |
6.79 |
0.75 |
0.86 |
| 13 |
Bg |
910 |
895 |
0.00 |
7.80 |
0.75 |
0.86 |
| 14 |
Bu |
3067 |
3016 |
5.88 |
0.00 |
0.00 |
0.00 |
| 15 |
Bu |
1459 |
1435 |
6.16 |
0.00 |
0.00 |
0.00 |
| 16 |
Bu |
1173 |
1154 |
51.87 |
0.00 |
0.00 |
0.00 |
| 17 |
Bu |
579 |
570 |
77.50 |
0.00 |
0.00 |
0.00 |
| 18 |
Bu |
173 |
170 |
7.78 |
0.00 |
0.00 |
0.00 |
Unscaled Zero Point Vibrational Energy (zpe) 12225.5 cm
-1
Scaled (by 0.9835) Zero Point Vibrational Energy (zpe) 12023.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBEPBEultrafine/6-31G*
Point Group is C2h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.499 |
0.568 |
0.000 |
| C2 |
-0.499 |
-0.568 |
0.000 |
| Br3 |
-0.499 |
2.277 |
0.000 |
| Br4 |
0.499 |
-2.277 |
0.000 |
| H5 |
1.127 |
0.573 |
0.901 |
| H6 |
1.127 |
0.573 |
-0.901 |
| H7 |
-1.127 |
-0.573 |
0.901 |
| H8 |
-1.127 |
-0.573 |
-0.901 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Br3 |
Br4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5124 | 1.9787 | 2.8454 | 1.0979 | 1.0979 | 2.1807 | 2.1807 |
C2 | 1.5124 | | 2.8454 | 1.9787 | 2.1807 | 2.1807 | 1.0979 | 1.0979 | Br3 | 1.9787 | 2.8454 | | 4.6621 | 2.5213 | 2.5213 | 3.0541 | 3.0541 | Br4 | 2.8454 | 1.9787 | 4.6621 | | 3.0541 | 3.0541 | 2.5213 | 2.5213 | H5 | 1.0979 | 2.1807 | 2.5213 | 3.0541 | | 1.8014 | 2.5277 | 3.1039 | H6 | 1.0979 | 2.1807 | 2.5213 | 3.0541 | 1.8014 | | 3.1039 | 2.5277 | H7 | 2.1807 | 1.0979 | 3.0541 | 2.5213 | 2.5277 | 3.1039 | | 1.8014 | H8 | 2.1807 | 1.0979 | 3.0541 | 2.5213 | 3.1039 | 2.5277 | 1.8014 | |
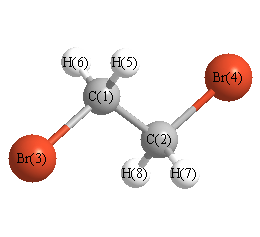 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
Br4 |
108.446 |
|
C1 |
C2 |
H7 |
112.353 |
| C1 |
C2 |
H8 |
112.353 |
|
C2 |
C1 |
Br3 |
108.446 |
| C2 |
C1 |
H5 |
112.353 |
|
C2 |
C1 |
H6 |
112.353 |
| Br3 |
C1 |
H5 |
106.539 |
|
Br3 |
C1 |
H6 |
106.539 |
| Br4 |
C2 |
H7 |
106.539 |
|
Br4 |
C2 |
H8 |
106.539 |
| H5 |
C1 |
H6 |
110.253 |
|
H7 |
C2 |
H8 |
110.253 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.330 |
|
|
|
| 2 |
C |
-0.330 |
|
|
|
| 3 |
Br |
-0.113 |
|
|
|
| 4 |
Br |
-0.113 |
|
|
|
| 5 |
H |
0.221 |
|
|
|
| 6 |
H |
0.221 |
|
|
|
| 7 |
H |
0.221 |
|
|
|
| 8 |
H |
0.221 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-48.112 |
1.141 |
0.000 |
| y |
1.141 |
-54.671 |
0.000 |
| z |
0.000 |
0.000 |
-48.884 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.666 |
1.141 |
0.000 |
| y |
1.141 |
-6.173 |
0.000 |
| z |
0.000 |
0.000 |
2.507 |
|
| Polar |
|---|
| 3z2-r2 | 5.015 |
|---|
| x2-y2 | 6.559 |
|---|
| xy | 1.141 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.908 |
-1.877 |
0.000 |
| y |
-1.877 |
12.821 |
0.000 |
| z |
0.000 |
0.000 |
5.997 |
<r2> (average value of r
2) Å
2
| <r2> |
428.438 |
| (<r2>)1/2 |
20.699 |
Jump to
S1C1
Energy calculated at PBEPBEultrafine/6-31G*
| | hartrees |
|---|
| Energy at 0K | -5221.109853 |
| Energy at 298.15K | |
| HF Energy | -5221.109853 |
| Nuclear repulsion energy | 453.115169 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBEPBEultrafine/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3102 |
3051 |
0.04 |
100.63 |
0.75 |
0.86 |
| 2 |
A |
3037 |
2987 |
14.29 |
225.78 |
0.02 |
0.03 |
| 3 |
A |
1444 |
1420 |
0.42 |
6.26 |
0.63 |
0.77 |
| 4 |
A |
1279 |
1258 |
23.51 |
5.50 |
0.64 |
0.78 |
| 5 |
A |
1161 |
1141 |
2.74 |
19.69 |
0.75 |
0.86 |
| 6 |
A |
1012 |
995 |
0.90 |
4.16 |
0.59 |
0.74 |
| 7 |
A |
888 |
873 |
7.33 |
7.76 |
0.48 |
0.65 |
| 8 |
A |
546 |
537 |
8.88 |
16.48 |
0.08 |
0.15 |
| 9 |
A |
222 |
218 |
1.10 |
1.05 |
0.28 |
0.44 |
| 10 |
A |
84 |
83 |
0.14 |
0.80 |
0.72 |
0.83 |
| 11 |
B |
3116 |
3065 |
1.91 |
31.79 |
0.75 |
0.86 |
| 12 |
B |
3030 |
2980 |
1.94 |
65.64 |
0.75 |
0.86 |
| 13 |
B |
1434 |
1411 |
10.63 |
18.57 |
0.75 |
0.86 |
| 14 |
B |
1242 |
1221 |
58.99 |
0.57 |
0.75 |
0.86 |
| 15 |
B |
1093 |
1075 |
2.15 |
6.16 |
0.75 |
0.86 |
| 16 |
B |
819 |
806 |
19.66 |
0.73 |
0.75 |
0.86 |
| 17 |
B |
576 |
567 |
12.66 |
8.01 |
0.75 |
0.86 |
| 18 |
B |
348 |
342 |
7.45 |
2.61 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 12216.4 cm
-1
Scaled (by 0.9835) Zero Point Vibrational Energy (zpe) 12014.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBEPBEultrafine/6-31G*
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.304 |
0.693 |
1.226 |
| C2 |
-0.304 |
-0.693 |
1.226 |
| Br3 |
-0.304 |
1.768 |
-0.304 |
| Br4 |
0.304 |
-1.768 |
-0.304 |
| H5 |
-0.009 |
1.244 |
2.127 |
| H6 |
1.400 |
0.663 |
1.167 |
| H7 |
0.009 |
-1.244 |
2.127 |
| H8 |
-1.400 |
-0.663 |
1.167 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Br3 |
Br4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5129 | 1.9662 | 2.8975 | 1.1015 | 1.0984 | 2.1562 | 2.1782 |
C2 | 1.5129 | | 2.8975 | 1.9662 | 2.1562 | 2.1782 | 1.1015 | 1.0984 | Br3 | 1.9662 | 2.8975 | | 3.5870 | 2.5047 | 2.5073 | 3.8830 | 3.0452 | Br4 | 2.8975 | 1.9662 | 3.5870 | | 3.8830 | 3.0452 | 2.5047 | 2.5073 | H5 | 1.1015 | 2.1562 | 2.5047 | 3.8830 | | 1.8015 | 2.4875 | 2.5483 | H6 | 1.0984 | 2.1782 | 2.5073 | 3.0452 | 1.8015 | | 2.5483 | 3.0983 | H7 | 2.1562 | 1.1015 | 3.8830 | 2.5047 | 2.4875 | 2.5483 | | 1.8015 | H8 | 2.1782 | 1.0984 | 3.0452 | 2.5073 | 2.5483 | 3.0983 | 1.8015 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
Br4 |
112.125 |
|
C1 |
C2 |
H7 |
110.127 |
| C1 |
C2 |
H8 |
112.076 |
|
C2 |
C1 |
Br3 |
112.125 |
| C2 |
C1 |
H5 |
110.127 |
|
C2 |
C1 |
H6 |
112.076 |
| Br3 |
C1 |
H5 |
106.002 |
|
Br3 |
C1 |
H6 |
106.328 |
| Br4 |
C2 |
H7 |
106.002 |
|
Br4 |
C2 |
H8 |
106.328 |
| H5 |
C1 |
H6 |
109.946 |
|
H7 |
C2 |
H8 |
109.946 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.333 |
|
|
|
| 2 |
C |
-0.333 |
|
|
|
| 3 |
Br |
-0.098 |
|
|
|
| 4 |
Br |
-0.098 |
|
|
|
| 5 |
H |
0.210 |
|
|
|
| 6 |
H |
0.221 |
|
|
|
| 7 |
H |
0.210 |
|
|
|
| 8 |
H |
0.221 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
2.600 |
2.600 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-48.603 |
0.815 |
0.000 |
| y |
0.815 |
-52.764 |
0.000 |
| z |
0.000 |
0.000 |
-44.085 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.179 |
0.815 |
0.000 |
| y |
0.815 |
-6.420 |
0.000 |
| z |
0.000 |
0.000 |
6.599 |
|
| Polar |
|---|
| 3z2-r2 | 13.198 |
|---|
| x2-y2 | 4.161 |
|---|
| xy | 0.815 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.283 |
-0.721 |
0.000 |
| y |
-0.721 |
9.432 |
0.000 |
| z |
0.000 |
0.000 |
8.317 |
<r2> (average value of r
2) Å
2
| <r2> |
306.501 |
| (<r2>)1/2 |
17.507 |
