return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH3CH2CH2Br (n-propyl bromide)

using model chemistry: CID/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at CID/6-31G*
 hartrees
Energy at 0K-2688.047162
Energy at 298.15K-2688.057712
HF Energy-2687.574175
Nuclear repulsion energy236.304269
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CID/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3232 2986 28.65      
2 A' 3218 2972 14.36      
3 A' 3171 2929 9.47      
4 A' 3149 2909 22.69      
5 A' 1598 1476 5.49      
6 A' 1583 1462 1.20      
7 A' 1577 1457 2.61      
8 A' 1507 1392 1.78      
9 A' 1455 1344 8.15      
10 A' 1329 1227 44.94      
11 A' 1178 1088 1.93      
12 A' 1103 1018 1.09      
13 A' 953 880 7.13      
14 A' 693 640 25.26      
15 A' 325 300 2.01      
16 A' 226 209 1.52      
17 A" 3286 3035 9.73      
18 A" 3229 2983 37.88      
19 A" 3208 2963 0.04      
20 A" 1588 1467 8.28      
21 A" 1390 1284 0.01      
22 A" 1314 1214 0.38      
23 A" 1121 1035 1.98      
24 A" 899 831 0.00      
25 A" 777 717 3.56      
26 A" 249 230 0.03      
27 A" 119 110 0.95      

Unscaled Zero Point Vibrational Energy (zpe) 21738.8 cm-1
Scaled (by 0.9237) Zero Point Vibrational Energy (zpe) 20080.1 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at CID/6-31G*
ABC
0.84813 0.05456 0.05277

See section I.F.4 to change rotational constant units
Geometric Data calculated at CID/6-31G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.841 0.000
C2 1.506 0.660 0.000
C3 2.222 2.009 0.000
Br4 -0.928 -0.886 0.000
H5 -0.350 1.362 0.886
H6 -0.350 1.362 -0.886
H7 1.796 0.079 0.875
H8 1.796 0.079 -0.875
H9 3.302 1.870 0.000
H10 1.961 2.595 -0.882
H11 1.961 2.595 0.882

Atom - Atom Distances (Å)
  C1 C2 C3 Br4 H5 H6 H7 H8 H9 H10 H11
C11.51662.51011.96021.08541.08542.13852.13853.45922.77482.7748
C21.51661.52702.88342.17232.17231.09021.09022.16622.17442.1744
C32.51011.52704.27792.79532.79532.16152.16151.08971.09071.0907
Br41.96022.88344.27792.48422.48423.01983.01985.04914.60854.6085
H51.08542.17232.79532.48421.77132.50003.05803.79203.15952.6186
H61.08542.17232.79532.48421.77133.05802.50003.79202.61863.1595
H72.13851.09022.16153.01982.50003.05801.75092.49883.07352.5213
H82.13851.09022.16153.01983.05802.50001.75092.49882.52133.0735
H93.45922.16621.08975.04913.79203.79202.49882.49881.76171.7617
H102.77482.17441.09074.60853.15952.61863.07352.52131.76171.7643
H112.77482.17441.09074.60852.61863.15952.52133.07351.76171.7643

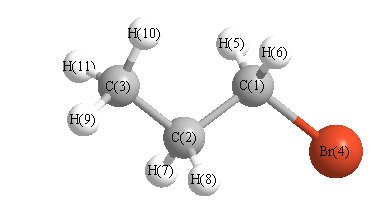
picture of n-propyl bromide state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 111.120 C1 C2 H7 109.145
C1 C2 H8 109.145 C2 C1 Br4 111.423
C2 C1 H5 112.140 C2 C1 H6 112.140
C2 C3 H9 110.644 C2 C3 H10 111.243
C2 C3 H11 111.243 C3 C2 H7 110.244
C3 C2 H8 110.244 Br4 C1 H5 105.690
Br4 C1 H6 105.690 H5 C1 H6 109.365
H7 C2 H8 106.836 H9 C3 H10 107.802
H9 C3 H11 107.802 H10 C3 H11 107.956
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability