Jump to
S1C2
Energy calculated at CID/6-31G*
| | hartrees |
|---|
| Energy at 0K | -157.796815 |
| Energy at 298.15K | -157.807606 |
| HF Energy | -157.298228 |
| Nuclear repulsion energy | 130.893597 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CID/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
|
Au |
126 |
116 |
0.01 |
|
|
|
|
Au |
238 |
219 |
0.00 |
|
|
|
|
Bu |
273 |
252 |
0.03 |
|
|
|
|
Bg |
274 |
253 |
0.00 |
|
|
|
|
Ag |
445 |
411 |
0.00 |
|
|
|
|
Au |
767 |
709 |
3.16 |
|
|
|
|
Bg |
853 |
788 |
0.00 |
|
|
|
|
Ag |
890 |
822 |
0.00 |
|
|
|
|
Au |
1011 |
934 |
0.94 |
|
|
|
|
Bu |
1030 |
952 |
5.78 |
|
|
|
|
Bu |
1083 |
1000 |
0.24 |
|
|
|
|
Ag |
1132 |
1046 |
0.00 |
|
|
|
|
Ag |
1237 |
1142 |
0.00 |
|
|
|
|
Bg |
1285 |
1187 |
0.00 |
|
|
|
|
Au |
1363 |
1259 |
0.19 |
|
|
|
|
Bu |
1398 |
1291 |
2.39 |
|
|
|
|
Bg |
1402 |
1295 |
0.00 |
|
|
|
|
Ag |
1490 |
1376 |
0.00 |
|
|
|
|
Bu |
1509 |
1394 |
5.54 |
|
|
|
|
Ag |
1511 |
1395 |
0.00 |
|
|
|
|
Ag |
1583 |
1462 |
0.00 |
|
|
|
|
Bu |
1583 |
1463 |
2.26 |
|
|
|
|
Bg |
1590 |
1469 |
0.00 |
|
|
|
|
Au |
1591 |
1469 |
13.16 |
|
|
|
|
Ag |
1596 |
1474 |
0.00 |
|
|
|
|
Bu |
1604 |
1482 |
5.96 |
|
|
|
|
Ag |
3133 |
2894 |
0.00 |
|
|
|
|
Bu |
3139 |
2900 |
45.07 |
|
|
|
|
Bu |
3145 |
2905 |
69.29 |
|
|
|
|
Ag |
3147 |
2907 |
0.00 |
|
|
|
|
Bg |
3163 |
2921 |
0.00 |
|
|
|
|
Au |
3183 |
2940 |
24.58 |
|
|
|
|
Bg |
3219 |
2974 |
0.00 |
|
|
|
|
Au |
3223 |
2977 |
111.20 |
|
|
|
|
Ag |
3223 |
2977 |
0.00 |
|
|
|
|
Bu |
3224 |
2978 |
77.40 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 30330.3 cm
-1
Scaled (by 0.9237) Zero Point Vibrational Energy (zpe) 28016.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CID/6-31G*
Point Group is C2h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.421 |
0.636 |
0.000 |
| C2 |
0.421 |
-0.636 |
0.000 |
| C3 |
0.421 |
1.907 |
0.000 |
| C4 |
-0.421 |
-1.907 |
0.000 |
| H5 |
-1.077 |
0.632 |
0.874 |
| H6 |
-1.077 |
0.632 |
-0.874 |
| H7 |
1.077 |
-0.632 |
0.874 |
| H8 |
1.077 |
-0.632 |
-0.874 |
| H9 |
-0.205 |
2.800 |
0.000 |
| H10 |
1.063 |
1.950 |
0.881 |
| H11 |
1.063 |
1.950 |
-0.881 |
| H12 |
0.205 |
-2.800 |
0.000 |
| H13 |
-1.063 |
-1.950 |
0.881 |
| H14 |
-1.063 |
-1.950 |
-0.881 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
| C1 | | 1.5256 | 1.5247 | 2.5434 | 1.0932 | 1.0932 | 2.1488 | 2.1488 | 2.1745 | 2.1692 | 2.1692 | 3.4926 | 2.8067 | 2.8067 |
C2 | 1.5256 | | 2.5434 | 1.5247 | 2.1488 | 2.1488 | 1.0932 | 1.0932 | 3.4926 | 2.8067 | 2.8067 | 2.1745 | 2.1692 | 2.1692 | C3 | 1.5247 | 2.5434 | | 3.9064 | 2.1530 | 2.1530 | 2.7647 | 2.7647 | 1.0903 | 1.0909 | 1.0909 | 4.7122 | 4.2260 | 4.2260 | C4 | 2.5434 | 1.5247 | 3.9064 | | 2.7647 | 2.7647 | 2.1530 | 2.1530 | 4.7122 | 4.2260 | 4.2260 | 1.0903 | 1.0909 | 1.0909 | H5 | 1.0932 | 2.1488 | 2.1530 | 2.7647 | | 1.7486 | 2.4980 | 3.0492 | 2.4950 | 2.5139 | 3.0659 | 3.7666 | 2.5823 | 3.1222 | H6 | 1.0932 | 2.1488 | 2.1530 | 2.7647 | 1.7486 | | 3.0492 | 2.4980 | 2.4950 | 3.0659 | 2.5139 | 3.7666 | 3.1222 | 2.5823 | H7 | 2.1488 | 1.0932 | 2.7647 | 2.1530 | 2.4980 | 3.0492 | | 1.7486 | 3.7666 | 2.5823 | 3.1222 | 2.4950 | 2.5139 | 3.0659 | H8 | 2.1488 | 1.0932 | 2.7647 | 2.1530 | 3.0492 | 2.4980 | 1.7486 | | 3.7666 | 3.1222 | 2.5823 | 2.4950 | 3.0659 | 2.5139 | H9 | 2.1745 | 3.4926 | 1.0903 | 4.7122 | 2.4950 | 2.4950 | 3.7666 | 3.7666 | | 1.7625 | 1.7625 | 5.6148 | 4.9067 | 4.9067 | H10 | 2.1692 | 2.8067 | 1.0909 | 4.2260 | 2.5139 | 3.0659 | 2.5823 | 3.1222 | 1.7625 | | 1.7616 | 4.9067 | 4.4424 | 4.7790 | H11 | 2.1692 | 2.8067 | 1.0909 | 4.2260 | 3.0659 | 2.5139 | 3.1222 | 2.5823 | 1.7625 | 1.7616 | | 4.9067 | 4.7790 | 4.4424 | H12 | 3.4926 | 2.1745 | 4.7122 | 1.0903 | 3.7666 | 3.7666 | 2.4950 | 2.4950 | 5.6148 | 4.9067 | 4.9067 | | 1.7625 | 1.7625 | H13 | 2.8067 | 2.1692 | 4.2260 | 1.0909 | 2.5823 | 3.1222 | 2.5139 | 3.0659 | 4.9067 | 4.4424 | 4.7790 | 1.7625 | | 1.7616 | H14 | 2.8067 | 2.1692 | 4.2260 | 1.0909 | 3.1222 | 2.5823 | 3.0659 | 2.5139 | 4.9067 | 4.7790 | 4.4424 | 1.7625 | 1.7616 | |
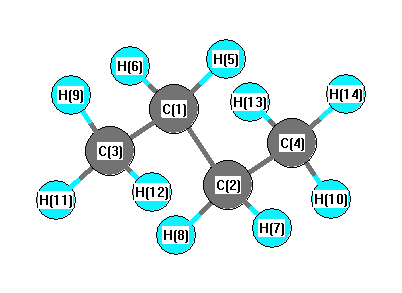 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C4 |
112.987 |
|
C1 |
C2 |
H7 |
109.158 |
| C1 |
C2 |
H8 |
109.158 |
|
C1 |
C3 |
H9 |
111.439 |
| C1 |
C3 |
H11 |
110.971 |
|
C1 |
C3 |
H12 |
30.892 |
| C2 |
C1 |
C3 |
112.987 |
|
C2 |
C1 |
H5 |
109.158 |
| C2 |
C1 |
H6 |
109.158 |
|
C2 |
C4 |
H10 |
17.263 |
| C2 |
C4 |
H13 |
110.971 |
|
C2 |
C4 |
H14 |
110.971 |
| C3 |
C1 |
H5 |
109.551 |
|
C3 |
C1 |
H6 |
109.551 |
| C4 |
C2 |
H7 |
109.551 |
|
C4 |
C2 |
H8 |
109.551 |
| H5 |
C1 |
H6 |
106.208 |
|
H7 |
C2 |
H8 |
106.208 |
| H9 |
C3 |
H11 |
107.809 |
|
H9 |
C3 |
H12 |
142.330 |
| H10 |
C4 |
H13 |
94.267 |
|
H10 |
C4 |
H14 |
114.264 |
| H11 |
C3 |
H12 |
93.800 |
|
H13 |
C4 |
H14 |
107.679 |
Electronic energy levels
Electronic state
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at CID/6-31G*
| | hartrees |
|---|
| Energy at 0K | -157.795548 |
| Energy at 298.15K | |
| Nuclear repulsion energy | 132.707953 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CID/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3236 |
2989 |
37.32 |
|
|
|
| 2 |
A |
3220 |
2975 |
0.40 |
|
|
|
| 3 |
A |
3175 |
2933 |
26.58 |
|
|
|
| 4 |
A |
3151 |
2911 |
14.31 |
|
|
|
| 5 |
A |
3139 |
2899 |
30.22 |
|
|
|
| 6 |
A |
1603 |
1481 |
0.50 |
|
|
|
| 7 |
A |
1599 |
1477 |
4.77 |
|
|
|
| 8 |
A |
1581 |
1460 |
0.23 |
|
|
|
| 9 |
A |
1510 |
1394 |
3.80 |
|
|
|
| 10 |
A |
1472 |
1360 |
0.12 |
|
|
|
| 11 |
A |
1386 |
1280 |
0.39 |
|
|
|
| 12 |
A |
1270 |
1173 |
0.08 |
|
|
|
| 13 |
A |
1154 |
1065 |
0.04 |
|
|
|
| 14 |
A |
1051 |
970 |
0.17 |
|
|
|
| 15 |
A |
881 |
813 |
0.02 |
|
|
|
| 16 |
A |
837 |
773 |
1.52 |
|
|
|
| 17 |
A |
336 |
311 |
0.02 |
|
|
|
| 18 |
A |
281 |
259 |
0.02 |
|
|
|
| 19 |
A |
116 |
107 |
0.00 |
|
|
|
| 20 |
B |
3227 |
2981 |
55.06 |
|
|
|
| 21 |
B |
3222 |
2976 |
85.11 |
|
|
|
| 22 |
B |
3180 |
2938 |
30.72 |
|
|
|
| 23 |
B |
3147 |
2907 |
22.16 |
|
|
|
| 24 |
B |
3136 |
2896 |
27.16 |
|
|
|
| 25 |
B |
1598 |
1476 |
10.88 |
|
|
|
| 26 |
B |
1590 |
1469 |
6.47 |
|
|
|
| 27 |
B |
1579 |
1459 |
0.67 |
|
|
|
| 28 |
B |
1513 |
1397 |
6.27 |
|
|
|
| 29 |
B |
1453 |
1342 |
2.14 |
|
|
|
| 30 |
B |
1361 |
1257 |
0.31 |
|
|
|
| 31 |
B |
1222 |
1129 |
2.49 |
|
|
|
| 32 |
B |
1025 |
947 |
0.10 |
|
|
|
| 33 |
B |
1018 |
940 |
3.29 |
|
|
|
| 34 |
B |
786 |
726 |
3.59 |
|
|
|
| 35 |
B |
456 |
421 |
0.33 |
|
|
|
| 36 |
B |
226 |
209 |
0.03 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 30366.6 cm
-1
Scaled (by 0.9237) Zero Point Vibrational Energy (zpe) 28049.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CID/6-31G*
Point Group is C2h
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
