Jump to
S1C2
Energy calculated at QCISD/6-31G*
| | hartrees |
|---|
| Energy at 0K | -157.883108 |
| Energy at 298.15K | -157.893807 |
| HF Energy | -157.297439 |
| Nuclear repulsion energy | 130.320208 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
|
Au |
123 |
118 |
0.00 |
|
|
|
|
Au |
232 |
221 |
0.00 |
|
|
|
|
Bu |
267 |
254 |
0.03 |
|
|
|
|
Bg |
269 |
256 |
0.00 |
|
|
|
|
Ag |
436 |
415 |
0.00 |
|
|
|
|
Au |
749 |
713 |
3.00 |
|
|
|
|
Bg |
833 |
793 |
0.00 |
|
|
|
|
Ag |
869 |
827 |
0.00 |
|
|
|
|
Au |
986 |
939 |
0.92 |
|
|
|
|
Bu |
1004 |
956 |
5.62 |
|
|
|
|
Bu |
1057 |
1007 |
0.16 |
|
|
|
|
Ag |
1108 |
1055 |
0.00 |
|
|
|
|
Ag |
1204 |
1147 |
0.00 |
|
|
|
|
Bg |
1253 |
1194 |
0.00 |
|
|
|
|
Au |
1327 |
1263 |
0.20 |
|
|
|
|
Bu |
1359 |
1294 |
2.51 |
|
|
|
|
Bg |
1364 |
1299 |
0.00 |
|
|
|
|
Ag |
1448 |
1379 |
0.00 |
|
|
|
|
Bu |
1467 |
1397 |
5.05 |
|
|
|
|
Ag |
1467 |
1397 |
0.00 |
|
|
|
|
Ag |
1542 |
1469 |
0.00 |
|
|
|
|
Bu |
1543 |
1469 |
2.27 |
|
|
|
|
Bg |
1549 |
1475 |
0.00 |
|
|
|
|
Au |
1550 |
1476 |
12.40 |
|
|
|
|
Ag |
1555 |
1481 |
0.00 |
|
|
|
|
Bu |
1564 |
1489 |
5.30 |
|
|
|
|
Ag |
3043 |
2898 |
0.00 |
|
|
|
|
Bu |
3050 |
2904 |
49.69 |
|
|
|
|
Bu |
3058 |
2912 |
63.30 |
|
|
|
|
Ag |
3059 |
2913 |
0.00 |
|
|
|
|
Bg |
3072 |
2925 |
0.00 |
|
|
|
|
Au |
3092 |
2945 |
27.23 |
|
|
|
|
Bg |
3130 |
2981 |
0.00 |
|
|
|
|
Au |
3133 |
2984 |
107.56 |
|
|
|
|
Ag |
3135 |
2985 |
0.00 |
|
|
|
|
Bu |
3135 |
2986 |
76.85 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 29514.7 cm
-1
Scaled (by 0.9523) Zero Point Vibrational Energy (zpe) 28106.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD/6-31G*
Point Group is C2h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.423 |
0.638 |
0.000 |
| C2 |
0.423 |
-0.638 |
0.000 |
| C3 |
0.423 |
1.913 |
0.000 |
| C4 |
-0.423 |
-1.913 |
0.000 |
| H5 |
-1.083 |
0.633 |
0.880 |
| H6 |
-1.083 |
0.633 |
-0.880 |
| H7 |
1.083 |
-0.633 |
0.880 |
| H8 |
1.083 |
-0.633 |
-0.880 |
| H9 |
-0.207 |
2.812 |
0.000 |
| H10 |
1.069 |
1.955 |
0.887 |
| H11 |
1.069 |
1.955 |
-0.887 |
| H12 |
0.207 |
-2.812 |
0.000 |
| H13 |
-1.069 |
-1.955 |
0.887 |
| H14 |
-1.069 |
-1.955 |
-0.887 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
| C1 | | 1.5307 | 1.5300 | 2.5516 | 1.1006 | 1.1006 | 2.1584 | 2.1584 | 2.1848 | 2.1785 | 2.1785 | 3.5074 | 2.8160 | 2.8160 |
C2 | 1.5307 | | 2.5516 | 1.5300 | 2.1584 | 2.1584 | 1.1006 | 1.1006 | 3.5074 | 2.8160 | 2.8160 | 2.1848 | 2.1785 | 2.1785 | C3 | 1.5300 | 2.5516 | | 3.9192 | 2.1637 | 2.1637 | 2.7743 | 2.7743 | 1.0973 | 1.0981 | 1.0981 | 4.7307 | 4.2401 | 4.2401 | C4 | 2.5516 | 1.5300 | 3.9192 | | 2.7743 | 2.7743 | 2.1637 | 2.1637 | 4.7307 | 4.2401 | 4.2401 | 1.0973 | 1.0981 | 1.0981 | H5 | 1.1006 | 2.1584 | 2.1637 | 2.7743 | | 1.7604 | 2.5096 | 3.0655 | 2.5082 | 2.5260 | 3.0826 | 3.7829 | 2.5885 | 3.1340 | H6 | 1.1006 | 2.1584 | 2.1637 | 2.7743 | 1.7604 | | 3.0655 | 2.5096 | 2.5082 | 3.0826 | 2.5260 | 3.7829 | 3.1340 | 2.5885 | H7 | 2.1584 | 1.1006 | 2.7743 | 2.1637 | 2.5096 | 3.0655 | | 1.7604 | 3.7829 | 2.5885 | 3.1340 | 2.5082 | 2.5260 | 3.0826 | H8 | 2.1584 | 1.1006 | 2.7743 | 2.1637 | 3.0655 | 2.5096 | 1.7604 | | 3.7829 | 3.1340 | 2.5885 | 2.5082 | 3.0826 | 2.5260 | H9 | 2.1848 | 3.5074 | 1.0973 | 4.7307 | 2.5082 | 2.5082 | 3.7829 | 3.7829 | | 1.7744 | 1.7744 | 5.6397 | 4.9253 | 4.9253 | H10 | 2.1785 | 2.8160 | 1.0981 | 4.2401 | 2.5260 | 3.0826 | 2.5885 | 3.1340 | 1.7744 | | 1.7733 | 4.9253 | 4.4569 | 4.7967 | H11 | 2.1785 | 2.8160 | 1.0981 | 4.2401 | 3.0826 | 2.5260 | 3.1340 | 2.5885 | 1.7744 | 1.7733 | | 4.9253 | 4.7967 | 4.4569 | H12 | 3.5074 | 2.1848 | 4.7307 | 1.0973 | 3.7829 | 3.7829 | 2.5082 | 2.5082 | 5.6397 | 4.9253 | 4.9253 | | 1.7744 | 1.7744 | H13 | 2.8160 | 2.1785 | 4.2401 | 1.0981 | 2.5885 | 3.1340 | 2.5260 | 3.0826 | 4.9253 | 4.4569 | 4.7967 | 1.7744 | | 1.7733 | H14 | 2.8160 | 2.1785 | 4.2401 | 1.0981 | 3.1340 | 2.5885 | 3.0826 | 2.5260 | 4.9253 | 4.7967 | 4.4569 | 1.7744 | 1.7733 | |
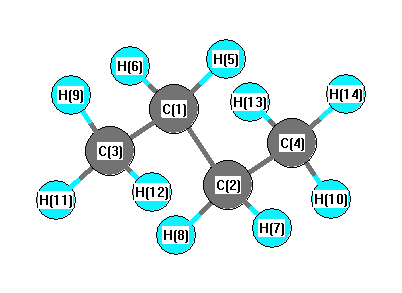 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C4 |
112.956 |
|
C1 |
C2 |
H7 |
109.130 |
| C1 |
C2 |
H8 |
109.130 |
|
C1 |
C3 |
H9 |
111.460 |
| C1 |
C3 |
H11 |
110.912 |
|
C1 |
C3 |
H12 |
30.919 |
| C2 |
C1 |
C3 |
112.956 |
|
C2 |
C1 |
H5 |
109.130 |
| C2 |
C1 |
H6 |
109.130 |
|
C2 |
C4 |
H10 |
17.272 |
| C2 |
C4 |
H13 |
110.912 |
|
C2 |
C4 |
H14 |
110.912 |
| C3 |
C1 |
H5 |
109.597 |
|
C3 |
C1 |
H6 |
109.597 |
| C4 |
C2 |
H7 |
109.597 |
|
C4 |
C2 |
H8 |
109.597 |
| H5 |
C1 |
H6 |
106.204 |
|
H7 |
C2 |
H8 |
106.204 |
| H9 |
C3 |
H11 |
107.851 |
|
H9 |
C3 |
H12 |
142.379 |
| H10 |
C4 |
H13 |
94.185 |
|
H10 |
C4 |
H14 |
114.247 |
| H11 |
C3 |
H12 |
93.719 |
|
H13 |
C4 |
H14 |
107.697 |
Electronic energy levels
Electronic state
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at QCISD/6-31G*
| | hartrees |
|---|
| Energy at 0K | -157.881962 |
| Energy at 298.15K | |
| Nuclear repulsion energy | 132.148322 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3147 |
2997 |
36.74 |
|
|
|
| 2 |
A |
3131 |
2982 |
0.42 |
|
|
|
| 3 |
A |
3083 |
2936 |
26.14 |
|
|
|
| 4 |
A |
3063 |
2917 |
18.37 |
|
|
|
| 5 |
A |
3048 |
2903 |
25.83 |
|
|
|
| 6 |
A |
1562 |
1488 |
0.79 |
|
|
|
| 7 |
A |
1558 |
1483 |
4.09 |
|
|
|
| 8 |
A |
1540 |
1467 |
0.19 |
|
|
|
| 9 |
A |
1467 |
1397 |
3.43 |
|
|
|
| 10 |
A |
1430 |
1361 |
0.24 |
|
|
|
| 11 |
A |
1348 |
1284 |
0.33 |
|
|
|
| 12 |
A |
1237 |
1178 |
0.06 |
|
|
|
| 13 |
A |
1128 |
1074 |
0.03 |
|
|
|
| 14 |
A |
1023 |
974 |
0.16 |
|
|
|
| 15 |
A |
859 |
818 |
0.01 |
|
|
|
| 16 |
A |
818 |
779 |
1.46 |
|
|
|
| 17 |
A |
329 |
313 |
0.02 |
|
|
|
| 18 |
A |
277 |
264 |
0.02 |
|
|
|
| 19 |
A |
113 |
108 |
0.01 |
|
|
|
| 20 |
B |
3138 |
2988 |
53.44 |
|
|
|
| 21 |
B |
3132 |
2983 |
83.64 |
|
|
|
| 22 |
B |
3088 |
2941 |
32.89 |
|
|
|
| 23 |
B |
3060 |
2914 |
21.61 |
|
|
|
| 24 |
B |
3045 |
2900 |
26.96 |
|
|
|
| 25 |
B |
1557 |
1483 |
10.15 |
|
|
|
| 26 |
B |
1549 |
1475 |
6.24 |
|
|
|
| 27 |
B |
1539 |
1465 |
0.62 |
|
|
|
| 28 |
B |
1470 |
1400 |
6.03 |
|
|
|
| 29 |
B |
1413 |
1346 |
2.27 |
|
|
|
| 30 |
B |
1325 |
1262 |
0.31 |
|
|
|
| 31 |
B |
1190 |
1133 |
2.35 |
|
|
|
| 32 |
B |
1001 |
953 |
0.26 |
|
|
|
| 33 |
B |
993 |
945 |
3.02 |
|
|
|
| 34 |
B |
766 |
730 |
3.43 |
|
|
|
| 35 |
B |
447 |
425 |
0.34 |
|
|
|
| 36 |
B |
222 |
211 |
0.03 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 29548.3 cm
-1
Scaled (by 0.9523) Zero Point Vibrational Energy (zpe) 28138.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD/6-31G*
Point Group is C2h
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
