return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH2CHCH2CH3 (1-Butene)

using model chemistry: QCISD/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at QCISD/6-31G*
 hartrees
Energy at 0K-156.672294
Energy at 298.15K-156.680272
HF Energy-156.104681
Nuclear repulsion energy116.785745
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3251 3096 21.42      
2 A 3172 3021 4.80      
3 A 3153 3003 33.83      
4 A 3144 2994 34.68      
5 A 3137 2987 31.72      
6 A 3092 2945 11.63      
7 A 3063 2917 25.94      
8 A 3049 2904 29.40      
9 A 1744 1661 4.94      
10 A 1556 1482 4.83      
11 A 1548 1474 5.32      
12 A 1538 1464 2.09      
13 A 1494 1423 1.76      
14 A 1463 1393 2.01      
15 A 1389 1322 2.53      
16 A 1346 1282 1.13      
17 A 1324 1260 0.06      
18 A 1233 1174 0.20      
19 A 1125 1072 3.48      
20 A 1066 1015 1.19      
21 A 1027 978 14.12      
22 A 1011 963 2.30      
23 A 930 886 36.94      
24 A 883 841 1.08      
25 A 811 773 2.28      
26 A 655 623 7.96      
27 A 438 417 0.63      
28 A 324 308 0.45      
29 A 240 229 0.03      
30 A 105 100 0.06      

Unscaled Zero Point Vibrational Energy (zpe) 24153.8 cm-1
Scaled (by 0.9523) Zero Point Vibrational Energy (zpe) 23001.7 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD/6-31G*
ABC
0.74942 0.13788 0.13472

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD/6-31G*

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 1.723 -0.245 -0.298
C2 0.543 0.521 0.313
C3 -0.718 -0.302 0.342
C4 -1.859 0.021 -0.280
H5 -0.667 -1.235 0.910
H6 1.506 -0.528 -1.336
H7 2.636 0.364 -0.292
H8 1.927 -1.165 0.265
H9 0.802 0.826 1.338
H10 0.362 1.445 -0.255
H11 -1.955 0.940 -0.859
H12 -2.740 -0.616 -0.232

Atom - Atom Distances (Å)
  C1 C2 C3 C4 H5 H6 H7 H8 H9 H10 H11 H12
C11.53422.52453.59232.85471.09691.09721.09782.16192.17063.90454.4792
C21.53421.50622.52382.21442.17812.18402.18271.10081.09912.79013.5166
C32.52451.50621.33881.09362.79493.47752.78412.13932.13912.12422.1250
C43.59232.52381.33882.10153.56944.50794.00563.21652.63851.08981.0882
H52.85472.21441.09362.10153.20363.86062.67402.56683.09803.08502.4470
H61.09692.17812.79493.56943.20361.77791.77383.07842.52343.78894.3882
H71.09722.18403.47754.50793.86061.77791.77462.49642.51784.66105.4647
H81.09782.18272.78414.00562.67401.77381.77462.52683.08784.55694.7263
H92.16191.10082.13933.21652.56683.07842.49642.52681.76443.52684.1340
H102.17061.09912.13912.63853.09802.52342.51783.08781.76442.44713.7244
H113.90452.79012.12421.08983.08503.78894.66104.55693.52682.44711.8516
H124.47923.51662.12501.08822.44704.38825.46474.72634.13403.72441.8516

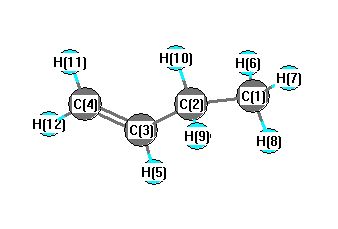
picture of 1-Butene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 112.255 C1 C2 H9 109.153
C1 C2 H10 109.933 C2 C1 H6 110.653
C2 C1 H7 111.109 C2 C1 H8 110.964
C2 C3 C4 124.914 C2 C3 H5 115.897
C3 C2 H9 109.303 C3 C2 H10 109.384
C3 C4 H11 121.669 C3 C4 H12 121.881
C4 C3 H5 119.184 H6 C1 H7 108.252
H6 C1 H8 107.842 H7 C1 H8 107.894
H9 C2 H10 106.652 H11 C4 H12 116.450
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability