Jump to
S1C2
Energy calculated at CID/6-31G*
| | hartrees |
|---|
| Energy at 0K | -155.386564 |
| Energy at 298.15K | -155.392003 |
| HF Energy | -154.919307 |
| Nuclear repulsion energy | 103.988407 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CID/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3350 |
3094 |
0.00 |
|
|
|
| 2 |
Ag |
3270 |
3021 |
0.00 |
|
|
|
| 3 |
Ag |
3253 |
3005 |
0.00 |
|
|
|
| 4 |
Ag |
1833 |
1693 |
0.00 |
|
|
|
| 5 |
Ag |
1553 |
1434 |
0.00 |
|
|
|
| 6 |
Ag |
1383 |
1277 |
0.00 |
|
|
|
| 7 |
Ag |
1289 |
1191 |
0.00 |
|
|
|
| 8 |
Ag |
939 |
867 |
0.00 |
|
|
|
| 9 |
Ag |
535 |
494 |
0.00 |
|
|
|
| 10 |
Au |
1094 |
1011 |
35.35 |
|
|
|
| 11 |
Au |
987 |
912 |
88.78 |
|
|
|
| 12 |
Au |
553 |
511 |
10.82 |
|
|
|
| 13 |
Au |
158 |
146 |
0.14 |
|
|
|
| 14 |
Bg |
1045 |
965 |
0.00 |
|
|
|
| 15 |
Bg |
988 |
912 |
0.00 |
|
|
|
| 16 |
Bg |
808 |
746 |
0.00 |
|
|
|
| 17 |
Bu |
3350 |
3095 |
30.05 |
|
|
|
| 18 |
Bu |
3269 |
3020 |
8.30 |
|
|
|
| 19 |
Bu |
3260 |
3011 |
22.91 |
|
|
|
| 20 |
Bu |
1757 |
1623 |
10.71 |
|
|
|
| 21 |
Bu |
1491 |
1377 |
3.97 |
|
|
|
| 22 |
Bu |
1386 |
1280 |
2.15 |
|
|
|
| 23 |
Bu |
1051 |
970 |
2.41 |
|
|
|
| 24 |
Bu |
307 |
284 |
2.38 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 19453.3 cm
-1
Scaled (by 0.9237) Zero Point Vibrational Energy (zpe) 17969.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CID/6-31G*
Point Group is C2h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.731 |
0.000 |
| C2 |
0.000 |
-0.731 |
0.000 |
| C3 |
1.105 |
1.473 |
0.000 |
| C4 |
-1.105 |
-1.473 |
0.000 |
| H5 |
-0.972 |
1.214 |
0.000 |
| H6 |
0.972 |
-1.214 |
0.000 |
| H7 |
1.065 |
2.553 |
0.000 |
| H8 |
2.090 |
1.025 |
0.000 |
| H9 |
-1.065 |
-2.553 |
0.000 |
| H10 |
-2.090 |
-1.025 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| C1 | | 1.4629 | 1.3306 | 2.4658 | 1.0849 | 2.1747 | 2.1102 | 2.1107 | 3.4529 | 2.7302 |
C2 | 1.4629 | | 2.4658 | 1.3306 | 2.1747 | 1.0849 | 3.4529 | 2.7302 | 2.1102 | 2.1107 | C3 | 1.3306 | 2.4658 | | 3.6826 | 2.0926 | 2.6904 | 1.0806 | 1.0825 | 4.5736 | 4.0556 | C4 | 2.4658 | 1.3306 | 3.6826 | | 2.6904 | 2.0926 | 4.5736 | 4.0556 | 1.0806 | 1.0825 | H5 | 1.0849 | 2.1747 | 2.0926 | 2.6904 | | 3.1101 | 2.4377 | 3.0677 | 3.7682 | 2.5028 | H6 | 2.1747 | 1.0849 | 2.6904 | 2.0926 | 3.1101 | | 3.7682 | 2.5028 | 2.4377 | 3.0677 | H7 | 2.1102 | 3.4529 | 1.0806 | 4.5736 | 2.4377 | 3.7682 | | 1.8398 | 5.5326 | 4.7706 | H8 | 2.1107 | 2.7302 | 1.0825 | 4.0556 | 3.0677 | 2.5028 | 1.8398 | | 4.7706 | 4.6559 | H9 | 3.4529 | 2.1102 | 4.5736 | 1.0806 | 3.7682 | 2.4377 | 5.5326 | 4.7706 | | 1.8398 | H10 | 2.7302 | 2.1107 | 4.0556 | 1.0825 | 2.5028 | 3.0677 | 4.7706 | 4.6559 | 1.8398 | |
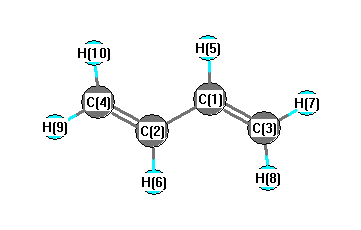 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C4 |
123.870 |
|
C1 |
C2 |
H6 |
116.412 |
| C1 |
C3 |
H7 |
121.783 |
|
C1 |
C3 |
H8 |
121.678 |
| C2 |
C1 |
C3 |
123.870 |
|
C2 |
C1 |
H5 |
116.412 |
| C2 |
C4 |
H9 |
121.783 |
|
C2 |
C4 |
H10 |
121.678 |
| C3 |
C1 |
H5 |
119.718 |
|
C4 |
C2 |
H6 |
119.718 |
| H7 |
C3 |
H8 |
116.539 |
|
H9 |
C4 |
H10 |
116.539 |
Electronic energy levels
Electronic state
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at CID/6-31G*
| | hartrees |
|---|
| Energy at 0K | -155.382269 |
| Energy at 298.15K | -155.387730 |
| HF Energy | -154.914427 |
| Nuclear repulsion energy | 105.148813 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CID/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3353 |
3097 |
6.27 |
|
|
|
| 2 |
A |
3276 |
3026 |
1.70 |
|
|
|
| 3 |
A |
3259 |
3011 |
24.85 |
|
|
|
| 4 |
A |
1799 |
1662 |
1.10 |
|
|
|
| 5 |
A |
1543 |
1426 |
8.23 |
|
|
|
| 6 |
A |
1406 |
1299 |
0.00 |
|
|
|
| 7 |
A |
1124 |
1039 |
0.17 |
|
|
|
| 8 |
A |
1060 |
979 |
2.93 |
|
|
|
| 9 |
A |
996 |
920 |
7.87 |
|
|
|
| 10 |
A |
929 |
858 |
0.45 |
|
|
|
| 11 |
A |
791 |
730 |
5.17 |
|
|
|
| 12 |
A |
288 |
266 |
0.00 |
|
|
|
| 13 |
A |
178 |
165 |
0.04 |
|
|
|
| 14 |
B |
3351 |
3096 |
22.94 |
|
|
|
| 15 |
B |
3269 |
3019 |
9.45 |
|
|
|
| 16 |
B |
3251 |
3003 |
8.74 |
|
|
|
| 17 |
B |
1797 |
1660 |
2.47 |
|
|
|
| 18 |
B |
1512 |
1396 |
1.06 |
|
|
|
| 19 |
B |
1374 |
1269 |
0.74 |
|
|
|
| 20 |
B |
1155 |
1067 |
5.04 |
|
|
|
| 21 |
B |
1079 |
997 |
30.35 |
|
|
|
| 22 |
B |
997 |
921 |
76.38 |
|
|
|
| 23 |
B |
652 |
602 |
6.89 |
|
|
|
| 24 |
B |
483 |
446 |
10.71 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 19460.3 cm
-1
Scaled (by 0.9237) Zero Point Vibrational Energy (zpe) 17975.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CID/6-31G*
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.119 |
0.727 |
0.547 |
| C2 |
-0.119 |
-0.727 |
0.547 |
| C3 |
-0.119 |
1.531 |
-0.485 |
| C4 |
0.119 |
-1.531 |
-0.485 |
| H5 |
0.501 |
1.151 |
1.470 |
| H6 |
-0.501 |
-1.151 |
1.470 |
| H7 |
0.092 |
2.590 |
-0.437 |
| H8 |
-0.544 |
1.160 |
-1.408 |
| H9 |
-0.092 |
-2.590 |
-0.437 |
| H10 |
0.544 |
-1.160 |
-1.408 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| C1 | | 1.4732 | 1.3298 | 2.4826 | 1.0850 | 2.1824 | 2.1074 | 2.1092 | 3.4664 | 2.7500 |
C2 | 1.4732 | | 2.4826 | 1.3298 | 2.1824 | 1.0850 | 3.4664 | 2.7500 | 2.1074 | 2.1092 | C3 | 1.3298 | 2.4826 | | 3.0714 | 2.0856 | 3.3408 | 1.0809 | 1.0817 | 4.1216 | 2.9211 | C4 | 2.4826 | 1.3298 | 3.0714 | | 3.3408 | 2.0856 | 4.1216 | 2.9211 | 1.0809 | 1.0817 | H5 | 1.0850 | 2.1824 | 2.0856 | 3.3408 | | 2.5108 | 2.4237 | 3.0615 | 4.2408 | 3.6910 | H6 | 2.1824 | 1.0850 | 3.3408 | 2.0856 | 2.5108 | | 4.2408 | 3.6910 | 2.4237 | 3.0615 | H7 | 2.1074 | 3.4664 | 1.0809 | 4.1216 | 2.4237 | 4.2408 | | 1.8418 | 5.1836 | 3.9000 | H8 | 2.1092 | 2.7500 | 1.0817 | 2.9211 | 3.0615 | 3.6910 | 1.8418 | | 3.9000 | 2.5622 | H9 | 3.4664 | 2.1074 | 4.1216 | 1.0809 | 4.2408 | 2.4237 | 5.1836 | 3.9000 | | 1.8418 | H10 | 2.7500 | 2.1092 | 2.9211 | 1.0817 | 3.6910 | 3.0615 | 3.9000 | 2.5622 | 1.8418 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C4 |
124.599 |
|
C1 |
C2 |
H6 |
116.276 |
| C1 |
C3 |
H7 |
121.548 |
|
C1 |
C3 |
H8 |
121.667 |
| C2 |
C1 |
C3 |
124.599 |
|
C2 |
C1 |
H5 |
116.276 |
| C2 |
C4 |
H9 |
121.548 |
|
C2 |
C4 |
H10 |
121.667 |
| C3 |
C1 |
H5 |
119.117 |
|
C4 |
C2 |
H6 |
119.117 |
| H7 |
C3 |
H8 |
116.781 |
|
H9 |
C4 |
H10 |
116.781 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
