Jump to
S1C2
Energy calculated at QCISD/6-31G*
| | hartrees |
|---|
| Energy at 0K | -155.467208 |
| Energy at 298.15K | -155.472526 |
| HF Energy | -154.917782 |
| Nuclear repulsion energy | 103.277100 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3260 |
3105 |
0.00 |
|
|
|
| 2 |
Ag |
3179 |
3028 |
0.00 |
|
|
|
| 3 |
Ag |
3163 |
3012 |
0.00 |
|
|
|
| 4 |
Ag |
1755 |
1671 |
0.00 |
|
|
|
| 5 |
Ag |
1511 |
1439 |
0.00 |
|
|
|
| 6 |
Ag |
1340 |
1276 |
0.00 |
|
|
|
| 7 |
Ag |
1256 |
1196 |
0.00 |
|
|
|
| 8 |
Ag |
914 |
870 |
0.00 |
|
|
|
| 9 |
Ag |
520 |
495 |
0.00 |
|
|
|
| 10 |
Au |
1043 |
993 |
36.09 |
|
|
|
| 11 |
Au |
922 |
878 |
76.95 |
|
|
|
| 12 |
Au |
531 |
505 |
9.82 |
|
|
|
| 13 |
Au |
155 |
148 |
0.15 |
|
|
|
| 14 |
Bg |
993 |
946 |
0.00 |
|
|
|
| 15 |
Bg |
923 |
879 |
0.00 |
|
|
|
| 16 |
Bg |
767 |
730 |
0.00 |
|
|
|
| 17 |
Bu |
3261 |
3105 |
33.23 |
|
|
|
| 18 |
Bu |
3179 |
3027 |
7.18 |
|
|
|
| 19 |
Bu |
3170 |
3019 |
28.16 |
|
|
|
| 20 |
Bu |
1684 |
1604 |
9.58 |
|
|
|
| 21 |
Bu |
1446 |
1377 |
2.88 |
|
|
|
| 22 |
Bu |
1347 |
1283 |
2.23 |
|
|
|
| 23 |
Bu |
1023 |
974 |
2.06 |
|
|
|
| 24 |
Bu |
299 |
285 |
1.84 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 18819.7 cm
-1
Scaled (by 0.9523) Zero Point Vibrational Energy (zpe) 17922.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD/6-31G*
Point Group is C2h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.733 |
0.000 |
| C2 |
0.000 |
-0.733 |
0.000 |
| C3 |
1.116 |
1.481 |
0.000 |
| C4 |
-1.116 |
-1.481 |
0.000 |
| H5 |
-0.978 |
1.220 |
0.000 |
| H6 |
0.978 |
-1.220 |
0.000 |
| H7 |
1.078 |
2.568 |
0.000 |
| H8 |
2.107 |
1.028 |
0.000 |
| H9 |
-1.078 |
-2.568 |
0.000 |
| H10 |
-2.107 |
-1.028 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| C1 | | 1.4665 | 1.3437 | 2.4798 | 1.0923 | 2.1846 | 2.1278 | 2.1278 | 3.4726 | 2.7465 |
C2 | 1.4665 | | 2.4798 | 1.3437 | 2.1846 | 1.0923 | 3.4726 | 2.7465 | 2.1278 | 2.1278 | C3 | 1.3437 | 2.4798 | | 3.7093 | 2.1100 | 2.7051 | 1.0874 | 1.0895 | 4.6052 | 4.0851 | C4 | 2.4798 | 1.3437 | 3.7093 | | 2.7051 | 2.1100 | 4.6052 | 4.0851 | 1.0874 | 1.0895 | H5 | 1.0923 | 2.1846 | 2.1100 | 2.7051 | | 3.1274 | 2.4577 | 3.0908 | 3.7896 | 2.5165 | H6 | 2.1846 | 1.0923 | 2.7051 | 2.1100 | 3.1274 | | 3.7896 | 2.5165 | 2.4577 | 3.0908 | H7 | 2.1278 | 3.4726 | 1.0874 | 4.6052 | 2.4577 | 3.7896 | | 1.8519 | 5.5698 | 4.8038 | H8 | 2.1278 | 2.7465 | 1.0895 | 4.0851 | 3.0908 | 2.5165 | 1.8519 | | 4.8038 | 4.6894 | H9 | 3.4726 | 2.1278 | 4.6052 | 1.0874 | 3.7896 | 2.4577 | 5.5698 | 4.8038 | | 1.8519 | H10 | 2.7465 | 2.1278 | 4.0851 | 1.0895 | 2.5165 | 3.0908 | 4.8038 | 4.6894 | 1.8519 | |
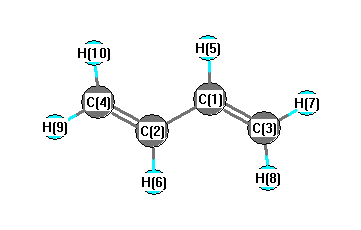 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C4 |
123.820 |
|
C1 |
C2 |
H6 |
116.491 |
| C1 |
C3 |
H7 |
121.794 |
|
C1 |
C3 |
H8 |
121.621 |
| C2 |
C1 |
C3 |
123.820 |
|
C2 |
C1 |
H5 |
116.491 |
| C2 |
C4 |
H9 |
121.794 |
|
C2 |
C4 |
H10 |
121.621 |
| C3 |
C1 |
H5 |
119.689 |
|
C4 |
C2 |
H6 |
119.689 |
| H7 |
C3 |
H8 |
116.584 |
|
H9 |
C4 |
H10 |
116.584 |
Electronic energy levels
Electronic state
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at QCISD/6-31G*
| | hartrees |
|---|
| Energy at 0K | -155.463064 |
| Energy at 298.15K | -155.468392 |
| HF Energy | -154.912940 |
| Nuclear repulsion energy | 104.460081 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3263 |
3108 |
7.23 |
|
|
|
| 2 |
A |
3186 |
3034 |
1.03 |
|
|
|
| 3 |
A |
3171 |
3020 |
28.11 |
|
|
|
| 4 |
A |
1725 |
1643 |
1.20 |
|
|
|
| 5 |
A |
1502 |
1430 |
7.08 |
|
|
|
| 6 |
A |
1366 |
1300 |
0.00 |
|
|
|
| 7 |
A |
1092 |
1040 |
0.07 |
|
|
|
| 8 |
A |
1009 |
961 |
3.19 |
|
|
|
| 9 |
A |
931 |
887 |
6.03 |
|
|
|
| 10 |
A |
903 |
860 |
0.66 |
|
|
|
| 11 |
A |
753 |
718 |
4.32 |
|
|
|
| 12 |
A |
279 |
266 |
0.00 |
|
|
|
| 13 |
A |
168 |
160 |
0.03 |
|
|
|
| 14 |
B |
3261 |
3106 |
25.69 |
|
|
|
| 15 |
B |
3180 |
3028 |
9.48 |
|
|
|
| 16 |
B |
3163 |
3012 |
9.26 |
|
|
|
| 17 |
B |
1724 |
1642 |
2.26 |
|
|
|
| 18 |
B |
1470 |
1400 |
0.82 |
|
|
|
| 19 |
B |
1333 |
1269 |
0.69 |
|
|
|
| 20 |
B |
1122 |
1069 |
4.26 |
|
|
|
| 21 |
B |
1028 |
979 |
32.42 |
|
|
|
| 22 |
B |
933 |
888 |
66.06 |
|
|
|
| 23 |
B |
627 |
597 |
5.81 |
|
|
|
| 24 |
B |
467 |
445 |
9.57 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 18827.8 cm
-1
Scaled (by 0.9523) Zero Point Vibrational Energy (zpe) 17929.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD/6-31G*
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.117 |
0.730 |
0.555 |
| C2 |
-0.117 |
-0.730 |
0.555 |
| C3 |
-0.117 |
1.537 |
-0.491 |
| C4 |
0.117 |
-1.537 |
-0.491 |
| H5 |
0.493 |
1.161 |
1.485 |
| H6 |
-0.493 |
-1.161 |
1.485 |
| H7 |
0.092 |
2.604 |
-0.446 |
| H8 |
-0.535 |
1.158 |
-1.423 |
| H9 |
-0.092 |
-2.604 |
-0.446 |
| H10 |
0.535 |
-1.158 |
-1.423 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| C1 | | 1.4782 | 1.3424 | 2.4969 | 1.0920 | 2.1937 | 2.1248 | 2.1262 | 3.4869 | 2.7663 |
C2 | 1.4782 | | 2.4969 | 1.3424 | 2.1937 | 1.0920 | 3.4869 | 2.7663 | 2.1248 | 2.1262 | C3 | 1.3424 | 2.4969 | | 3.0831 | 2.1027 | 3.3659 | 1.0877 | 1.0887 | 4.1412 | 2.9255 | C4 | 2.4969 | 1.3424 | 3.0831 | | 3.3659 | 2.1027 | 4.1412 | 2.9255 | 1.0877 | 1.0887 | H5 | 1.0920 | 2.1937 | 2.1027 | 3.3659 | | 2.5228 | 2.4439 | 3.0843 | 4.2714 | 3.7199 | H6 | 2.1937 | 1.0920 | 3.3659 | 2.1027 | 2.5228 | | 4.2714 | 3.7199 | 2.4439 | 3.0843 | H7 | 2.1248 | 3.4869 | 1.0877 | 4.1412 | 2.4439 | 4.2714 | | 1.8534 | 5.2107 | 3.9121 | H8 | 2.1262 | 2.7663 | 1.0887 | 2.9255 | 3.0843 | 3.7199 | 1.8534 | | 3.9121 | 2.5521 | H9 | 3.4869 | 2.1248 | 4.1412 | 1.0877 | 4.2714 | 2.4439 | 5.2107 | 3.9121 | | 1.8534 | H10 | 2.7663 | 2.1262 | 2.9255 | 1.0887 | 3.7199 | 3.0843 | 3.9121 | 2.5521 | 1.8534 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C4 |
124.491 |
|
C1 |
C2 |
H6 |
116.385 |
| C1 |
C3 |
H7 |
121.582 |
|
C1 |
C3 |
H8 |
121.642 |
| C2 |
C1 |
C3 |
124.491 |
|
C2 |
C1 |
H5 |
116.385 |
| C2 |
C4 |
H9 |
121.582 |
|
C2 |
C4 |
H10 |
121.642 |
| C3 |
C1 |
H5 |
119.119 |
|
C4 |
C2 |
H6 |
119.119 |
| H7 |
C3 |
H8 |
116.772 |
|
H9 |
C4 |
H10 |
116.772 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
