Jump to
S1C2
Energy calculated at PBEPBEultrafine/6-31G*
| | hartrees |
|---|
| Energy at 0K | -155.767327 |
| Energy at 298.15K | -155.772578 |
| HF Energy | -155.767327 |
| Nuclear repulsion energy | 103.030356 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBEPBEultrafine/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3182 |
3129 |
0.00 |
|
|
|
| 2 |
Ag |
3096 |
3045 |
0.00 |
|
|
|
| 3 |
Ag |
3076 |
3025 |
0.00 |
|
|
|
| 4 |
Ag |
1674 |
1647 |
0.00 |
|
|
|
| 5 |
Ag |
1448 |
1424 |
0.00 |
|
|
|
| 6 |
Ag |
1287 |
1266 |
0.00 |
|
|
|
| 7 |
Ag |
1211 |
1191 |
0.00 |
|
|
|
| 8 |
Ag |
889 |
875 |
0.00 |
|
|
|
| 9 |
Ag |
501 |
493 |
0.00 |
|
|
|
| 10 |
Au |
1029 |
1012 |
28.60 |
|
|
|
| 11 |
Au |
879 |
864 |
79.49 |
|
|
|
| 12 |
Au |
522 |
514 |
8.36 |
|
|
|
| 13 |
Au |
178 |
175 |
0.35 |
|
|
|
| 14 |
Bg |
962 |
946 |
0.00 |
|
|
|
| 15 |
Bg |
884 |
869 |
0.00 |
|
|
|
| 16 |
Bg |
758 |
745 |
0.00 |
|
|
|
| 17 |
Bu |
3182 |
3130 |
28.39 |
|
|
|
| 18 |
Bu |
3095 |
3044 |
7.79 |
|
|
|
| 19 |
Bu |
3086 |
3035 |
31.73 |
|
|
|
| 20 |
Bu |
1625 |
1599 |
12.36 |
|
|
|
| 21 |
Bu |
1389 |
1366 |
3.49 |
|
|
|
| 22 |
Bu |
1287 |
1266 |
2.18 |
|
|
|
| 23 |
Bu |
977 |
960 |
2.66 |
|
|
|
| 24 |
Bu |
285 |
280 |
2.21 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 18251.4 cm
-1
Scaled (by 0.9835) Zero Point Vibrational Energy (zpe) 17950.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBEPBEultrafine/6-31G*
Point Group is C2h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.327 |
0.650 |
0.000 |
| C2 |
0.327 |
-0.650 |
0.000 |
| C3 |
0.327 |
1.832 |
0.000 |
| C4 |
-0.327 |
-1.832 |
0.000 |
| H5 |
-1.426 |
0.642 |
0.000 |
| H6 |
1.426 |
-0.642 |
0.000 |
| H7 |
-0.206 |
2.787 |
0.000 |
| H8 |
1.422 |
1.878 |
0.000 |
| H9 |
0.206 |
-2.787 |
0.000 |
| H10 |
-1.422 |
-1.878 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| C1 | | 1.4557 | 1.3505 | 2.4818 | 1.0991 | 2.1781 | 2.1400 | 2.1373 | 3.4780 | 2.7552 |
C2 | 1.4557 | | 2.4818 | 1.3505 | 2.1781 | 1.0991 | 3.4780 | 2.7552 | 2.1400 | 2.1373 | C3 | 1.3505 | 2.4818 | | 3.7212 | 2.1187 | 2.7069 | 1.0938 | 1.0961 | 4.6199 | 4.1014 | C4 | 2.4818 | 1.3505 | 3.7212 | | 2.7069 | 2.1187 | 4.6199 | 4.1014 | 1.0938 | 1.0961 | H5 | 1.0991 | 2.1781 | 2.1187 | 2.7069 | | 3.1281 | 2.4675 | 3.1050 | 3.7974 | 2.5201 | H6 | 2.1781 | 1.0991 | 2.7069 | 2.1187 | 3.1281 | | 3.7974 | 2.5201 | 2.4675 | 3.1050 | H7 | 2.1400 | 3.4780 | 1.0938 | 4.6199 | 2.4675 | 3.7974 | | 1.8646 | 5.5886 | 4.8207 | H8 | 2.1373 | 2.7552 | 1.0961 | 4.1014 | 3.1050 | 2.5201 | 1.8646 | | 4.8207 | 4.7115 | H9 | 3.4780 | 2.1400 | 4.6199 | 1.0938 | 3.7974 | 2.4675 | 5.5886 | 4.8207 | | 1.8646 | H10 | 2.7552 | 2.1373 | 4.1014 | 1.0961 | 2.5201 | 3.1050 | 4.8207 | 4.7115 | 1.8646 | |
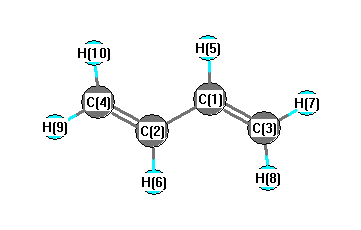 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C4 |
124.315 |
|
C1 |
C2 |
H6 |
116.287 |
| C1 |
C3 |
H7 |
121.861 |
|
C1 |
C3 |
H8 |
121.402 |
| C2 |
C1 |
C3 |
124.315 |
|
C2 |
C1 |
H5 |
116.287 |
| C2 |
C4 |
H9 |
121.861 |
|
C2 |
C4 |
H10 |
121.402 |
| C3 |
C1 |
H5 |
119.398 |
|
C4 |
C2 |
H6 |
119.398 |
| H7 |
C3 |
H8 |
116.738 |
|
H9 |
C4 |
H10 |
116.738 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.069 |
|
|
|
| 2 |
C |
-0.069 |
|
|
|
| 3 |
C |
-0.371 |
|
|
|
| 4 |
C |
-0.371 |
|
|
|
| 5 |
H |
0.135 |
|
|
|
| 6 |
H |
0.135 |
|
|
|
| 7 |
H |
0.155 |
|
|
|
| 8 |
H |
0.151 |
|
|
|
| 9 |
H |
0.155 |
|
|
|
| 10 |
H |
0.151 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-22.510 |
-0.044 |
0.000 |
| y |
-0.044 |
-22.662 |
0.000 |
| z |
0.000 |
0.000 |
-28.010 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.825 |
-0.044 |
0.000 |
| y |
-0.044 |
2.598 |
0.000 |
| z |
0.000 |
0.000 |
-5.424 |
|
| Polar |
|---|
| 3z2-r2 | -10.847 |
|---|
| x2-y2 | 0.151 |
|---|
| xy | -0.044 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.204 |
1.407 |
0.000 |
| y |
1.407 |
11.539 |
0.000 |
| z |
0.000 |
0.000 |
2.345 |
<r2> (average value of r
2) Å
2
| <r2> |
94.743 |
| (<r2>)1/2 |
9.734 |
Jump to
S1C1
Energy calculated at PBEPBEultrafine/6-31G*
| | hartrees |
|---|
| Energy at 0K | -155.761473 |
| Energy at 298.15K | -155.766664 |
| HF Energy | -155.761473 |
| Nuclear repulsion energy | 104.101846 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBEPBEultrafine/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3183 |
3130 |
7.76 |
|
|
|
| 2 |
A |
3100 |
3049 |
0.47 |
|
|
|
| 3 |
A |
3086 |
3035 |
34.27 |
|
|
|
| 4 |
A |
1643 |
1616 |
1.71 |
|
|
|
| 5 |
A |
1439 |
1415 |
6.99 |
|
|
|
| 6 |
A |
1317 |
1296 |
0.22 |
|
|
|
| 7 |
A |
1046 |
1029 |
0.00 |
|
|
|
| 8 |
A |
989 |
973 |
1.06 |
|
|
|
| 9 |
A |
885 |
871 |
2.38 |
|
|
|
| 10 |
A |
875 |
861 |
1.50 |
|
|
|
| 11 |
A |
732 |
720 |
2.45 |
|
|
|
| 12 |
A |
267 |
263 |
0.00 |
|
|
|
| 13 |
A |
151 |
148 |
0.15 |
|
|
|
| 14 |
B |
3181 |
3128 |
19.65 |
|
|
|
| 15 |
B |
3096 |
3045 |
4.60 |
|
|
|
| 16 |
B |
3073 |
3022 |
12.02 |
|
|
|
| 17 |
B |
1667 |
1640 |
2.04 |
|
|
|
| 18 |
B |
1407 |
1384 |
1.68 |
|
|
|
| 19 |
B |
1282 |
1261 |
0.14 |
|
|
|
| 20 |
B |
1079 |
1061 |
6.47 |
|
|
|
| 21 |
B |
998 |
982 |
28.59 |
|
|
|
| 22 |
B |
886 |
871 |
70.73 |
|
|
|
| 23 |
B |
592 |
582 |
4.98 |
|
|
|
| 24 |
B |
465 |
457 |
10.72 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 18219.4 cm
-1
Scaled (by 0.9835) Zero Point Vibrational Energy (zpe) 17918.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBEPBEultrafine/6-31G*
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.087 |
0.729 |
0.560 |
| C2 |
-0.087 |
-0.729 |
0.560 |
| C3 |
-0.087 |
1.546 |
-0.499 |
| C4 |
0.087 |
-1.546 |
-0.499 |
| H5 |
0.361 |
1.177 |
1.526 |
| H6 |
-0.361 |
-1.177 |
1.526 |
| H7 |
0.083 |
2.624 |
-0.423 |
| H8 |
-0.417 |
1.165 |
-1.471 |
| H9 |
-0.083 |
-2.624 |
-0.423 |
| H10 |
0.417 |
-1.165 |
-1.471 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| C1 | | 1.4689 | 1.3489 | 2.5099 | 1.0989 | 2.1832 | 2.1348 | 2.1382 | 3.4988 | 2.7974 |
C2 | 1.4689 | | 2.5099 | 1.3489 | 2.1832 | 1.0989 | 3.4988 | 2.7974 | 2.1348 | 2.1382 | C3 | 1.3489 | 2.5099 | | 3.0973 | 2.1062 | 3.4043 | 1.0940 | 1.0956 | 4.1713 | 2.9242 | C4 | 2.5099 | 1.3489 | 3.0973 | | 3.4043 | 2.1062 | 4.1713 | 2.9242 | 1.0940 | 1.0956 | H5 | 1.0989 | 2.1832 | 2.1062 | 3.4043 | | 2.4622 | 2.4431 | 3.0966 | 4.2945 | 3.8040 | H6 | 2.1832 | 1.0989 | 3.4043 | 2.1062 | 2.4622 | | 4.2945 | 3.8040 | 2.4431 | 3.0966 | H7 | 2.1348 | 3.4988 | 1.0940 | 4.1713 | 2.4431 | 4.2945 | | 1.8653 | 5.2513 | 3.9460 | H8 | 2.1382 | 2.7974 | 1.0956 | 2.9242 | 3.0966 | 3.8040 | 1.8653 | | 3.9460 | 2.4749 | H9 | 3.4988 | 2.1348 | 4.1713 | 1.0940 | 4.2945 | 2.4431 | 5.2513 | 3.9460 | | 1.8653 | H10 | 2.7974 | 2.1382 | 2.9242 | 1.0956 | 3.8040 | 3.0966 | 3.9460 | 2.4749 | 1.8653 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C4 |
125.884 |
|
C1 |
C2 |
H6 |
115.722 |
| C1 |
C3 |
H7 |
121.478 |
|
C1 |
C3 |
H8 |
121.682 |
| C2 |
C1 |
C3 |
125.884 |
|
C2 |
C1 |
H5 |
115.722 |
| C2 |
C4 |
H9 |
121.478 |
|
C2 |
C4 |
H10 |
121.682 |
| C3 |
C1 |
H5 |
118.379 |
|
C4 |
C2 |
H6 |
118.379 |
| H7 |
C3 |
H8 |
116.835 |
|
H9 |
C4 |
H10 |
116.835 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.095 |
|
|
|
| 2 |
C |
-0.095 |
|
|
|
| 3 |
C |
-0.349 |
|
|
|
| 4 |
C |
-0.349 |
|
|
|
| 5 |
H |
0.140 |
|
|
|
| 6 |
H |
0.140 |
|
|
|
| 7 |
H |
0.153 |
|
|
|
| 8 |
H |
0.152 |
|
|
|
| 9 |
H |
0.153 |
|
|
|
| 10 |
H |
0.152 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.119 |
0.119 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-27.559 |
0.617 |
0.000 |
| y |
0.617 |
-22.711 |
0.000 |
| z |
0.000 |
0.000 |
-22.663 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-4.872 |
0.617 |
0.000 |
| y |
0.617 |
2.400 |
0.000 |
| z |
0.000 |
0.000 |
2.472 |
|
| Polar |
|---|
| 3z2-r2 | 4.944 |
|---|
| x2-y2 | -4.848 |
|---|
| xy | 0.617 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.681 |
0.067 |
0.000 |
| y |
0.067 |
9.349 |
0.000 |
| z |
0.000 |
0.000 |
6.740 |
<r2> (average value of r
2) Å
2
| <r2> |
86.416 |
| (<r2>)1/2 |
9.296 |
