Jump to
S1C2
Energy calculated at QCISD/6-31G*
| | hartrees |
|---|
| Energy at 0K | -191.343094 |
| Energy at 298.15K | |
| HF Energy | -190.759493 |
| Nuclear repulsion energy | 102.530022 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3266 |
3110 |
8.75 |
|
|
|
| 2 |
A' |
3218 |
3064 |
1.56 |
|
|
|
| 3 |
A' |
3176 |
3024 |
5.85 |
|
|
|
| 4 |
A' |
2957 |
2816 |
88.55 |
|
|
|
| 5 |
A' |
1810 |
1723 |
166.96 |
|
|
|
| 6 |
A' |
1710 |
1629 |
4.46 |
|
|
|
| 7 |
A' |
1489 |
1418 |
8.02 |
|
|
|
| 8 |
A' |
1424 |
1356 |
3.75 |
|
|
|
| 9 |
A' |
1322 |
1259 |
2.50 |
|
|
|
| 10 |
A' |
1202 |
1144 |
32.37 |
|
|
|
| 11 |
A' |
942 |
897 |
19.83 |
|
|
|
| 12 |
A' |
576 |
548 |
5.33 |
|
|
|
| 13 |
A' |
324 |
308 |
10.36 |
|
|
|
| 14 |
A" |
1029 |
980 |
4.71 |
|
|
|
| 15 |
A" |
1016 |
968 |
24.81 |
|
|
|
| 16 |
A" |
978 |
932 |
29.44 |
|
|
|
| 17 |
A" |
602 |
573 |
9.93 |
|
|
|
| 18 |
A" |
157 |
150 |
5.26 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13597.9 cm
-1
Scaled (by 0.9523) Zero Point Vibrational Energy (zpe) 12949.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD/6-31G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.145 |
-0.752 |
0.000 |
| C2 |
0.000 |
0.722 |
0.000 |
| C3 |
1.214 |
1.294 |
0.000 |
| O4 |
-1.221 |
-1.329 |
0.000 |
| H5 |
0.810 |
-1.321 |
0.000 |
| H6 |
-0.921 |
1.303 |
0.000 |
| H7 |
1.347 |
2.373 |
0.000 |
| H8 |
2.123 |
0.692 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.4808 | 2.4554 | 1.2212 | 1.1120 | 2.1964 | 3.4623 | 2.6883 |
C2 | 1.4808 | | 1.3415 | 2.3871 | 2.1982 | 1.0890 | 2.1305 | 2.1229 | C3 | 2.4554 | 1.3415 | | 3.5786 | 2.6460 | 2.1347 | 1.0873 | 1.0901 | O4 | 1.2212 | 2.3871 | 3.5786 | | 2.0311 | 2.6492 | 4.5053 | 3.9072 | H5 | 1.1120 | 2.1982 | 2.6460 | 2.0311 | | 3.1440 | 3.7329 | 2.4037 | H6 | 2.1964 | 1.0890 | 2.1347 | 2.6492 | 3.1440 | | 2.5076 | 3.1045 | H7 | 3.4623 | 2.1305 | 1.0873 | 4.5053 | 3.7329 | 2.5076 | | 1.8509 | H8 | 2.6883 | 2.1229 | 1.0901 | 3.9072 | 2.4037 | 3.1045 | 1.8509 | |
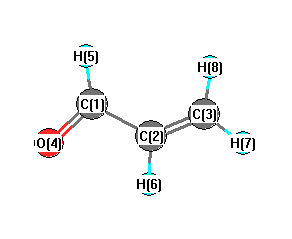 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
120.835 |
|
C1 |
C2 |
H6 |
116.626 |
| C2 |
C1 |
O4 |
123.834 |
|
C2 |
C1 |
H5 |
115.206 |
| C2 |
C3 |
H7 |
122.265 |
|
C2 |
C3 |
H8 |
121.290 |
| C3 |
C2 |
H6 |
122.538 |
|
O4 |
C1 |
H5 |
120.959 |
| H7 |
C3 |
H8 |
116.446 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at QCISD/6-31G*
| | hartrees |
|---|
| Energy at 0K | -191.341016 |
| Energy at 298.15K | |
| HF Energy | -190.756866 |
| Nuclear repulsion energy | 104.196460 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3280 |
3124 |
4.07 |
|
|
|
| 2 |
A' |
3203 |
3050 |
9.69 |
|
|
|
| 3 |
A' |
3186 |
3034 |
6.37 |
|
|
|
| 4 |
A' |
2983 |
2841 |
128.76 |
|
|
|
| 5 |
A' |
1800 |
1714 |
78.48 |
|
|
|
| 6 |
A' |
1709 |
1627 |
44.88 |
|
|
|
| 7 |
A' |
1470 |
1399 |
35.97 |
|
|
|
| 8 |
A' |
1458 |
1388 |
0.39 |
|
|
|
| 9 |
A' |
1336 |
1273 |
1.96 |
|
|
|
| 10 |
A' |
1092 |
1040 |
4.49 |
|
|
|
| 11 |
A' |
957 |
912 |
57.25 |
|
|
|
| 12 |
A' |
690 |
657 |
12.02 |
|
|
|
| 13 |
A' |
298 |
284 |
7.37 |
|
|
|
| 14 |
A" |
1025 |
977 |
0.04 |
|
|
|
| 15 |
A" |
1018 |
969 |
44.93 |
|
|
|
| 16 |
A" |
982 |
936 |
7.15 |
|
|
|
| 17 |
A" |
563 |
536 |
8.93 |
|
|
|
| 18 |
A" |
165 |
157 |
8.03 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13607.3 cm
-1
Scaled (by 0.9523) Zero Point Vibrational Energy (zpe) 12958.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD/6-31G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.896 |
-0.285 |
0.000 |
| C2 |
0.000 |
0.905 |
0.000 |
| C3 |
1.334 |
0.764 |
0.000 |
| O4 |
-0.494 |
-1.439 |
0.000 |
| H5 |
-1.985 |
-0.071 |
0.000 |
| H6 |
-0.474 |
1.887 |
0.000 |
| H7 |
2.004 |
1.621 |
0.000 |
| H8 |
1.780 |
-0.228 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.4892 | 2.4643 | 1.2227 | 1.1094 | 2.2120 | 3.4696 | 2.6765 |
C2 | 1.4892 | | 1.3415 | 2.3960 | 2.2116 | 1.0900 | 2.1277 | 2.1101 | C3 | 2.4643 | 1.3415 | | 2.8635 | 3.4222 | 2.1279 | 1.0872 | 1.0879 | O4 | 1.2227 | 2.3960 | 2.8635 | | 2.0231 | 3.3263 | 3.9504 | 2.5769 | H5 | 1.1094 | 2.2116 | 3.4222 | 2.0231 | | 2.4731 | 4.3323 | 3.7679 | H6 | 2.2120 | 1.0900 | 2.1279 | 3.3263 | 2.4731 | | 2.4916 | 3.0906 | H7 | 3.4696 | 2.1277 | 1.0872 | 3.9504 | 4.3323 | 2.4916 | | 1.8623 | H8 | 2.6765 | 2.1101 | 1.0879 | 2.5769 | 3.7679 | 3.0906 | 1.8623 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
120.962 |
|
C1 |
C2 |
H6 |
117.264 |
| C2 |
C1 |
O4 |
123.847 |
|
C2 |
C1 |
H5 |
115.891 |
| C2 |
C3 |
H7 |
121.997 |
|
C2 |
C3 |
H8 |
120.220 |
| C3 |
C2 |
H6 |
121.774 |
|
O4 |
C1 |
H5 |
120.262 |
| H7 |
C3 |
H8 |
117.783 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
