Jump to
S1C2
Energy calculated at PBE1PBE/6-31G*
| | hartrees |
|---|
| Energy at 0K | -191.684109 |
| Energy at 298.15K | |
| HF Energy | -191.684109 |
| Nuclear repulsion energy | 103.099526 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBE1PBE/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3279 |
3116 |
5.24 |
64.40 |
0.53 |
0.70 |
| 2 |
A' |
3227 |
3067 |
1.66 |
93.80 |
0.32 |
0.48 |
| 3 |
A' |
3183 |
3025 |
5.17 |
66.68 |
0.11 |
0.19 |
| 4 |
A' |
2930 |
2785 |
101.03 |
84.88 |
0.29 |
0.45 |
| 5 |
A' |
1843 |
1751 |
221.48 |
59.95 |
0.38 |
0.55 |
| 6 |
A' |
1720 |
1634 |
2.65 |
27.19 |
0.20 |
0.34 |
| 7 |
A' |
1474 |
1401 |
12.23 |
6.98 |
0.53 |
0.69 |
| 8 |
A' |
1410 |
1340 |
6.30 |
20.92 |
0.44 |
0.61 |
| 9 |
A' |
1309 |
1244 |
3.60 |
6.80 |
0.34 |
0.51 |
| 10 |
A' |
1188 |
1129 |
35.36 |
11.86 |
0.62 |
0.77 |
| 11 |
A' |
934 |
888 |
20.49 |
1.18 |
0.12 |
0.21 |
| 12 |
A' |
577 |
548 |
6.43 |
5.21 |
0.35 |
0.52 |
| 13 |
A' |
318 |
302 |
9.32 |
0.91 |
0.36 |
0.53 |
| 14 |
A" |
1048 |
995 |
8.86 |
2.59 |
0.75 |
0.86 |
| 15 |
A" |
1029 |
978 |
8.90 |
4.09 |
0.75 |
0.86 |
| 16 |
A" |
999 |
950 |
43.32 |
0.51 |
0.75 |
0.86 |
| 17 |
A" |
613 |
582 |
9.32 |
4.48 |
0.75 |
0.86 |
| 18 |
A" |
173 |
165 |
3.91 |
1.61 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 13627.1 cm
-1
Scaled (by 0.9503) Zero Point Vibrational Energy (zpe) 12949.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBE1PBE/6-31G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.151 |
-0.744 |
0.000 |
| C2 |
0.000 |
0.720 |
0.000 |
| C3 |
1.212 |
1.280 |
0.000 |
| O4 |
-1.216 |
-1.320 |
0.000 |
| H5 |
0.810 |
-1.307 |
0.000 |
| H6 |
-0.915 |
1.307 |
0.000 |
| H7 |
1.355 |
2.356 |
0.000 |
| H8 |
2.112 |
0.666 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.4709 | 2.4399 | 1.2110 | 1.1137 | 2.1881 | 3.4466 | 2.6657 |
C2 | 1.4709 | | 1.3354 | 2.3745 | 2.1824 | 1.0875 | 2.1251 | 2.1122 | C3 | 2.4399 | 1.3354 | | 3.5575 | 2.6181 | 2.1277 | 1.0859 | 1.0889 | O4 | 1.2110 | 2.3745 | 3.5575 | | 2.0257 | 2.6438 | 4.4863 | 3.8752 | H5 | 1.1137 | 2.1824 | 2.6181 | 2.0257 | | 3.1316 | 3.7039 | 2.3639 | H6 | 2.1881 | 1.0875 | 2.1277 | 2.6438 | 3.1316 | | 2.5016 | 3.0939 | H7 | 3.4466 | 2.1251 | 1.0859 | 4.4863 | 3.7039 | 2.5016 | | 1.8518 | H8 | 2.6657 | 2.1122 | 1.0889 | 3.8752 | 2.3639 | 3.0939 | 1.8518 | |
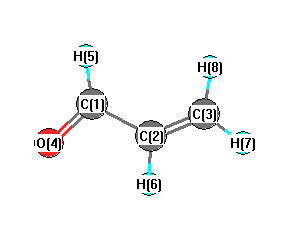 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
120.704 |
|
C1 |
C2 |
H6 |
116.782 |
| C2 |
C1 |
O4 |
124.314 |
|
C2 |
C1 |
H5 |
114.502 |
| C2 |
C3 |
H7 |
122.385 |
|
C2 |
C3 |
H8 |
120.872 |
| C3 |
C2 |
H6 |
122.513 |
|
O4 |
C1 |
H5 |
121.184 |
| H7 |
C3 |
H8 |
116.743 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBE1PBE/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.217 |
|
|
|
| 2 |
C |
-0.152 |
|
|
|
| 3 |
C |
-0.375 |
|
|
|
| 4 |
O |
-0.405 |
|
|
|
| 5 |
H |
0.129 |
|
|
|
| 6 |
H |
0.196 |
|
|
|
| 7 |
H |
0.198 |
|
|
|
| 8 |
H |
0.191 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
2.409 |
2.074 |
0.000 |
3.179 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-23.847 |
-2.505 |
0.000 |
| y |
-2.505 |
-23.471 |
0.000 |
| z |
0.000 |
0.000 |
-24.427 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.102 |
-2.505 |
0.000 |
| y |
-2.505 |
0.666 |
0.000 |
| z |
0.000 |
0.000 |
-0.768 |
|
| Polar |
|---|
| 3z2-r2 | -1.536 |
|---|
| x2-y2 | -0.376 |
|---|
| xy | -2.505 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.653 |
1.681 |
0.000 |
| y |
1.681 |
6.093 |
0.000 |
| z |
0.000 |
0.000 |
2.080 |
<r2> (average value of r
2) Å
2
| <r2> |
83.112 |
| (<r2>)1/2 |
9.117 |
Jump to
S1C1
Energy calculated at PBE1PBE/6-31G*
| | hartrees |
|---|
| Energy at 0K | -191.681440 |
| Energy at 298.15K | |
| HF Energy | -191.681440 |
| Nuclear repulsion energy | 104.729608 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBE1PBE/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3292 |
3128 |
1.94 |
52.46 |
0.65 |
0.79 |
| 2 |
A' |
3213 |
3054 |
9.97 |
123.19 |
0.22 |
0.36 |
| 3 |
A' |
3193 |
3035 |
5.17 |
42.66 |
0.18 |
0.30 |
| 4 |
A' |
2961 |
2814 |
145.90 |
142.91 |
0.32 |
0.49 |
| 5 |
A' |
1840 |
1749 |
107.08 |
13.52 |
0.49 |
0.66 |
| 6 |
A' |
1710 |
1625 |
66.36 |
43.96 |
0.22 |
0.37 |
| 7 |
A' |
1455 |
1383 |
31.87 |
9.71 |
0.74 |
0.85 |
| 8 |
A' |
1443 |
1371 |
13.20 |
8.42 |
0.25 |
0.40 |
| 9 |
A' |
1326 |
1260 |
2.14 |
15.46 |
0.44 |
0.61 |
| 10 |
A' |
1076 |
1023 |
5.26 |
4.76 |
0.73 |
0.84 |
| 11 |
A' |
957 |
909 |
60.66 |
4.66 |
0.21 |
0.35 |
| 12 |
A' |
688 |
653 |
11.15 |
0.71 |
0.74 |
0.85 |
| 13 |
A' |
288 |
274 |
7.17 |
3.07 |
0.47 |
0.64 |
| 14 |
A" |
1046 |
994 |
7.88 |
9.40 |
0.75 |
0.86 |
| 15 |
A" |
1036 |
984 |
28.38 |
0.76 |
0.75 |
0.86 |
| 16 |
A" |
1010 |
959 |
16.39 |
0.27 |
0.75 |
0.86 |
| 17 |
A" |
565 |
537 |
8.31 |
6.49 |
0.75 |
0.86 |
| 18 |
A" |
183 |
174 |
7.64 |
0.86 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 13640.2 cm
-1
Scaled (by 0.9503) Zero Point Vibrational Energy (zpe) 12962.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBE1PBE/6-31G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.888 |
-0.285 |
0.000 |
| C2 |
0.000 |
0.899 |
0.000 |
| C3 |
1.328 |
0.763 |
0.000 |
| O4 |
-0.495 |
-1.432 |
0.000 |
| H5 |
-1.976 |
-0.064 |
0.000 |
| H6 |
-0.474 |
1.879 |
0.000 |
| H7 |
1.998 |
1.618 |
0.000 |
| H8 |
1.768 |
-0.232 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.4803 | 2.4513 | 1.2123 | 1.1107 | 2.2037 | 3.4564 | 2.6563 |
C2 | 1.4803 | | 1.3353 | 2.3835 | 2.1983 | 1.0883 | 2.1229 | 2.0988 | C3 | 2.4513 | 1.3353 | | 2.8534 | 3.4062 | 2.1198 | 1.0859 | 1.0872 | O4 | 1.2123 | 2.3835 | 2.8534 | | 2.0164 | 3.3114 | 3.9390 | 2.5619 | H5 | 1.1107 | 2.1983 | 3.4062 | 2.0164 | | 2.4561 | 4.3149 | 3.7479 | H6 | 2.2037 | 1.0883 | 2.1198 | 3.3114 | 2.4561 | | 2.4852 | 3.0790 | H7 | 3.4564 | 2.1229 | 1.0859 | 3.9390 | 4.3149 | 2.4852 | | 1.8634 | H8 | 2.6563 | 2.0988 | 1.0872 | 2.5619 | 3.7479 | 3.0790 | 1.8634 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
120.972 |
|
C1 |
C2 |
H6 |
117.353 |
| C2 |
C1 |
O4 |
124.255 |
|
C2 |
C1 |
H5 |
115.346 |
| C2 |
C3 |
H7 |
122.186 |
|
C2 |
C3 |
H8 |
119.734 |
| C3 |
C2 |
H6 |
121.675 |
|
O4 |
C1 |
H5 |
120.398 |
| H7 |
C3 |
H8 |
118.080 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBE1PBE/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.208 |
|
|
|
| 2 |
C |
-0.184 |
|
|
|
| 3 |
C |
-0.342 |
|
|
|
| 4 |
O |
-0.399 |
|
|
|
| 5 |
H |
0.134 |
|
|
|
| 6 |
H |
0.178 |
|
|
|
| 7 |
H |
0.191 |
|
|
|
| 8 |
H |
0.213 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.281 |
2.634 |
0.000 |
2.649 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-19.801 |
-0.653 |
0.000 |
| y |
-0.653 |
-25.529 |
0.000 |
| z |
0.000 |
0.000 |
-24.402 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
5.165 |
-0.653 |
0.000 |
| y |
-0.653 |
-3.428 |
0.000 |
| z |
0.000 |
0.000 |
-1.737 |
|
| Polar |
|---|
| 3z2-r2 | -3.474 |
|---|
| x2-y2 | 5.729 |
|---|
| xy | -0.653 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.822 |
0.507 |
0.000 |
| y |
0.507 |
5.477 |
0.000 |
| z |
0.000 |
0.000 |
2.077 |
<r2> (average value of r
2) Å
2
| <r2> |
74.486 |
| (<r2>)1/2 |
8.630 |
