Jump to
S1C2
Energy calculated at B2PLYP/6-31G*
| | hartrees |
|---|
| Energy at 0K | -191.706767 |
| Energy at 298.15K | |
| HF Energy | -191.521828 |
| Nuclear repulsion energy | 102.799038 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3285 |
3119 |
6.85 |
64.16 |
0.54 |
0.70 |
| 2 |
A' |
3232 |
3068 |
1.89 |
90.69 |
0.31 |
0.48 |
| 3 |
A' |
3192 |
3030 |
5.27 |
67.24 |
0.11 |
0.20 |
| 4 |
A' |
2943 |
2793 |
101.87 |
84.51 |
0.28 |
0.44 |
| 5 |
A' |
1786 |
1696 |
183.02 |
59.56 |
0.37 |
0.54 |
| 6 |
A' |
1700 |
1614 |
2.64 |
13.71 |
0.17 |
0.29 |
| 7 |
A' |
1487 |
1411 |
9.27 |
7.12 |
0.50 |
0.67 |
| 8 |
A' |
1420 |
1348 |
4.61 |
21.15 |
0.43 |
0.60 |
| 9 |
A' |
1320 |
1253 |
2.70 |
7.88 |
0.34 |
0.51 |
| 10 |
A' |
1194 |
1133 |
35.79 |
12.68 |
0.63 |
0.77 |
| 11 |
A' |
940 |
892 |
21.00 |
0.99 |
0.10 |
0.17 |
| 12 |
A' |
575 |
545 |
5.51 |
5.37 |
0.35 |
0.52 |
| 13 |
A' |
323 |
306 |
9.66 |
0.69 |
0.33 |
0.49 |
| 14 |
A" |
1046 |
993 |
10.14 |
2.16 |
0.75 |
0.86 |
| 15 |
A" |
1025 |
973 |
12.49 |
4.44 |
0.75 |
0.86 |
| 16 |
A" |
992 |
941 |
32.62 |
0.41 |
0.75 |
0.86 |
| 17 |
A" |
614 |
583 |
8.36 |
4.83 |
0.75 |
0.86 |
| 18 |
A" |
171 |
162 |
4.18 |
1.46 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 13622.0 cm
-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 12930.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP/6-31G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.148 |
-0.746 |
0.000 |
| C2 |
0.000 |
0.721 |
0.000 |
| C3 |
1.214 |
1.287 |
0.000 |
| O4 |
-1.220 |
-1.326 |
0.000 |
| H5 |
0.811 |
-1.308 |
0.000 |
| H6 |
-0.915 |
1.305 |
0.000 |
| H7 |
1.351 |
2.362 |
0.000 |
| H8 |
2.114 |
0.679 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.4736 | 2.4462 | 1.2194 | 1.1110 | 2.1892 | 3.4502 | 2.6731 |
C2 | 1.4736 | | 1.3394 | 2.3827 | 2.1845 | 1.0857 | 2.1262 | 2.1148 | C3 | 2.4462 | 1.3394 | | 3.5708 | 2.6260 | 2.1290 | 1.0839 | 1.0867 | O4 | 1.2194 | 2.3827 | 3.5708 | | 2.0306 | 2.6484 | 4.4961 | 3.8907 | H5 | 1.1110 | 2.1845 | 2.6260 | 2.0306 | | 3.1311 | 3.7095 | 2.3764 | H6 | 2.1892 | 1.0857 | 2.1290 | 2.6484 | 3.1311 | | 2.5012 | 3.0936 | H7 | 3.4502 | 2.1262 | 1.0839 | 4.4961 | 3.7095 | 2.5012 | | 1.8480 | H8 | 2.6731 | 2.1148 | 1.0867 | 3.8907 | 2.3764 | 3.0936 | 1.8480 | |
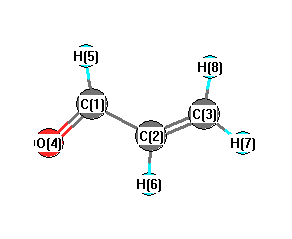 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
120.759 |
|
C1 |
C2 |
H6 |
116.797 |
| C2 |
C1 |
O4 |
124.178 |
|
C2 |
C1 |
H5 |
114.663 |
| C2 |
C3 |
H7 |
122.316 |
|
C2 |
C3 |
H8 |
120.963 |
| C3 |
C2 |
H6 |
122.444 |
|
O4 |
C1 |
H5 |
121.159 |
| H7 |
C3 |
H8 |
116.721 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.225 |
|
|
|
| 2 |
C |
-0.130 |
|
|
|
| 3 |
C |
-0.334 |
|
|
|
| 4 |
O |
-0.399 |
|
|
|
| 5 |
H |
0.115 |
|
|
|
| 6 |
H |
0.175 |
|
|
|
| 7 |
H |
0.178 |
|
|
|
| 8 |
H |
0.171 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
2.387 |
2.040 |
0.000 |
3.140 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-24.175 |
-2.531 |
-0.002 |
| y |
-2.531 |
-23.821 |
-0.002 |
| z |
-0.002 |
-0.002 |
-24.453 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.038 |
-2.531 |
-0.002 |
| y |
-2.531 |
0.493 |
-0.002 |
| z |
-0.002 |
-0.002 |
-0.455 |
|
| Polar |
|---|
| 3z2-r2 | -0.910 |
|---|
| x2-y2 | -0.354 |
|---|
| xy | -2.531 |
|---|
| xz | -0.002 |
|---|
| yz | -0.002 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.593 |
1.651 |
0.000 |
| y |
1.651 |
6.038 |
0.000 |
| z |
0.000 |
0.000 |
2.045 |
<r2> (average value of r
2) Å
2
| <r2> |
83.660 |
| (<r2>)1/2 |
9.147 |
Jump to
S1C1
Energy calculated at B2PLYP/6-31G*
| | hartrees |
|---|
| Energy at 0K | -191.704305 |
| Energy at 298.15K | |
| HF Energy | -191.519232 |
| Nuclear repulsion energy | 104.415034 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3299 |
3131 |
2.97 |
51.71 |
0.66 |
0.79 |
| 2 |
A' |
3219 |
3055 |
10.81 |
127.17 |
0.20 |
0.34 |
| 3 |
A' |
3202 |
3040 |
5.45 |
37.48 |
0.22 |
0.36 |
| 4 |
A' |
2975 |
2824 |
146.44 |
142.14 |
0.32 |
0.48 |
| 5 |
A' |
1787 |
1696 |
75.55 |
9.11 |
0.52 |
0.68 |
| 6 |
A' |
1690 |
1604 |
67.34 |
41.40 |
0.24 |
0.39 |
| 7 |
A' |
1467 |
1392 |
40.45 |
9.13 |
0.58 |
0.73 |
| 8 |
A' |
1455 |
1382 |
0.46 |
10.72 |
0.34 |
0.50 |
| 9 |
A' |
1336 |
1268 |
2.29 |
15.04 |
0.44 |
0.61 |
| 10 |
A' |
1086 |
1031 |
4.68 |
4.94 |
0.73 |
0.84 |
| 11 |
A' |
953 |
905 |
59.85 |
4.34 |
0.20 |
0.34 |
| 12 |
A' |
688 |
653 |
10.86 |
0.86 |
0.72 |
0.84 |
| 13 |
A' |
293 |
278 |
7.18 |
2.90 |
0.47 |
0.64 |
| 14 |
A" |
1037 |
985 |
14.89 |
8.10 |
0.75 |
0.86 |
| 15 |
A" |
1036 |
983 |
21.10 |
1.62 |
0.75 |
0.86 |
| 16 |
A" |
1000 |
949 |
11.82 |
0.55 |
0.75 |
0.86 |
| 17 |
A" |
567 |
538 |
7.44 |
6.97 |
0.75 |
0.86 |
| 18 |
A" |
183 |
174 |
7.32 |
0.87 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 13635.7 cm
-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 12943.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP/6-31G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.891 |
-0.284 |
0.000 |
| C2 |
0.000 |
0.902 |
0.000 |
| C3 |
1.332 |
0.765 |
0.000 |
| O4 |
-0.496 |
-1.439 |
0.000 |
| H5 |
-1.977 |
-0.063 |
0.000 |
| H6 |
-0.473 |
1.880 |
0.000 |
| H7 |
1.999 |
1.620 |
0.000 |
| H8 |
1.774 |
-0.225 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.4831 | 2.4585 | 1.2208 | 1.1079 | 2.2037 | 3.4607 | 2.6661 |
C2 | 1.4831 | | 1.3392 | 2.3928 | 2.1994 | 1.0865 | 2.1240 | 2.1021 | C3 | 2.4585 | 1.3392 | | 2.8641 | 3.4110 | 2.1215 | 1.0838 | 1.0849 | O4 | 1.2208 | 2.3928 | 2.8641 | | 2.0214 | 3.3189 | 3.9475 | 2.5747 | H5 | 1.1079 | 2.1994 | 3.4110 | 2.0214 | | 2.4564 | 4.3170 | 3.7546 | H6 | 2.2037 | 1.0865 | 2.1215 | 3.3189 | 2.4564 | | 2.4856 | 3.0794 | H7 | 3.4607 | 2.1240 | 1.0838 | 3.9475 | 4.3170 | 2.4856 | | 1.8588 | H8 | 2.6661 | 2.1021 | 1.0849 | 2.5747 | 3.7546 | 3.0794 | 1.8588 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
121.088 |
|
C1 |
C2 |
H6 |
117.261 |
| C2 |
C1 |
O4 |
124.203 |
|
C2 |
C1 |
H5 |
115.412 |
| C2 |
C3 |
H7 |
122.120 |
|
C2 |
C3 |
H8 |
119.894 |
| C3 |
C2 |
H6 |
121.651 |
|
O4 |
C1 |
H5 |
120.385 |
| H7 |
C3 |
H8 |
117.986 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.218 |
|
|
|
| 2 |
C |
-0.159 |
|
|
|
| 3 |
C |
-0.305 |
|
|
|
| 4 |
O |
-0.393 |
|
|
|
| 5 |
H |
0.119 |
|
|
|
| 6 |
H |
0.158 |
|
|
|
| 7 |
H |
0.170 |
|
|
|
| 8 |
H |
0.193 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.231 |
2.610 |
0.000 |
2.620 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-20.140 |
-0.687 |
-0.001 |
| y |
-0.687 |
-25.856 |
-0.002 |
| z |
-0.001 |
-0.002 |
-24.430 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
5.003 |
-0.687 |
-0.001 |
| y |
-0.687 |
-3.571 |
-0.002 |
| z |
-0.001 |
-0.002 |
-1.432 |
|
| Polar |
|---|
| 3z2-r2 | -2.863 |
|---|
| x2-y2 | 5.716 |
|---|
| xy | -0.687 |
|---|
| xz | -0.001 |
|---|
| yz | -0.002 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.729 |
0.481 |
0.000 |
| y |
0.481 |
5.475 |
0.000 |
| z |
0.000 |
0.000 |
2.041 |
<r2> (average value of r
2) Å
2
| <r2> |
74.977 |
| (<r2>)1/2 |
8.659 |
