Jump to
S1C2
Energy calculated at HSEh1PBE/6-31G*
| | hartrees |
|---|
| Energy at 0K | -191.700891 |
| Energy at 298.15K | |
| HF Energy | -191.700891 |
| Nuclear repulsion energy | 103.112476 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HSEh1PBE/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3278 |
3117 |
5.69 |
64.95 |
0.53 |
0.69 |
| 2 |
A' |
3226 |
3068 |
1.87 |
94.18 |
0.32 |
0.48 |
| 3 |
A' |
3182 |
3026 |
5.52 |
67.18 |
0.11 |
0.19 |
| 4 |
A' |
2926 |
2782 |
102.91 |
85.20 |
0.29 |
0.45 |
| 5 |
A' |
1839 |
1749 |
221.52 |
59.81 |
0.39 |
0.56 |
| 6 |
A' |
1718 |
1634 |
2.70 |
27.04 |
0.20 |
0.34 |
| 7 |
A' |
1474 |
1402 |
12.04 |
6.97 |
0.53 |
0.69 |
| 8 |
A' |
1410 |
1341 |
6.30 |
21.04 |
0.44 |
0.61 |
| 9 |
A' |
1309 |
1245 |
3.60 |
6.86 |
0.34 |
0.51 |
| 10 |
A' |
1188 |
1130 |
35.49 |
11.89 |
0.62 |
0.77 |
| 11 |
A' |
934 |
888 |
20.31 |
1.20 |
0.12 |
0.21 |
| 12 |
A' |
577 |
548 |
6.44 |
5.26 |
0.35 |
0.52 |
| 13 |
A' |
319 |
303 |
9.39 |
0.91 |
0.36 |
0.53 |
| 14 |
A" |
1048 |
996 |
8.61 |
2.57 |
0.75 |
0.86 |
| 15 |
A" |
1029 |
979 |
8.94 |
4.08 |
0.75 |
0.86 |
| 16 |
A" |
999 |
950 |
42.58 |
0.50 |
0.75 |
0.86 |
| 17 |
A" |
613 |
583 |
9.09 |
4.48 |
0.75 |
0.86 |
| 18 |
A" |
174 |
166 |
3.90 |
1.61 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 13620.7 cm
-1
Scaled (by 0.951) Zero Point Vibrational Energy (zpe) 12953.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HSEh1PBE/6-31G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.151 |
-0.743 |
0.000 |
| C2 |
0.000 |
0.719 |
0.000 |
| C3 |
1.212 |
1.280 |
0.000 |
| O4 |
-1.216 |
-1.320 |
0.000 |
| H5 |
0.810 |
-1.306 |
0.000 |
| H6 |
-0.915 |
1.307 |
0.000 |
| H7 |
1.356 |
2.356 |
0.000 |
| H8 |
2.111 |
0.665 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.4703 | 2.4392 | 1.2111 | 1.1137 | 2.1874 | 3.4458 | 2.6648 |
C2 | 1.4703 | | 1.3354 | 2.3743 | 2.1814 | 1.0873 | 2.1251 | 2.1118 | C3 | 2.4392 | 1.3354 | | 3.5571 | 2.6168 | 2.1276 | 1.0857 | 1.0887 | O4 | 1.2111 | 2.3743 | 3.5571 | | 2.0259 | 2.6434 | 4.4860 | 3.8743 | H5 | 1.1137 | 2.1814 | 2.6168 | 2.0259 | | 3.1307 | 3.7023 | 2.3622 | H6 | 2.1874 | 1.0873 | 2.1276 | 2.6434 | 3.1307 | | 2.5019 | 3.0936 | H7 | 3.4458 | 2.1251 | 1.0857 | 4.4860 | 3.7023 | 2.5019 | | 1.8514 | H8 | 2.6648 | 2.1118 | 1.0887 | 3.8743 | 2.3622 | 3.0936 | 1.8514 | |
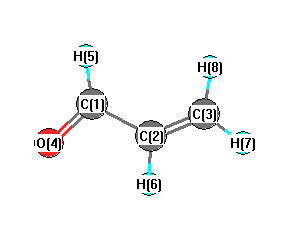 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
120.697 |
|
C1 |
C2 |
H6 |
116.778 |
| C2 |
C1 |
O4 |
124.334 |
|
C2 |
C1 |
H5 |
114.471 |
| C2 |
C3 |
H7 |
122.410 |
|
C2 |
C3 |
H8 |
120.856 |
| C3 |
C2 |
H6 |
122.525 |
|
O4 |
C1 |
H5 |
121.195 |
| H7 |
C3 |
H8 |
116.734 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.217 |
|
|
|
| 2 |
C |
-0.145 |
|
|
|
| 3 |
C |
-0.371 |
|
|
|
| 4 |
O |
-0.405 |
|
|
|
| 5 |
H |
0.127 |
|
|
|
| 6 |
H |
0.193 |
|
|
|
| 7 |
H |
0.196 |
|
|
|
| 8 |
H |
0.189 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
2.408 |
2.081 |
0.000 |
3.182 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-23.865 |
-2.499 |
0.000 |
| y |
-2.499 |
-23.481 |
0.000 |
| z |
0.000 |
0.000 |
-24.409 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.080 |
-2.499 |
0.000 |
| y |
-2.499 |
0.656 |
0.000 |
| z |
0.000 |
0.000 |
-0.736 |
|
| Polar |
|---|
| 3z2-r2 | -1.471 |
|---|
| x2-y2 | -0.384 |
|---|
| xy | -2.499 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.661 |
1.682 |
0.000 |
| y |
1.682 |
6.104 |
0.000 |
| z |
0.000 |
0.000 |
2.078 |
<r2> (average value of r
2) Å
2
| <r2> |
83.093 |
| (<r2>)1/2 |
9.116 |
Jump to
S1C1
Energy calculated at HSEh1PBE/6-31G*
| | hartrees |
|---|
| Energy at 0K | -191.698170 |
| Energy at 298.15K | |
| HF Energy | -191.698170 |
| Nuclear repulsion energy | 104.742968 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HSEh1PBE/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3290 |
3128 |
2.21 |
52.85 |
0.65 |
0.79 |
| 2 |
A' |
3211 |
3054 |
10.48 |
123.95 |
0.22 |
0.36 |
| 3 |
A' |
3192 |
3035 |
5.54 |
42.91 |
0.18 |
0.30 |
| 4 |
A' |
2958 |
2813 |
148.55 |
143.44 |
0.32 |
0.49 |
| 5 |
A' |
1838 |
1748 |
106.42 |
13.32 |
0.49 |
0.66 |
| 6 |
A' |
1708 |
1624 |
67.37 |
44.20 |
0.23 |
0.37 |
| 7 |
A' |
1455 |
1384 |
31.52 |
9.78 |
0.74 |
0.85 |
| 8 |
A' |
1443 |
1372 |
13.45 |
8.41 |
0.25 |
0.39 |
| 9 |
A' |
1326 |
1261 |
2.20 |
15.57 |
0.44 |
0.61 |
| 10 |
A' |
1076 |
1023 |
5.23 |
4.82 |
0.73 |
0.84 |
| 11 |
A' |
956 |
910 |
60.62 |
4.69 |
0.21 |
0.34 |
| 12 |
A' |
688 |
654 |
11.15 |
0.70 |
0.74 |
0.85 |
| 13 |
A' |
288 |
274 |
7.23 |
3.08 |
0.47 |
0.64 |
| 14 |
A" |
1046 |
994 |
7.74 |
9.43 |
0.75 |
0.86 |
| 15 |
A" |
1036 |
985 |
27.80 |
0.72 |
0.75 |
0.86 |
| 16 |
A" |
1009 |
959 |
16.24 |
0.27 |
0.75 |
0.86 |
| 17 |
A" |
565 |
537 |
8.09 |
6.53 |
0.75 |
0.86 |
| 18 |
A" |
184 |
175 |
7.57 |
0.87 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 13633.0 cm
-1
Scaled (by 0.951) Zero Point Vibrational Energy (zpe) 12965.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HSEh1PBE/6-31G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.888 |
-0.285 |
0.000 |
| C2 |
0.000 |
0.899 |
0.000 |
| C3 |
1.328 |
0.762 |
0.000 |
| O4 |
-0.495 |
-1.432 |
0.000 |
| H5 |
-1.976 |
-0.063 |
0.000 |
| H6 |
-0.474 |
1.879 |
0.000 |
| H7 |
1.998 |
1.616 |
0.000 |
| H8 |
1.767 |
-0.232 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.4799 | 2.4507 | 1.2124 | 1.1106 | 2.2031 | 3.4557 | 2.6553 |
C2 | 1.4799 | | 1.3352 | 2.3832 | 2.1978 | 1.0881 | 2.1229 | 2.0984 | C3 | 2.4507 | 1.3352 | | 2.8528 | 3.4055 | 2.1197 | 1.0856 | 1.0870 | O4 | 1.2124 | 2.3832 | 2.8528 | | 2.0165 | 3.3110 | 3.9381 | 2.5608 | H5 | 1.1106 | 2.1978 | 3.4055 | 2.0165 | | 2.4554 | 4.3143 | 3.7469 | H6 | 2.2031 | 1.0881 | 2.1197 | 3.3110 | 2.4554 | | 2.4855 | 3.0786 | H7 | 3.4557 | 2.1229 | 1.0856 | 3.9381 | 4.3143 | 2.4855 | | 1.8630 | H8 | 2.6553 | 2.0984 | 1.0870 | 2.5608 | 3.7469 | 3.0786 | 1.8630 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
120.960 |
|
C1 |
C2 |
H6 |
117.351 |
| C2 |
C1 |
O4 |
124.259 |
|
C2 |
C1 |
H5 |
115.337 |
| C2 |
C3 |
H7 |
122.210 |
|
C2 |
C3 |
H8 |
119.716 |
| C3 |
C2 |
H6 |
121.689 |
|
O4 |
C1 |
H5 |
120.404 |
| H7 |
C3 |
H8 |
118.073 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.209 |
|
|
|
| 2 |
C |
-0.177 |
|
|
|
| 3 |
C |
-0.338 |
|
|
|
| 4 |
O |
-0.399 |
|
|
|
| 5 |
H |
0.131 |
|
|
|
| 6 |
H |
0.175 |
|
|
|
| 7 |
H |
0.188 |
|
|
|
| 8 |
H |
0.211 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.285 |
2.637 |
0.000 |
2.652 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-19.822 |
-0.649 |
0.000 |
| y |
-0.649 |
-25.542 |
0.000 |
| z |
0.000 |
0.000 |
-24.384 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
5.141 |
-0.649 |
0.000 |
| y |
-0.649 |
-3.439 |
0.000 |
| z |
0.000 |
0.000 |
-1.702 |
|
| Polar |
|---|
| 3z2-r2 | -3.404 |
|---|
| x2-y2 | 5.720 |
|---|
| xy | -0.649 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.831 |
0.509 |
0.000 |
| y |
0.509 |
5.486 |
0.000 |
| z |
0.000 |
0.000 |
2.075 |
<r2> (average value of r
2) Å
2
| <r2> |
74.466 |
| (<r2>)1/2 |
8.629 |
