return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C3H7SH (1-Propanethiol)

using model chemistry: CID/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at CID/6-31G*
 hartrees
Energy at 0K-516.255675
Energy at 298.15K-516.264299
HF Energy-515.770356
Nuclear repulsion energy163.331641
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CID/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3230 2984 32.07      
2 A' 3179 2936 33.43      
3 A' 3157 2916 10.04      
4 A' 3149 2908 22.59      
5 A' 2830 2614 23.41      
6 A' 1601 1479 4.39      
7 A' 1586 1465 0.11      
8 A' 1584 1463 5.10      
9 A' 1510 1395 1.61      
10 A' 1474 1361 8.12      
11 A' 1363 1259 28.87      
12 A' 1201 1109 5.86      
13 A' 1114 1029 0.02      
14 A' 993 917 1.08      
15 A' 916 846 4.04      
16 A' 791 730 1.92      
17 A' 374 346 0.46      
18 A' 238 220 2.45      
19 A" 3238 2991 45.47      
20 A" 3223 2977 15.52      
21 A" 3194 2951 1.92      
22 A" 1590 1468 7.68      
23 A" 1398 1291 0.01      
24 A" 1324 1223 0.51      
25 A" 1143 1055 2.44      
26 A" 921 851 0.00      
27 A" 788 728 3.11      
28 A" 257 237 0.26      
29 A" 188 174 19.42      
30 A" 112 103 5.48      

Unscaled Zero Point Vibrational Energy (zpe) 23831.7 cm-1
Scaled (by 0.9237) Zero Point Vibrational Energy (zpe) 22013.4 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at CID/6-31G*
ABC
0.80906 0.07976 0.07566

See section I.F.4 to change rotational constant units
Geometric Data calculated at CID/6-31G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
H1 3.046 -1.079 0.000
H2 2.636 0.390 0.882
H3 2.636 0.390 -0.882
C4 2.396 -0.205 0.000
H5 0.717 -1.235 0.876
H6 0.717 -1.235 -0.876
C7 0.928 -0.619 0.000
H8 0.185 1.200 0.883
H9 0.185 1.200 -0.883
C10 0.000 0.590 0.000
H11 -2.300 1.245 0.000
S12 -1.735 0.033 0.000

Atom - Atom Distances (Å)
  H1 H2 H3 C4 H5 H6 C7 H8 H9 C10 H11 S12
H11.76191.76191.08972.49292.49292.16773.76293.76293.47345.82924.9093
H21.76191.76351.09072.51463.06792.17112.58163.12692.78715.08644.4740
H31.76191.76351.09073.06792.51462.17113.12692.58162.78715.08644.4740
C41.08971.09071.09072.15512.15511.52492.76402.76402.52404.91404.1379
H52.49292.51463.06792.15511.75161.09142.49253.05032.14744.00252.8969
H62.49293.06792.51462.15511.75161.09143.05032.49252.14744.00252.8969
C72.16772.17112.17111.52491.09141.09142.15362.15361.52363.72702.7420
H83.76292.58163.12692.76402.49253.05032.15361.76501.08882.63702.4141
H93.76293.12692.58162.76403.05032.49252.15361.76501.08882.63702.4141
C103.47342.78712.78712.52402.14742.14741.52361.08881.08882.39111.8224
H115.82925.08645.08644.91404.00254.00253.72702.63702.63702.39111.3367
S124.90934.47404.47404.13792.89692.89692.74202.41412.41411.82241.3367

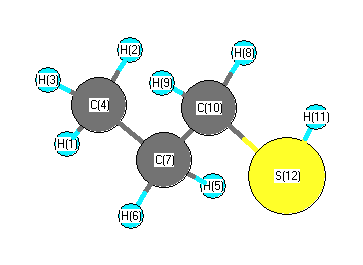
picture of 1-Propanethiol state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
H1 C4 H2 107.816 H1 C4 H3 107.816
H1 C4 C7 110.917 H2 C4 H3 107.886
H2 C4 C7 111.128 H3 C4 C7 111.128
C4 C7 H5 109.806 C4 C7 H6 109.806
C4 C7 C10 111.782 H5 C7 H6 106.725
H5 C7 C10 109.294 H6 C7 C10 109.294
C7 C10 H8 109.939 C7 C10 H9 109.939
C7 C10 S12 109.746 H8 C10 H9 108.294
H8 C10 S12 109.449 H9 C10 S12 109.449
C10 S12 H11 97.188
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability