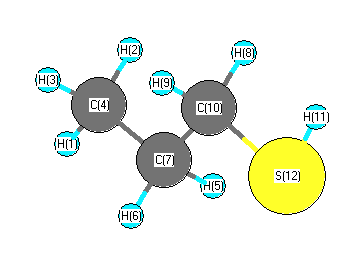
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3136 |
3001 |
30.82 |
|
|
|
| 2 |
A' |
3076 |
2942 |
41.59 |
|
|
|
| 3 |
A' |
3059 |
2927 |
8.14 |
|
|
|
| 4 |
A' |
3054 |
2922 |
21.40 |
|
|
|
| 5 |
A' |
2713 |
2595 |
25.08 |
|
|
|
| 6 |
A' |
1532 |
1466 |
5.22 |
|
|
|
| 7 |
A' |
1518 |
1453 |
0.93 |
|
|
|
| 8 |
A' |
1512 |
1446 |
3.53 |
|
|
|
| 9 |
A' |
1435 |
1373 |
2.34 |
|
|
|
| 10 |
A' |
1394 |
1334 |
5.43 |
|
|
|
| 11 |
A' |
1289 |
1233 |
32.70 |
|
|
|
| 12 |
A' |
1147 |
1098 |
5.76 |
|
|
|
| 13 |
A' |
1066 |
1020 |
0.09 |
|
|
|
| 14 |
A' |
947 |
906 |
2.57 |
|
|
|
| 15 |
A' |
862 |
824 |
3.42 |
|
|
|
| 16 |
A' |
746 |
714 |
2.63 |
|
|
|
| 17 |
A' |
353 |
337 |
0.55 |
|
|
|
| 18 |
A' |
221 |
212 |
2.56 |
|
|
|
| 19 |
A" |
3138 |
3002 |
57.12 |
|
|
|
| 20 |
A" |
3125 |
2990 |
4.02 |
|
|
|
| 21 |
A" |
3095 |
2961 |
1.28 |
|
|
|
| 22 |
A" |
1523 |
1457 |
8.49 |
|
|
|
| 23 |
A" |
1339 |
1281 |
0.01 |
|
|
|
| 24 |
A" |
1259 |
1204 |
0.42 |
|
|
|
| 25 |
A" |
1083 |
1036 |
3.14 |
|
|
|
| 26 |
A" |
879 |
841 |
0.00 |
|
|
|
| 27 |
A" |
758 |
725 |
4.64 |
|
|
|
| 28 |
A" |
234 |
224 |
0.69 |
|
|
|
| 29 |
A" |
187 |
178 |
19.89 |
|
|
|
| 30 |
A" |
108 |
103 |
4.45 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 22893.7 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 21902.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
H |
0.177 |
|
|
0.120 |
| 2 |
H |
0.170 |
|
|
0.133 |
| 3 |
H |
0.170 |
|
|
0.133 |
| 4 |
C |
-0.511 |
|
|
-0.505 |
| 5 |
H |
0.182 |
|
|
0.045 |
| 6 |
H |
0.182 |
|
|
0.045 |
| 7 |
C |
-0.320 |
|
|
0.209 |
| 8 |
H |
0.197 |
|
|
0.147 |
| 9 |
H |
0.197 |
|
|
0.147 |
| 10 |
C |
-0.457 |
|
|
-0.331 |
| 11 |
H |
0.102 |
|
|
0.215 |
| 12 |
S |
-0.089 |
|
|
-0.358 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.388 |
1.202 |
0.000 |
1.836 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
1.423 |
1.241 |
0.000 |
1.888 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-34.681 |
-2.904 |
0.000 |
| y |
-2.904 |
-32.190 |
0.000 |
| z |
0.000 |
0.000 |
-35.124 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.024 |
-2.904 |
0.000 |
| y |
-2.904 |
2.713 |
0.000 |
| z |
0.000 |
0.000 |
-1.689 |
|
| Polar |
|---|
| 3z2-r2 | -3.378 |
|---|
| x2-y2 | -2.492 |
|---|
| xy | -2.904 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
157.965 |
| (<r2>)1/2 |
12.568 |