Jump to
S1C2
Energy calculated at CID/6-31G*
| | hartrees |
|---|
| Energy at 0K | -3107.905293 |
| Energy at 298.15K | |
| HF Energy | -3107.438937 |
| Nuclear repulsion energy | 289.230641 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CID/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
208 |
192 |
7.47 |
|
|
|
| 2 |
A' |
263 |
243 |
0.86 |
|
|
|
| 3 |
A' |
686 |
634 |
52.61 |
|
|
|
| 4 |
A' |
814 |
752 |
28.96 |
|
|
|
| 5 |
A' |
1127 |
1041 |
1.43 |
|
|
|
| 6 |
A' |
1313 |
1212 |
49.02 |
|
|
|
| 7 |
A' |
1423 |
1315 |
2.81 |
|
|
|
| 8 |
A' |
1577 |
1457 |
7.53 |
|
|
|
| 9 |
A' |
1581 |
1461 |
0.82 |
|
|
|
| 10 |
A' |
3230 |
2984 |
4.01 |
|
|
|
| 11 |
A' |
3240 |
2993 |
7.42 |
|
|
|
| 12 |
A" |
119 |
110 |
6.52 |
|
|
|
| 13 |
A" |
801 |
740 |
3.15 |
|
|
|
| 14 |
A" |
1038 |
959 |
0.20 |
|
|
|
| 15 |
A" |
1206 |
1114 |
2.19 |
|
|
|
| 16 |
A" |
1369 |
1265 |
0.08 |
|
|
|
| 17 |
A" |
3292 |
3041 |
0.76 |
|
|
|
| 18 |
A" |
3317 |
3064 |
1.57 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13302.4 cm
-1
Scaled (by 0.9237) Zero Point Vibrational Energy (zpe) 12287.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CID/6-31G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.650 |
0.000 |
| C2 |
1.233 |
-0.225 |
0.000 |
| Br3 |
-1.589 |
-0.487 |
0.000 |
| Cl4 |
2.693 |
0.804 |
0.000 |
| H5 |
-0.057 |
1.268 |
0.888 |
| H6 |
-0.057 |
1.268 |
-0.888 |
| H7 |
1.269 |
-0.851 |
0.885 |
| H8 |
1.269 |
-0.851 |
-0.885 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Br3 |
Cl4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5115 | 1.9537 | 2.6974 | 1.0837 | 1.0837 | 2.1554 | 2.1554 |
C2 | 1.5115 | | 2.8334 | 1.7863 | 2.1637 | 2.1637 | 1.0847 | 1.0847 | Br3 | 1.9537 | 2.8334 | | 4.4721 | 2.4933 | 2.4933 | 3.0130 | 3.0130 | Cl4 | 2.6974 | 1.7863 | 4.4721 | | 2.9270 | 2.9270 | 2.3563 | 2.3563 | H5 | 1.0837 | 2.1637 | 2.4933 | 2.9270 | | 1.7762 | 2.5000 | 3.0648 | H6 | 1.0837 | 2.1637 | 2.4933 | 2.9270 | 1.7762 | | 3.0648 | 2.5000 | H7 | 2.1554 | 1.0847 | 3.0130 | 2.3563 | 2.5000 | 3.0648 | | 1.7695 | H8 | 2.1554 | 1.0847 | 3.0130 | 2.3563 | 3.0648 | 2.5000 | 1.7695 | |
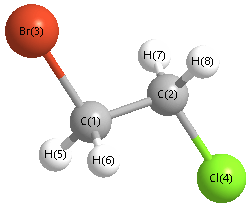 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
Cl4 |
109.475 |
|
C1 |
C2 |
H7 |
111.173 |
| C1 |
C2 |
H8 |
111.173 |
|
C2 |
C1 |
Br3 |
109.040 |
| C2 |
C1 |
Cl4 |
38.635 |
|
C2 |
C1 |
H6 |
111.914 |
| Br3 |
C1 |
H5 |
106.817 |
|
Br3 |
C1 |
H6 |
106.817 |
| Cl4 |
C2 |
H7 |
107.798 |
|
Cl4 |
C2 |
H8 |
107.798 |
| H5 |
C1 |
H6 |
110.074 |
|
H7 |
C2 |
H8 |
109.304 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at CID/6-31G*
| | hartrees |
|---|
| Energy at 0K | -3107.902535 |
| Energy at 298.15K | |
| HF Energy | -3107.435592 |
| Nuclear repulsion energy | 305.850481 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CID/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3297 |
3045 |
1.54 |
|
|
|
| 2 |
A |
3275 |
3025 |
1.88 |
|
|
|
| 3 |
A |
3224 |
2978 |
12.07 |
|
|
|
| 4 |
A |
3204 |
2959 |
9.31 |
|
|
|
| 5 |
A |
1568 |
1448 |
1.52 |
|
|
|
| 6 |
A |
1560 |
1441 |
10.18 |
|
|
|
| 7 |
A |
1433 |
1323 |
27.65 |
|
|
|
| 8 |
A |
1382 |
1276 |
48.76 |
|
|
|
| 9 |
A |
1291 |
1193 |
2.90 |
|
|
|
| 10 |
A |
1218 |
1125 |
1.65 |
|
|
|
| 11 |
A |
1106 |
1022 |
1.36 |
|
|
|
| 12 |
A |
997 |
921 |
8.59 |
|
|
|
| 13 |
A |
928 |
857 |
18.55 |
|
|
|
| 14 |
A |
734 |
678 |
18.09 |
|
|
|
| 15 |
A |
609 |
563 |
13.00 |
|
|
|
| 16 |
A |
408 |
377 |
7.54 |
|
|
|
| 17 |
A |
254 |
235 |
1.19 |
|
|
|
| 18 |
A |
108 |
99 |
0.66 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13296.6 cm
-1
Scaled (by 0.9237) Zero Point Vibrational Energy (zpe) 12282.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CID/6-31G*
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.012 |
1.108 |
-0.383 |
| C2 |
1.284 |
0.895 |
0.403 |
| Br3 |
-1.347 |
-0.225 |
0.036 |
| Cl4 |
2.151 |
-0.578 |
-0.088 |
| H5 |
-0.421 |
2.071 |
-0.130 |
| H6 |
0.184 |
1.052 |
-1.452 |
| H7 |
1.955 |
1.734 |
0.230 |
| H8 |
1.077 |
0.819 |
1.466 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Br3 |
Cl4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5110 | 1.9486 | 2.7403 | 1.0858 | 1.0841 | 2.1312 | 2.1536 |
C2 | 1.5110 | | 2.8829 | 1.7789 | 2.1390 | 2.1625 | 1.0879 | 1.0852 | Br3 | 1.9486 | 2.8829 | | 3.5182 | 2.4811 | 2.4874 | 3.8436 | 3.0015 | Cl4 | 2.7403 | 1.7789 | 3.5182 | | 3.6931 | 2.8960 | 2.3425 | 2.3493 | H5 | 1.0858 | 2.1390 | 2.4811 | 3.6931 | | 1.7758 | 2.4261 | 2.5220 | H6 | 1.0841 | 2.1625 | 2.4874 | 2.8960 | 1.7758 | | 2.5351 | 3.0602 | H7 | 2.1312 | 1.0879 | 3.8436 | 2.3425 | 2.4261 | 2.5351 | | 1.7710 | H8 | 2.1536 | 1.0852 | 3.0015 | 2.3493 | 2.5220 | 3.0602 | 1.7710 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
Cl4 |
112.549 |
|
C1 |
C2 |
H7 |
109.089 |
| C1 |
C2 |
H8 |
111.033 |
|
C2 |
C1 |
Br3 |
112.260 |
| C2 |
C1 |
Cl4 |
36.836 |
|
C2 |
C1 |
H6 |
111.822 |
| Br3 |
C1 |
H5 |
106.185 |
|
Br3 |
C1 |
H6 |
106.709 |
| Cl4 |
C2 |
H7 |
107.122 |
|
Cl4 |
C2 |
H8 |
107.755 |
| H5 |
C1 |
H6 |
109.838 |
|
H7 |
C2 |
H8 |
109.170 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
