Jump to
S1C2
Energy calculated at B3PW91/6-31G*
| | hartrees |
|---|
| Energy at 0K | -3110.432298 |
| Energy at 298.15K | |
| HF Energy | -3110.432298 |
| Nuclear repulsion energy | 288.274297 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
193 |
185 |
8.06 |
|
|
|
| 2 |
A' |
249 |
239 |
0.94 |
|
|
|
| 3 |
A' |
649 |
621 |
58.11 |
|
|
|
| 4 |
A' |
750 |
718 |
33.10 |
|
|
|
| 5 |
A' |
1081 |
1034 |
1.36 |
|
|
|
| 6 |
A' |
1244 |
1190 |
42.02 |
|
|
|
| 7 |
A' |
1340 |
1282 |
3.24 |
|
|
|
| 8 |
A' |
1508 |
1442 |
6.55 |
|
|
|
| 9 |
A' |
1513 |
1447 |
1.25 |
|
|
|
| 10 |
A' |
3128 |
2992 |
3.32 |
|
|
|
| 11 |
A' |
3142 |
3006 |
7.36 |
|
|
|
| 12 |
A" |
113 |
108 |
5.92 |
|
|
|
| 13 |
A" |
771 |
738 |
4.52 |
|
|
|
| 14 |
A" |
983 |
940 |
0.10 |
|
|
|
| 15 |
A" |
1144 |
1094 |
2.48 |
|
|
|
| 16 |
A" |
1307 |
1250 |
0.09 |
|
|
|
| 17 |
A" |
3190 |
3052 |
0.75 |
|
|
|
| 18 |
A" |
3219 |
3079 |
1.65 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 12762.0 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 12209.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3PW91/6-31G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.654 |
0.000 |
| C2 |
1.227 |
-0.229 |
0.000 |
| Br3 |
-1.593 |
-0.486 |
0.000 |
| Cl4 |
2.705 |
0.801 |
0.000 |
| H5 |
-0.059 |
1.275 |
0.894 |
| H6 |
-0.059 |
1.275 |
-0.894 |
| H7 |
1.266 |
-0.859 |
0.890 |
| H8 |
1.266 |
-0.859 |
-0.890 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Br3 |
Cl4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5118 | 1.9588 | 2.7092 | 1.0898 | 1.0898 | 2.1641 | 2.1641 |
C2 | 1.5118 | | 2.8324 | 1.8016 | 2.1713 | 2.1713 | 1.0912 | 1.0912 | Br3 | 1.9588 | 2.8324 | | 4.4871 | 2.5003 | 2.5003 | 3.0181 | 3.0181 | Cl4 | 2.7092 | 1.8016 | 4.4871 | | 2.9434 | 2.9434 | 2.3706 | 2.3706 | H5 | 1.0898 | 2.1713 | 2.5003 | 2.9434 | | 1.7873 | 2.5116 | 3.0806 | H6 | 1.0898 | 2.1713 | 2.5003 | 2.9434 | 1.7873 | | 3.0806 | 2.5116 | H7 | 2.1641 | 1.0912 | 3.0181 | 2.3706 | 2.5116 | 3.0806 | | 1.7805 | H8 | 2.1641 | 1.0912 | 3.0181 | 2.3706 | 3.0806 | 2.5116 | 1.7805 | |
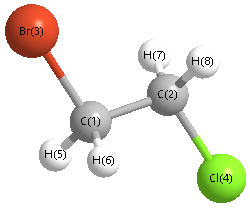 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
Cl4 |
109.391 |
|
C1 |
C2 |
H7 |
111.464 |
| C1 |
C2 |
H8 |
111.464 |
|
C2 |
C1 |
Br3 |
108.710 |
| C2 |
C1 |
Cl4 |
38.849 |
|
C2 |
C1 |
H6 |
112.128 |
| Br3 |
C1 |
H5 |
106.698 |
|
Br3 |
C1 |
H6 |
106.698 |
| Cl4 |
C2 |
H7 |
107.511 |
|
Cl4 |
C2 |
H8 |
107.511 |
| H5 |
C1 |
H6 |
110.170 |
|
H7 |
C2 |
H8 |
109.343 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.344 |
|
|
|
| 2 |
C |
-0.427 |
|
|
|
| 3 |
Br |
-0.118 |
|
|
|
| 4 |
Cl |
-0.060 |
|
|
|
| 5 |
H |
0.235 |
|
|
|
| 6 |
H |
0.235 |
|
|
|
| 7 |
H |
0.239 |
|
|
|
| 8 |
H |
0.239 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.195 |
-0.056 |
0.000 |
0.203 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-49.961 |
-2.274 |
0.000 |
| y |
-2.274 |
-42.762 |
0.000 |
| z |
0.000 |
0.000 |
-43.021 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-7.070 |
-2.274 |
0.000 |
| y |
-2.274 |
3.729 |
0.000 |
| z |
0.000 |
0.000 |
3.340 |
|
| Polar |
|---|
| 3z2-r2 | 6.681 |
|---|
| x2-y2 | -7.199 |
|---|
| xy | -2.274 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.033 |
1.935 |
0.000 |
| y |
1.935 |
6.283 |
0.000 |
| z |
0.000 |
0.000 |
5.130 |
<r2> (average value of r
2) Å
2
| <r2> |
283.741 |
| (<r2>)1/2 |
16.845 |
Jump to
S1C1
Energy calculated at B3PW91/6-31G*
| | hartrees |
|---|
| Energy at 0K | -3110.429428 |
| Energy at 298.15K | |
| HF Energy | -3110.429428 |
| Nuclear repulsion energy | 304.772769 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3196 |
3057 |
1.80 |
|
|
|
| 2 |
A |
3170 |
3033 |
1.79 |
|
|
|
| 3 |
A |
3121 |
2986 |
10.93 |
|
|
|
| 4 |
A |
3099 |
2965 |
10.22 |
|
|
|
| 5 |
A |
1494 |
1430 |
2.09 |
|
|
|
| 6 |
A |
1486 |
1422 |
10.81 |
|
|
|
| 7 |
A |
1356 |
1298 |
26.15 |
|
|
|
| 8 |
A |
1309 |
1252 |
47.88 |
|
|
|
| 9 |
A |
1228 |
1175 |
3.66 |
|
|
|
| 10 |
A |
1161 |
1111 |
1.66 |
|
|
|
| 11 |
A |
1053 |
1008 |
1.86 |
|
|
|
| 12 |
A |
945 |
904 |
9.75 |
|
|
|
| 13 |
A |
877 |
839 |
22.55 |
|
|
|
| 14 |
A |
687 |
658 |
20.21 |
|
|
|
| 15 |
A |
582 |
557 |
12.10 |
|
|
|
| 16 |
A |
389 |
373 |
8.23 |
|
|
|
| 17 |
A |
243 |
232 |
1.30 |
|
|
|
| 18 |
A |
100 |
96 |
0.57 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 12748.1 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 12196.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3PW91/6-31G*
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.007 |
1.114 |
-0.380 |
| C2 |
1.284 |
0.905 |
0.399 |
| Br3 |
-1.350 |
-0.227 |
0.036 |
| Cl4 |
2.160 |
-0.582 |
-0.087 |
| H5 |
-0.431 |
2.080 |
-0.118 |
| H6 |
0.172 |
1.064 |
-1.457 |
| H7 |
1.960 |
1.746 |
0.215 |
| H8 |
1.087 |
0.832 |
1.470 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Br3 |
Cl4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5109 | 1.9522 | 2.7561 | 1.0924 | 1.0903 | 2.1375 | 2.1612 |
C2 | 1.5109 | | 2.8896 | 1.7920 | 2.1421 | 2.1691 | 1.0947 | 1.0917 | Br3 | 1.9522 | 2.8896 | | 3.5296 | 2.4877 | 2.4924 | 3.8573 | 3.0193 | Cl4 | 2.7561 | 1.7920 | 3.5296 | | 3.7141 | 2.9214 | 2.3555 | 2.3607 | H5 | 1.0924 | 2.1421 | 2.4877 | 3.7141 | | 1.7857 | 2.4368 | 2.5262 | H6 | 1.0903 | 2.1691 | 2.4924 | 2.9214 | 1.7857 | | 2.5407 | 3.0754 | H7 | 2.1375 | 1.0947 | 3.8573 | 2.3555 | 2.4368 | 2.5407 | | 1.7814 | H8 | 2.1612 | 1.0917 | 3.0193 | 2.3607 | 2.5262 | 3.0754 | 1.7814 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
Cl4 |
112.840 |
|
C1 |
C2 |
H7 |
109.201 |
| C1 |
C2 |
H8 |
111.263 |
|
C2 |
C1 |
Br3 |
112.484 |
| C2 |
C1 |
Cl4 |
36.815 |
|
C2 |
C1 |
H6 |
111.991 |
| Br3 |
C1 |
H5 |
106.119 |
|
Br3 |
C1 |
H6 |
106.547 |
| Cl4 |
C2 |
H7 |
106.864 |
|
Cl4 |
C2 |
H8 |
107.396 |
| H5 |
C1 |
H6 |
109.792 |
|
H7 |
C2 |
H8 |
109.125 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.349 |
|
|
|
| 2 |
C |
-0.422 |
|
|
|
| 3 |
Br |
-0.105 |
|
|
|
| 4 |
Cl |
-0.048 |
|
|
|
| 5 |
H |
0.224 |
|
|
|
| 6 |
H |
0.236 |
|
|
|
| 7 |
H |
0.227 |
|
|
|
| 8 |
H |
0.239 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.308 |
2.788 |
0.124 |
2.808 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-47.123 |
2.903 |
0.457 |
| y |
2.903 |
-39.905 |
-0.123 |
| z |
0.457 |
-0.123 |
-42.618 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-5.862 |
2.903 |
0.457 |
| y |
2.903 |
4.965 |
-0.123 |
| z |
0.457 |
-0.123 |
0.896 |
|
| Polar |
|---|
| 3z2-r2 | 1.793 |
|---|
| x2-y2 | -7.218 |
|---|
| xy | 2.903 |
|---|
| xz | 0.457 |
|---|
| yz | -0.123 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.076 |
0.193 |
-0.290 |
| y |
0.193 |
7.170 |
0.083 |
| z |
-0.290 |
0.083 |
5.315 |
<r2> (average value of r
2) Å
2
| <r2> |
220.634 |
| (<r2>)1/2 |
14.854 |
