Jump to
S1C2
Energy calculated at B2PLYP/6-31G*
| | hartrees |
|---|
| Energy at 0K | -3109.777705 |
| Energy at 298.15K | |
| HF Energy | -3109.611227 |
| Nuclear repulsion energy | 287.537076 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3166 |
3005 |
7.86 |
29.32 |
0.08 |
0.15 |
| 2 |
A' |
3154 |
2994 |
2.93 |
112.99 |
0.09 |
0.16 |
| 3 |
A' |
1538 |
1460 |
1.68 |
13.72 |
0.75 |
0.86 |
| 4 |
A' |
1534 |
1456 |
4.86 |
4.25 |
0.74 |
0.85 |
| 5 |
A' |
1363 |
1294 |
3.63 |
13.82 |
0.55 |
0.71 |
| 6 |
A' |
1261 |
1197 |
48.29 |
2.86 |
0.44 |
0.61 |
| 7 |
A' |
1086 |
1031 |
1.28 |
8.31 |
0.75 |
0.86 |
| 8 |
A' |
756 |
718 |
30.43 |
47.76 |
0.30 |
0.46 |
| 9 |
A' |
647 |
614 |
56.98 |
12.11 |
0.26 |
0.41 |
| 10 |
A' |
251 |
238 |
0.89 |
5.26 |
0.33 |
0.50 |
| 11 |
A' |
197 |
187 |
7.67 |
0.20 |
0.29 |
0.45 |
| 12 |
A" |
3242 |
3078 |
1.96 |
10.81 |
0.75 |
0.86 |
| 13 |
A" |
3217 |
3053 |
0.76 |
84.52 |
0.75 |
0.86 |
| 14 |
A" |
1326 |
1258 |
0.07 |
9.74 |
0.75 |
0.86 |
| 15 |
A" |
1157 |
1099 |
2.30 |
0.41 |
0.75 |
0.86 |
| 16 |
A" |
998 |
947 |
0.15 |
8.50 |
0.75 |
0.86 |
| 17 |
A" |
782 |
742 |
3.47 |
0.04 |
0.75 |
0.86 |
| 18 |
A" |
114 |
108 |
6.03 |
0.02 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 12894.2 cm
-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 12239.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP/6-31G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.656 |
0.000 |
| C2 |
1.234 |
-0.221 |
0.000 |
| Br3 |
-1.595 |
-0.499 |
0.000 |
| Cl4 |
2.706 |
0.823 |
0.000 |
| H5 |
-0.063 |
1.274 |
0.892 |
| H6 |
-0.063 |
1.274 |
-0.892 |
| H7 |
1.276 |
-0.848 |
0.889 |
| H8 |
1.276 |
-0.848 |
-0.889 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Br3 |
Cl4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5135 | 1.9690 | 2.7109 | 1.0874 | 1.0874 | 2.1628 | 2.1628 |
C2 | 1.5135 | | 2.8424 | 1.8044 | 2.1710 | 2.1710 | 1.0886 | 1.0886 | Br3 | 1.9690 | 2.8424 | | 4.4992 | 2.5070 | 2.5070 | 3.0252 | 3.0252 | Cl4 | 2.7109 | 1.8044 | 4.4992 | | 2.9441 | 2.9441 | 2.3721 | 2.3721 | H5 | 1.0874 | 2.1710 | 2.5070 | 2.9441 | | 1.7848 | 2.5089 | 3.0769 | H6 | 1.0874 | 2.1710 | 2.5070 | 2.9441 | 1.7848 | | 3.0769 | 2.5089 | H7 | 2.1628 | 1.0886 | 3.0252 | 2.3721 | 2.5089 | 3.0769 | | 1.7778 | H8 | 2.1628 | 1.0886 | 3.0252 | 2.3721 | 3.0769 | 2.5089 | 1.7778 | |
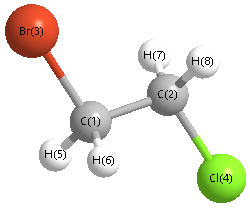 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
Cl4 |
109.268 |
|
C1 |
C2 |
H7 |
111.400 |
| C1 |
C2 |
H8 |
111.400 |
|
C2 |
C1 |
Br3 |
108.709 |
| C2 |
C1 |
Cl4 |
38.928 |
|
C2 |
C1 |
H6 |
112.131 |
| Br3 |
C1 |
H5 |
106.628 |
|
Br3 |
C1 |
H6 |
106.628 |
| Cl4 |
C2 |
H7 |
107.569 |
|
Cl4 |
C2 |
H8 |
107.569 |
| H5 |
C1 |
H6 |
110.296 |
|
H7 |
C2 |
H8 |
109.485 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.296 |
|
|
|
| 2 |
C |
-0.372 |
|
|
|
| 3 |
Br |
-0.133 |
|
|
|
| 4 |
Cl |
-0.072 |
|
|
|
| 5 |
H |
0.217 |
|
|
|
| 6 |
H |
0.217 |
|
|
|
| 7 |
H |
0.220 |
|
|
|
| 8 |
H |
0.220 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.149 |
-0.037 |
0.000 |
0.154 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-50.449 |
-2.341 |
-0.007 |
| y |
-2.341 |
-43.227 |
-0.002 |
| z |
-0.007 |
-0.002 |
-43.428 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-7.122 |
-2.341 |
-0.007 |
| y |
-2.341 |
3.712 |
-0.002 |
| z |
-0.007 |
-0.002 |
3.410 |
|
| Polar |
|---|
| 3z2-r2 | 6.819 |
|---|
| x2-y2 | -7.222 |
|---|
| xy | -2.341 |
|---|
| xz | -0.007 |
|---|
| yz | -0.002 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.831 |
1.985 |
-0.000 |
| y |
1.985 |
6.257 |
-0.000 |
| z |
-0.000 |
-0.000 |
5.074 |
<r2> (average value of r
2) Å
2
| <r2> |
285.385 |
| (<r2>)1/2 |
16.893 |
Jump to
S1C1
Energy calculated at B2PLYP/6-31G*
| | hartrees |
|---|
| Energy at 0K | -3109.774846 |
| Energy at 298.15K | |
| HF Energy | -3109.608126 |
| Nuclear repulsion energy | 304.184809 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3220 |
3056 |
1.97 |
42.25 |
0.75 |
0.86 |
| 2 |
A |
3197 |
3034 |
1.87 |
80.78 |
0.75 |
0.86 |
| 3 |
A |
3145 |
2985 |
11.29 |
131.66 |
0.06 |
0.11 |
| 4 |
A |
3126 |
2967 |
9.87 |
102.64 |
0.22 |
0.36 |
| 5 |
A |
1521 |
1444 |
1.65 |
8.10 |
0.70 |
0.83 |
| 6 |
A |
1513 |
1436 |
8.72 |
16.20 |
0.75 |
0.86 |
| 7 |
A |
1378 |
1308 |
29.01 |
3.90 |
0.75 |
0.86 |
| 8 |
A |
1329 |
1262 |
51.54 |
1.65 |
0.54 |
0.70 |
| 9 |
A |
1241 |
1178 |
4.11 |
17.16 |
0.75 |
0.85 |
| 10 |
A |
1174 |
1114 |
1.64 |
6.09 |
0.73 |
0.84 |
| 11 |
A |
1062 |
1008 |
1.53 |
3.91 |
0.75 |
0.86 |
| 12 |
A |
959 |
910 |
8.61 |
6.38 |
0.45 |
0.62 |
| 13 |
A |
890 |
845 |
19.19 |
1.53 |
0.71 |
0.83 |
| 14 |
A |
691 |
656 |
20.27 |
10.34 |
0.41 |
0.58 |
| 15 |
A |
579 |
549 |
12.56 |
15.82 |
0.21 |
0.34 |
| 16 |
A |
392 |
372 |
8.18 |
2.66 |
0.70 |
0.83 |
| 17 |
A |
245 |
233 |
1.24 |
1.07 |
0.32 |
0.48 |
| 18 |
A |
102 |
97 |
0.57 |
1.30 |
0.73 |
0.84 |
Unscaled Zero Point Vibrational Energy (zpe) 12881.1 cm
-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 12226.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP/6-31G*
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.013 |
1.119 |
-0.383 |
| C2 |
1.288 |
0.906 |
0.403 |
| Br3 |
-1.353 |
-0.227 |
0.036 |
| Cl4 |
2.161 |
-0.584 |
-0.087 |
| H5 |
-0.425 |
2.083 |
-0.124 |
| H6 |
0.182 |
1.064 |
-1.457 |
| H7 |
1.965 |
1.744 |
0.222 |
| H8 |
1.085 |
0.830 |
1.470 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Br3 |
Cl4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5131 | 1.9623 | 2.7570 | 1.0900 | 1.0879 | 2.1375 | 2.1604 |
C2 | 1.5131 | | 2.8968 | 1.7950 | 2.1439 | 2.1694 | 1.0922 | 1.0891 | Br3 | 1.9623 | 2.8968 | | 3.5336 | 2.4939 | 2.4994 | 3.8633 | 3.0192 | Cl4 | 2.7570 | 1.7950 | 3.5336 | | 3.7147 | 2.9167 | 2.3568 | 2.3628 | H5 | 1.0900 | 2.1439 | 2.4939 | 3.7147 | | 1.7841 | 2.4385 | 2.5276 | H6 | 1.0879 | 2.1694 | 2.4994 | 2.9167 | 1.7841 | | 2.5420 | 3.0717 | H7 | 2.1375 | 1.0922 | 3.8633 | 2.3568 | 2.4385 | 2.5420 | | 1.7794 | H8 | 2.1604 | 1.0891 | 3.0192 | 2.3628 | 2.5276 | 3.0717 | 1.7794 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
Cl4 |
112.622 |
|
C1 |
C2 |
H7 |
109.191 |
| C1 |
C2 |
H8 |
111.194 |
|
C2 |
C1 |
Br3 |
112.281 |
| C2 |
C1 |
Cl4 |
36.940 |
|
C2 |
C1 |
H6 |
111.992 |
| Br3 |
C1 |
H5 |
106.027 |
|
Br3 |
C1 |
H6 |
106.502 |
| Cl4 |
C2 |
H7 |
106.896 |
|
Cl4 |
C2 |
H8 |
107.480 |
| H5 |
C1 |
H6 |
110.001 |
|
H7 |
C2 |
H8 |
109.323 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.301 |
|
|
|
| 2 |
C |
-0.369 |
|
|
|
| 3 |
Br |
-0.119 |
|
|
|
| 4 |
Cl |
-0.060 |
|
|
|
| 5 |
H |
0.206 |
|
|
|
| 6 |
H |
0.217 |
|
|
|
| 7 |
H |
0.207 |
|
|
|
| 8 |
H |
0.219 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.341 |
2.791 |
0.118 |
2.814 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-47.581 |
2.899 |
0.451 |
| y |
2.899 |
-40.345 |
-0.114 |
| z |
0.451 |
-0.114 |
-43.033 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-5.892 |
2.899 |
0.451 |
| y |
2.899 |
4.962 |
-0.114 |
| z |
0.451 |
-0.114 |
0.930 |
|
| Polar |
|---|
| 3z2-r2 | 1.860 |
|---|
| x2-y2 | -7.236 |
|---|
| xy | 2.899 |
|---|
| xz | 0.451 |
|---|
| yz | -0.114 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.946 |
0.214 |
-0.319 |
| y |
0.214 |
7.122 |
0.082 |
| z |
-0.319 |
0.082 |
5.253 |
<r2> (average value of r
2) Å
2
| <r2> |
221.490 |
| (<r2>)1/2 |
14.883 |
