Jump to
S1C2
Energy calculated at CID/6-31G*
| | hartrees |
|---|
| Energy at 0K | -997.515730 |
| Energy at 298.15K | -997.520555 |
| HF Energy | -997.030722 |
| Nuclear repulsion energy | 195.230799 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CID/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3227 |
2981 |
0.00 |
|
|
|
| 2 |
Ag |
1583 |
1463 |
0.00 |
|
|
|
| 3 |
Ag |
1450 |
1340 |
0.00 |
|
|
|
| 4 |
Ag |
1128 |
1042 |
0.00 |
|
|
|
| 5 |
Ag |
844 |
779 |
0.00 |
|
|
|
| 6 |
Ag |
320 |
296 |
0.00 |
|
|
|
| 7 |
Au |
3306 |
3053 |
5.10 |
|
|
|
| 8 |
Au |
1230 |
1137 |
1.70 |
|
|
|
| 9 |
Au |
817 |
755 |
2.41 |
|
|
|
| 10 |
Au |
128 |
118 |
7.90 |
|
|
|
| 11 |
Bg |
3286 |
3035 |
0.00 |
|
|
|
| 12 |
Bg |
1376 |
1271 |
0.00 |
|
|
|
| 13 |
Bg |
1088 |
1005 |
0.00 |
|
|
|
| 14 |
Bu |
3232 |
2985 |
17.48 |
|
|
|
| 15 |
Bu |
1583 |
1462 |
8.39 |
|
|
|
| 16 |
Bu |
1353 |
1250 |
40.75 |
|
|
|
| 17 |
Bu |
795 |
734 |
92.34 |
|
|
|
| 18 |
Bu |
230 |
212 |
9.99 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13487.7 cm
-1
Scaled (by 0.9237) Zero Point Vibrational Energy (zpe) 12458.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CID/6-31G*
Point Group is C2h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.757 |
0.000 |
| C2 |
0.000 |
-0.757 |
0.000 |
| Cl3 |
-1.681 |
1.352 |
0.000 |
| Cl4 |
1.681 |
-1.352 |
0.000 |
| H5 |
0.491 |
1.148 |
0.885 |
| H6 |
0.491 |
1.148 |
-0.885 |
| H7 |
-0.491 |
-1.148 |
0.885 |
| H8 |
-0.491 |
-1.148 |
-0.885 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Cl3 |
Cl4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5136 | 1.7832 | 2.6968 | 1.0849 | 1.0849 | 2.1565 | 2.1565 |
C2 | 1.5136 | | 2.6968 | 1.7832 | 2.1565 | 2.1565 | 1.0849 | 1.0849 | Cl3 | 1.7832 | 2.6968 | | 4.3144 | 2.3545 | 2.3545 | 2.9061 | 2.9061 | Cl4 | 2.6968 | 1.7832 | 4.3144 | | 2.9061 | 2.9061 | 2.3545 | 2.3545 | H5 | 1.0849 | 2.1565 | 2.3545 | 2.9061 | | 1.7694 | 2.4967 | 3.0601 | H6 | 1.0849 | 2.1565 | 2.3545 | 2.9061 | 1.7694 | | 3.0601 | 2.4967 | H7 | 2.1565 | 1.0849 | 2.9061 | 2.3545 | 2.4967 | 3.0601 | | 1.7694 | H8 | 2.1565 | 1.0849 | 2.9061 | 2.3545 | 3.0601 | 2.4967 | 1.7694 | |
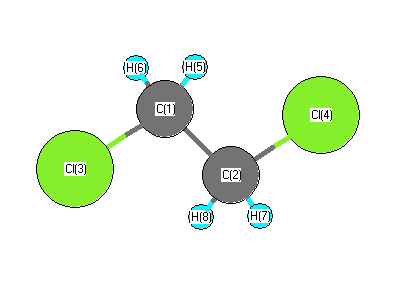 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
Cl4 |
109.503 |
|
C1 |
C2 |
H7 |
111.111 |
| C1 |
C2 |
H8 |
111.111 |
|
C2 |
C1 |
Cl3 |
109.503 |
| C2 |
C1 |
H5 |
111.111 |
|
C2 |
C1 |
H6 |
111.111 |
| Cl3 |
C1 |
H5 |
107.865 |
|
Cl3 |
C1 |
H6 |
107.865 |
| Cl4 |
C2 |
H7 |
107.865 |
|
Cl4 |
C2 |
H8 |
107.865 |
| H5 |
C1 |
H6 |
109.275 |
|
H7 |
C2 |
H8 |
109.275 |
Electronic energy levels
Electronic state
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at CID/6-31G*
| | hartrees |
|---|
| Energy at 0K | -997.513134 |
| Energy at 298.15K | -997.518094 |
| HF Energy | -997.027687 |
| Nuclear repulsion energy | 202.629264 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CID/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3274 |
3024 |
1.06 |
|
|
|
| 2 |
A |
3216 |
2971 |
22.36 |
|
|
|
| 3 |
A |
1568 |
1449 |
0.12 |
|
|
|
| 4 |
A |
1445 |
1335 |
24.69 |
|
|
|
| 5 |
A |
1316 |
1216 |
0.76 |
|
|
|
| 6 |
A |
1121 |
1035 |
0.52 |
|
|
|
| 7 |
A |
1026 |
947 |
12.21 |
|
|
|
| 8 |
A |
720 |
665 |
19.68 |
|
|
|
| 9 |
A |
275 |
254 |
0.82 |
|
|
|
| 10 |
A |
123 |
114 |
1.13 |
|
|
|
| 11 |
B |
3284 |
3034 |
5.44 |
|
|
|
| 12 |
B |
3207 |
2962 |
3.84 |
|
|
|
| 13 |
B |
1564 |
1445 |
11.96 |
|
|
|
| 14 |
B |
1421 |
1313 |
40.19 |
|
|
|
| 15 |
B |
1245 |
1150 |
1.15 |
|
|
|
| 16 |
B |
962 |
889 |
18.45 |
|
|
|
| 17 |
B |
751 |
693 |
23.87 |
|
|
|
| 18 |
B |
434 |
401 |
7.67 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13475.5 cm
-1
Scaled (by 0.9237) Zero Point Vibrational Energy (zpe) 12447.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CID/6-31G*
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.295 |
0.696 |
0.885 |
| C2 |
-0.295 |
-0.696 |
0.885 |
| Cl3 |
-0.295 |
1.692 |
-0.467 |
| Cl4 |
0.295 |
-1.692 |
-0.467 |
| H5 |
0.009 |
1.201 |
1.804 |
| H6 |
1.378 |
0.660 |
0.821 |
| H7 |
-0.009 |
-1.201 |
1.804 |
| H8 |
-1.378 |
-0.660 |
0.821 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Cl3 |
Cl4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5122 | 1.7790 | 2.7436 | 1.0871 | 1.0853 | 2.1296 | 2.1546 |
C2 | 1.5122 | | 2.7436 | 1.7790 | 2.1296 | 2.1546 | 1.0871 | 1.0853 | Cl3 | 1.7790 | 2.7436 | | 3.4342 | 2.3430 | 2.3493 | 3.6882 | 2.8918 | Cl4 | 2.7436 | 1.7790 | 3.4342 | | 3.6882 | 2.8918 | 2.3430 | 2.3493 | H5 | 1.0871 | 2.1296 | 2.3430 | 3.6882 | | 1.7699 | 2.4012 | 2.5204 | H6 | 1.0853 | 2.1546 | 2.3493 | 2.8918 | 1.7699 | | 2.5204 | 3.0558 | H7 | 2.1296 | 1.0871 | 3.6882 | 2.3430 | 2.4012 | 2.5204 | | 1.7699 | H8 | 2.1546 | 1.0853 | 2.8918 | 2.3493 | 2.5204 | 3.0558 | 1.7699 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
Cl4 |
112.697 |
|
C1 |
C2 |
H7 |
108.935 |
| C1 |
C2 |
H8 |
111.033 |
|
C2 |
C1 |
Cl3 |
112.697 |
| C2 |
C1 |
H5 |
108.935 |
|
C2 |
C1 |
H6 |
111.033 |
| Cl3 |
C1 |
H5 |
107.188 |
|
Cl3 |
C1 |
H6 |
107.742 |
| Cl4 |
C2 |
H7 |
107.188 |
|
Cl4 |
C2 |
H8 |
107.742 |
| H5 |
C1 |
H6 |
109.123 |
|
H7 |
C2 |
H8 |
109.123 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
