Jump to
S1C2
Energy calculated at B3PW91/6-31G*
| | hartrees |
|---|
| Energy at 0K | -998.885784 |
| Energy at 298.15K | -998.890463 |
| Nuclear repulsion energy | 194.059192 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3124 |
2989 |
0.00 |
|
|
|
| 2 |
Ag |
1514 |
1448 |
0.00 |
|
|
|
| 3 |
Ag |
1365 |
1306 |
0.00 |
|
|
|
| 4 |
Ag |
1080 |
1033 |
0.00 |
|
|
|
| 5 |
Ag |
778 |
744 |
0.00 |
|
|
|
| 6 |
Ag |
302 |
289 |
0.00 |
|
|
|
| 7 |
Au |
3205 |
3066 |
4.69 |
|
|
|
| 8 |
Au |
1168 |
1118 |
2.02 |
|
|
|
| 9 |
Au |
786 |
752 |
3.92 |
|
|
|
| 10 |
Au |
121 |
116 |
7.47 |
|
|
|
| 11 |
Bg |
3183 |
3046 |
0.00 |
|
|
|
| 12 |
Bg |
1313 |
1256 |
0.00 |
|
|
|
| 13 |
Bg |
1029 |
985 |
0.00 |
|
|
|
| 14 |
Bu |
3132 |
2996 |
16.29 |
|
|
|
| 15 |
Bu |
1513 |
1448 |
8.03 |
|
|
|
| 16 |
Bu |
1282 |
1226 |
36.28 |
|
|
|
| 17 |
Bu |
736 |
704 |
102.97 |
|
|
|
| 18 |
Bu |
215 |
205 |
10.88 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 12922.8 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 12363.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3PW91/6-31G*
Point Group is C2h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.758 |
0.000 |
| C2 |
0.000 |
-0.758 |
0.000 |
| Cl3 |
-1.697 |
1.353 |
0.000 |
| Cl4 |
1.697 |
-1.353 |
0.000 |
| H5 |
0.489 |
1.156 |
0.891 |
| H6 |
0.489 |
1.156 |
-0.891 |
| H7 |
-0.489 |
-1.156 |
0.891 |
| H8 |
-0.489 |
-1.156 |
-0.891 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Cl3 |
Cl4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5150 | 1.7984 | 2.7080 | 1.0914 | 1.0914 | 2.1663 | 2.1663 |
C2 | 1.5150 | | 2.7080 | 1.7984 | 2.1663 | 2.1663 | 1.0914 | 1.0914 | Cl3 | 1.7984 | 2.7080 | | 4.3405 | 2.3688 | 2.3688 | 2.9232 | 2.9232 | Cl4 | 2.7080 | 1.7984 | 4.3405 | | 2.9232 | 2.9232 | 2.3688 | 2.3688 | H5 | 1.0914 | 2.1663 | 2.3688 | 2.9232 | | 1.7813 | 2.5099 | 3.0778 | H6 | 1.0914 | 2.1663 | 2.3688 | 2.9232 | 1.7813 | | 3.0778 | 2.5099 | H7 | 2.1663 | 1.0914 | 2.9232 | 2.3688 | 2.5099 | 3.0778 | | 1.7813 | H8 | 2.1663 | 1.0914 | 2.9232 | 2.3688 | 3.0778 | 2.5099 | 1.7813 | |
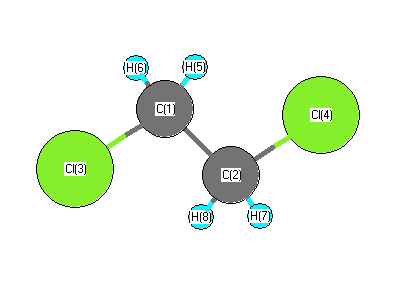 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
Cl4 |
109.331 |
|
C1 |
C2 |
H7 |
111.400 |
| C1 |
C2 |
H8 |
111.400 |
|
C2 |
C1 |
Cl3 |
109.331 |
| C2 |
C1 |
H5 |
111.400 |
|
C2 |
C1 |
H6 |
111.400 |
| Cl3 |
C1 |
H5 |
107.584 |
|
Cl3 |
C1 |
H6 |
107.584 |
| Cl4 |
C2 |
H7 |
107.584 |
|
Cl4 |
C2 |
H8 |
107.584 |
| H5 |
C1 |
H6 |
109.393 |
|
H7 |
C2 |
H8 |
109.393 |
Electronic energy levels
Electronic state
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.421 |
|
|
-0.152 |
| 2 |
C |
-0.421 |
|
|
-0.148 |
| 3 |
Cl |
-0.058 |
|
|
-0.157 |
| 4 |
Cl |
-0.058 |
|
|
-0.157 |
| 5 |
H |
0.239 |
|
|
0.154 |
| 6 |
H |
0.239 |
|
|
0.154 |
| 7 |
H |
0.239 |
|
|
0.153 |
| 8 |
H |
0.239 |
|
|
0.153 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
0.002 |
-0.001 |
0.000 |
0.002 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-42.230 |
4.435 |
0.000 |
| y |
4.435 |
-39.794 |
0.000 |
| z |
0.000 |
0.000 |
-37.278 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-3.693 |
4.435 |
0.000 |
| y |
4.435 |
-0.041 |
0.000 |
| z |
0.000 |
0.000 |
3.734 |
|
| Polar |
|---|
| 3z2-r2 | 7.468 |
|---|
| x2-y2 | -2.435 |
|---|
| xy | 4.435 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
201.335 |
| (<r2>)1/2 |
14.189 |
Jump to
S1C1
Energy calculated at B3PW91/6-31G*
| | hartrees |
|---|
| Energy at 0K | -998.883219 |
| Energy at 298.15K | -998.888033 |
| HF Energy | -998.883219 |
| Nuclear repulsion energy | 201.290309 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3169 |
3032 |
0.64 |
|
|
|
| 2 |
A |
3110 |
2975 |
22.05 |
|
|
|
| 3 |
A |
1494 |
1429 |
0.47 |
|
|
|
| 4 |
A |
1367 |
1307 |
23.56 |
|
|
|
| 5 |
A |
1253 |
1198 |
1.11 |
|
|
|
| 6 |
A |
1065 |
1019 |
0.82 |
|
|
|
| 7 |
A |
969 |
927 |
14.34 |
|
|
|
| 8 |
A |
675 |
646 |
19.81 |
|
|
|
| 9 |
A |
261 |
250 |
0.98 |
|
|
|
| 10 |
A |
115 |
110 |
1.01 |
|
|
|
| 11 |
B |
3182 |
3044 |
5.32 |
|
|
|
| 12 |
B |
3102 |
2967 |
3.37 |
|
|
|
| 13 |
B |
1490 |
1425 |
13.19 |
|
|
|
| 14 |
B |
1342 |
1284 |
41.68 |
|
|
|
| 15 |
B |
1185 |
1134 |
1.30 |
|
|
|
| 16 |
B |
908 |
869 |
22.63 |
|
|
|
| 17 |
B |
700 |
669 |
27.20 |
|
|
|
| 18 |
B |
412 |
395 |
8.69 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 12898.6 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 12340.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3PW91/6-31G*
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.294 |
0.697 |
0.890 |
| C2 |
-0.294 |
-0.697 |
0.890 |
| Cl3 |
-0.294 |
1.706 |
-0.469 |
| Cl4 |
0.294 |
-1.706 |
-0.469 |
| H5 |
0.003 |
1.205 |
1.814 |
| H6 |
1.384 |
0.666 |
0.826 |
| H7 |
-0.003 |
-1.205 |
1.814 |
| H8 |
-1.384 |
-0.666 |
0.826 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Cl3 |
Cl4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5128 | 1.7924 | 2.7608 | 1.0941 | 1.0918 | 2.1353 | 2.1631 |
C2 | 1.5128 | | 2.7608 | 1.7924 | 2.1353 | 2.1631 | 1.0941 | 1.0918 | Cl3 | 1.7924 | 2.7608 | | 3.4632 | 2.3564 | 2.3616 | 3.7113 | 2.9148 | Cl4 | 2.7608 | 1.7924 | 3.4632 | | 3.7113 | 2.9148 | 2.3564 | 2.3616 | H5 | 1.0941 | 2.1353 | 2.3564 | 3.7113 | | 1.7807 | 2.4100 | 2.5301 | H6 | 1.0918 | 2.1631 | 2.3616 | 2.9148 | 1.7807 | | 2.5301 | 3.0719 | H7 | 2.1353 | 1.0941 | 3.7113 | 2.3564 | 2.4100 | 2.5301 | | 1.7807 | H8 | 2.1631 | 1.0918 | 2.9148 | 2.3616 | 2.5301 | 3.0719 | 1.7807 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
Cl4 |
113.019 |
|
C1 |
C2 |
H7 |
108.927 |
| C1 |
C2 |
H8 |
111.267 |
|
C2 |
C1 |
Cl3 |
113.019 |
| C2 |
C1 |
H5 |
108.927 |
|
C2 |
C1 |
H6 |
111.267 |
| Cl3 |
C1 |
H5 |
106.941 |
|
Cl3 |
C1 |
H6 |
107.432 |
| Cl4 |
C2 |
H7 |
106.941 |
|
Cl4 |
C2 |
H8 |
107.432 |
| H5 |
C1 |
H6 |
109.106 |
|
H7 |
C2 |
H8 |
109.106 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.420 |
|
|
|
| 2 |
C |
-0.420 |
|
|
|
| 3 |
Cl |
-0.048 |
|
|
|
| 4 |
Cl |
-0.048 |
|
|
|
| 5 |
H |
0.228 |
|
|
|
| 6 |
H |
0.240 |
|
|
|
| 7 |
H |
0.228 |
|
|
|
| 8 |
H |
0.240 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
2.924 |
2.924 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-37.135 |
1.250 |
0.000 |
| y |
1.250 |
-42.159 |
0.000 |
| z |
0.000 |
0.000 |
-35.016 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.452 |
1.250 |
0.000 |
| y |
1.250 |
-6.083 |
0.000 |
| z |
0.000 |
0.000 |
4.631 |
|
| Polar |
|---|
| 3z2-r2 | 9.262 |
|---|
| x2-y2 | 5.023 |
|---|
| xy | 1.250 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.678 |
-0.596 |
0.000 |
| y |
-0.596 |
6.703 |
0.000 |
| z |
0.000 |
0.000 |
6.282 |
<r2> (average value of r
2) Å
2
| <r2> |
165.166 |
| (<r2>)1/2 |
12.852 |
