Jump to
S1C2
Energy calculated at B2PLYP/6-31G*
| | hartrees |
|---|
| Energy at 0K | -998.588320 |
| Energy at 298.15K | -998.593029 |
| HF Energy | -998.416899 |
| Nuclear repulsion energy | 193.883439 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3152 |
2992 |
0.00 |
|
|
|
| 2 |
Ag |
1539 |
1461 |
0.00 |
|
|
|
| 3 |
Ag |
1389 |
1318 |
0.00 |
|
|
|
| 4 |
Ag |
1086 |
1031 |
0.00 |
|
|
|
| 5 |
Ag |
785 |
745 |
0.00 |
|
|
|
| 6 |
Ag |
305 |
289 |
0.00 |
|
|
|
| 7 |
Au |
3232 |
3068 |
5.29 |
|
|
|
| 8 |
Au |
1182 |
1122 |
1.86 |
|
|
|
| 9 |
Au |
797 |
757 |
2.82 |
|
|
|
| 10 |
Au |
123 |
117 |
7.51 |
|
|
|
| 11 |
Bg |
3211 |
3048 |
0.00 |
|
|
|
| 12 |
Bg |
1332 |
1264 |
0.00 |
|
|
|
| 13 |
Bg |
1046 |
993 |
0.00 |
|
|
|
| 14 |
Bu |
3159 |
2999 |
16.41 |
|
|
|
| 15 |
Bu |
1540 |
1462 |
6.74 |
|
|
|
| 16 |
Bu |
1302 |
1236 |
41.71 |
|
|
|
| 17 |
Bu |
739 |
701 |
98.57 |
|
|
|
| 18 |
Bu |
219 |
208 |
10.33 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13067.9 cm
-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 12404.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP/6-31G*
Point Group is C2h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.476 |
0.590 |
0.000 |
| C2 |
-0.476 |
-0.590 |
0.000 |
| Cl3 |
-0.476 |
2.120 |
0.000 |
| Cl4 |
0.476 |
-2.120 |
0.000 |
| H5 |
1.104 |
0.593 |
0.889 |
| H6 |
1.104 |
0.593 |
-0.889 |
| H7 |
-1.104 |
-0.593 |
0.889 |
| H8 |
-1.104 |
-0.593 |
-0.889 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Cl3 |
Cl4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5164 | 1.8016 | 2.7107 | 1.0886 | 1.0886 | 2.1646 | 2.1646 |
C2 | 1.5164 | | 2.7107 | 1.8016 | 2.1646 | 2.1646 | 1.0886 | 1.0886 | Cl3 | 1.8016 | 2.7107 | | 4.3459 | 2.3704 | 2.3704 | 2.9233 | 2.9233 | Cl4 | 2.7107 | 1.8016 | 4.3459 | | 2.9233 | 2.9233 | 2.3704 | 2.3704 | H5 | 1.0886 | 2.1646 | 2.3704 | 2.9233 | | 1.7784 | 2.5058 | 3.0727 | H6 | 1.0886 | 2.1646 | 2.3704 | 2.9233 | 1.7784 | | 3.0727 | 2.5058 | H7 | 2.1646 | 1.0886 | 2.9233 | 2.3704 | 2.5058 | 3.0727 | | 1.7784 | H8 | 2.1646 | 1.0886 | 2.9233 | 2.3704 | 3.0727 | 2.5058 | 1.7784 | |
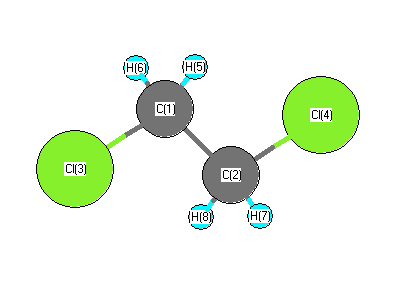 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
Cl4 |
109.261 |
|
C1 |
C2 |
H7 |
111.327 |
| C1 |
C2 |
H8 |
111.327 |
|
C2 |
C1 |
Cl3 |
109.261 |
| C2 |
C1 |
H5 |
111.327 |
|
C2 |
C1 |
H6 |
111.327 |
| Cl3 |
C1 |
H5 |
107.630 |
|
Cl3 |
C1 |
H6 |
107.630 |
| Cl4 |
C2 |
H7 |
107.630 |
|
Cl4 |
C2 |
H8 |
107.630 |
| H5 |
C1 |
H6 |
109.526 |
|
H7 |
C2 |
H8 |
109.526 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.371 |
|
|
|
| 2 |
C |
-0.371 |
|
|
|
| 3 |
Cl |
-0.079 |
|
|
|
| 4 |
Cl |
-0.079 |
|
|
|
| 5 |
H |
0.225 |
|
|
|
| 6 |
H |
0.225 |
|
|
|
| 7 |
H |
0.225 |
|
|
|
| 8 |
H |
0.225 |
|
|
|
Electric dipole moments
Electric Quadrupole moment
Quadrupole components in D Å
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.184 |
-1.490 |
0.000 |
| y |
-1.490 |
8.477 |
0.000 |
| z |
0.000 |
0.000 |
4.313 |
<r2> (average value of r
2) Å
2
| <r2> |
202.031 |
| (<r2>)1/2 |
14.214 |
Jump to
S1C1
Energy calculated at B2PLYP/6-31G*
| | hartrees |
|---|
| Energy at 0K | -998.585703 |
| Energy at 298.15K | -998.590549 |
| HF Energy | -998.414109 |
| Nuclear repulsion energy | 201.253436 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3196 |
3033 |
0.81 |
|
|
|
| 2 |
A |
3137 |
2977 |
22.36 |
|
|
|
| 3 |
A |
1521 |
1444 |
0.08 |
|
|
|
| 4 |
A |
1389 |
1318 |
25.19 |
|
|
|
| 5 |
A |
1267 |
1203 |
1.26 |
|
|
|
| 6 |
A |
1076 |
1021 |
0.52 |
|
|
|
| 7 |
A |
983 |
933 |
12.70 |
|
|
|
| 8 |
A |
678 |
644 |
20.36 |
|
|
|
| 9 |
A |
265 |
251 |
0.90 |
|
|
|
| 10 |
A |
117 |
111 |
1.01 |
|
|
|
| 11 |
B |
3207 |
3045 |
5.77 |
|
|
|
| 12 |
B |
3129 |
2970 |
3.39 |
|
|
|
| 13 |
B |
1517 |
1440 |
10.97 |
|
|
|
| 14 |
B |
1367 |
1298 |
45.09 |
|
|
|
| 15 |
B |
1200 |
1139 |
1.24 |
|
|
|
| 16 |
B |
923 |
876 |
18.81 |
|
|
|
| 17 |
B |
705 |
669 |
26.58 |
|
|
|
| 18 |
B |
417 |
396 |
8.45 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13046.2 cm
-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 12383.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP/6-31G*
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.296 |
0.697 |
0.893 |
| C2 |
-0.296 |
-0.697 |
0.893 |
| Cl3 |
-0.296 |
1.704 |
-0.471 |
| Cl4 |
0.296 |
-1.704 |
-0.471 |
| H5 |
0.006 |
1.206 |
1.814 |
| H6 |
1.383 |
0.663 |
0.827 |
| H7 |
-0.006 |
-1.206 |
1.814 |
| H8 |
-1.383 |
-0.663 |
0.827 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Cl3 |
Cl4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5149 | 1.7954 | 2.7611 | 1.0915 | 1.0892 | 2.1357 | 2.1621 |
C2 | 1.5149 | | 2.7611 | 1.7954 | 2.1357 | 2.1621 | 1.0915 | 1.0892 | Cl3 | 1.7954 | 2.7611 | | 3.4589 | 2.3576 | 2.3634 | 3.7108 | 2.9102 | Cl4 | 2.7611 | 1.7954 | 3.4589 | | 3.7108 | 2.9102 | 2.3576 | 2.3634 | H5 | 1.0915 | 2.1357 | 2.3576 | 3.7108 | | 1.7786 | 2.4118 | 2.5293 | H6 | 1.0892 | 2.1621 | 2.3634 | 2.9102 | 1.7786 | | 2.5293 | 3.0675 | H7 | 2.1357 | 1.0915 | 3.7108 | 2.3576 | 2.4118 | 2.5293 | | 1.7786 | H8 | 2.1621 | 1.0892 | 2.9102 | 2.3634 | 2.5293 | 3.0675 | 1.7786 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
Cl4 |
112.776 |
|
C1 |
C2 |
H7 |
108.973 |
| C1 |
C2 |
H8 |
111.200 |
|
C2 |
C1 |
Cl3 |
112.776 |
| C2 |
C1 |
H5 |
108.973 |
|
C2 |
C1 |
H6 |
111.200 |
| Cl3 |
C1 |
H5 |
106.968 |
|
Cl3 |
C1 |
H6 |
107.496 |
| Cl4 |
C2 |
H7 |
106.968 |
|
Cl4 |
C2 |
H8 |
107.496 |
| H5 |
C1 |
H6 |
109.295 |
|
H7 |
C2 |
H8 |
109.295 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.368 |
|
|
|
| 2 |
C |
-0.368 |
|
|
|
| 3 |
Cl |
-0.060 |
|
|
|
| 4 |
Cl |
-0.060 |
|
|
|
| 5 |
H |
0.208 |
|
|
|
| 6 |
H |
0.220 |
|
|
|
| 7 |
H |
0.208 |
|
|
|
| 8 |
H |
0.220 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
2.906 |
2.906 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-37.492 |
1.237 |
0.000 |
| y |
1.237 |
-42.525 |
0.000 |
| z |
0.000 |
0.000 |
-35.397 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.469 |
1.237 |
0.000 |
| y |
1.237 |
-6.080 |
0.000 |
| z |
0.000 |
0.000 |
4.611 |
|
| Polar |
|---|
| 3z2-r2 | 9.222 |
|---|
| x2-y2 | 5.033 |
|---|
| xy | 1.237 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.637 |
-0.611 |
0.000 |
| y |
-0.611 |
6.571 |
0.000 |
| z |
0.000 |
0.000 |
6.242 |
<r2> (average value of r
2) Å
2
| <r2> |
165.265 |
| (<r2>)1/2 |
12.856 |
