Jump to
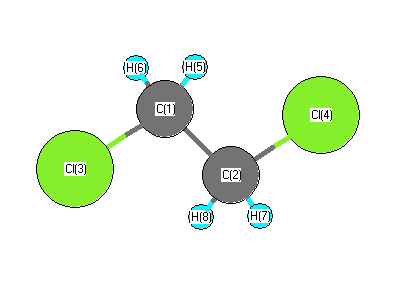
S1C2
Energy calculated at PBEPBEultrafine/6-31G*
| | hartrees |
|---|
| Energy at 0K | -998.500540 |
| Energy at 298.15K | -998.505131 |
| HF Energy | -998.500540 |
| Nuclear repulsion energy | 192.906942 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBEPBEultrafine/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3044 |
2994 |
0.00 |
|
|
|
| 2 |
Ag |
1468 |
1443 |
0.00 |
|
|
|
| 3 |
Ag |
1313 |
1292 |
0.00 |
|
|
|
| 4 |
Ag |
1051 |
1034 |
0.00 |
|
|
|
| 5 |
Ag |
742 |
730 |
0.00 |
|
|
|
| 6 |
Ag |
293 |
288 |
0.00 |
|
|
|
| 7 |
Au |
3124 |
3072 |
5.24 |
|
|
|
| 8 |
Au |
1124 |
1105 |
2.24 |
|
|
|
| 9 |
Au |
766 |
753 |
4.12 |
|
|
|
| 10 |
Au |
119 |
117 |
6.96 |
|
|
|
| 11 |
Bg |
3101 |
3050 |
0.00 |
|
|
|
| 12 |
Bg |
1270 |
1249 |
0.00 |
|
|
|
| 13 |
Bg |
995 |
978 |
0.00 |
|
|
|
| 14 |
Bu |
3052 |
3002 |
16.08 |
|
|
|
| 15 |
Bu |
1469 |
1444 |
6.67 |
|
|
|
| 16 |
Bu |
1235 |
1214 |
34.62 |
|
|
|
| 17 |
Bu |
705 |
693 |
104.27 |
|
|
|
| 18 |
Bu |
208 |
204 |
11.01 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 12538.5 cm
-1
Scaled (by 0.9835) Zero Point Vibrational Energy (zpe) 12331.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBEPBEultrafine/6-31G*
Point Group is C2h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.479 |
0.591 |
0.000 |
| C2 |
-0.479 |
-0.591 |
0.000 |
| Cl3 |
-0.479 |
2.129 |
0.000 |
| Cl4 |
0.479 |
-2.129 |
0.000 |
| H5 |
1.113 |
0.595 |
0.898 |
| H6 |
1.113 |
0.595 |
-0.898 |
| H7 |
-1.113 |
-0.595 |
0.898 |
| H8 |
-1.113 |
-0.595 |
-0.898 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Cl3 |
Cl4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5206 | 1.8118 | 2.7196 | 1.0996 | 1.0996 | 2.1786 | 2.1786 |
C2 | 1.5206 | | 2.7196 | 1.8118 | 2.1786 | 2.1786 | 1.0996 | 1.0996 | Cl3 | 1.8118 | 2.7196 | | 4.3641 | 2.3863 | 2.3863 | 2.9373 | 2.9373 | Cl4 | 2.7196 | 1.8118 | 4.3641 | | 2.9373 | 2.9373 | 2.3863 | 2.3863 | H5 | 1.0996 | 2.1786 | 2.3863 | 2.9373 | | 1.7959 | 2.5246 | 3.0982 | H6 | 1.0996 | 2.1786 | 2.3863 | 2.9373 | 1.7959 | | 3.0982 | 2.5246 | H7 | 2.1786 | 1.0996 | 2.9373 | 2.3863 | 2.5246 | 3.0982 | | 1.7959 | H8 | 2.1786 | 1.0996 | 2.9373 | 2.3863 | 3.0982 | 2.5246 | 1.7959 | |
 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
Cl4 |
109.081 |
|
C1 |
C2 |
H7 |
111.486 |
| C1 |
C2 |
H8 |
111.486 |
|
C2 |
C1 |
Cl3 |
109.081 |
| C2 |
C1 |
H5 |
111.486 |
|
C2 |
C1 |
H6 |
111.486 |
| Cl3 |
C1 |
H5 |
107.569 |
|
Cl3 |
C1 |
H6 |
107.569 |
| Cl4 |
C2 |
H7 |
107.569 |
|
Cl4 |
C2 |
H8 |
107.569 |
| H5 |
C1 |
H6 |
109.491 |
|
H7 |
C2 |
H8 |
109.491 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.402 |
|
|
|
| 2 |
C |
-0.402 |
|
|
|
| 3 |
Cl |
-0.053 |
|
|
|
| 4 |
Cl |
-0.053 |
|
|
|
| 5 |
H |
0.228 |
|
|
|
| 6 |
H |
0.228 |
|
|
|
| 7 |
H |
0.228 |
|
|
|
| 8 |
H |
0.228 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-37.051 |
2.064 |
0.000 |
| y |
2.064 |
-44.907 |
0.000 |
| z |
0.000 |
0.000 |
-37.404 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
4.105 |
2.064 |
0.000 |
| y |
2.064 |
-7.680 |
0.000 |
| z |
0.000 |
0.000 |
3.575 |
|
| Polar |
|---|
| 3z2-r2 | 7.150 |
|---|
| x2-y2 | 7.856 |
|---|
| xy | 2.064 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.343 |
-1.499 |
0.000 |
| y |
-1.499 |
9.064 |
0.000 |
| z |
0.000 |
0.000 |
4.434 |
<r2> (average value of r
2) Å
2
| <r2> |
203.273 |
| (<r2>)1/2 |
14.257 |
Jump to
S1C1
Energy calculated at PBEPBEultrafine/6-31G*
| | hartrees |
|---|
| Energy at 0K | -998.498097 |
| Energy at 298.15K | -998.502831 |
| HF Energy | -998.498097 |
| Nuclear repulsion energy | 200.119979 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBEPBEultrafine/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3084 |
3034 |
0.61 |
|
|
|
| 2 |
A |
3027 |
2977 |
21.49 |
|
|
|
| 3 |
A |
1447 |
1423 |
0.47 |
|
|
|
| 4 |
A |
1320 |
1298 |
23.16 |
|
|
|
| 5 |
A |
1210 |
1190 |
1.58 |
|
|
|
| 6 |
A |
1032 |
1015 |
0.94 |
|
|
|
| 7 |
A |
938 |
922 |
14.10 |
|
|
|
| 8 |
A |
651 |
640 |
18.97 |
|
|
|
| 9 |
A |
257 |
253 |
1.04 |
|
|
|
| 10 |
A |
113 |
112 |
0.87 |
|
|
|
| 11 |
B |
3098 |
3047 |
5.96 |
|
|
|
| 12 |
B |
3019 |
2969 |
3.18 |
|
|
|
| 13 |
B |
1443 |
1419 |
12.66 |
|
|
|
| 14 |
B |
1295 |
1273 |
42.49 |
|
|
|
| 15 |
B |
1146 |
1127 |
1.39 |
|
|
|
| 16 |
B |
877 |
863 |
23.16 |
|
|
|
| 17 |
B |
673 |
662 |
27.45 |
|
|
|
| 18 |
B |
402 |
395 |
8.90 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 12515.6 cm
-1
Scaled (by 0.9835) Zero Point Vibrational Energy (zpe) 12309.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBEPBEultrafine/6-31G*
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.295 |
0.700 |
0.897 |
| C2 |
-0.295 |
-0.700 |
0.897 |
| Cl3 |
-0.295 |
1.715 |
-0.473 |
| Cl4 |
0.295 |
-1.715 |
-0.473 |
| H5 |
-0.003 |
1.212 |
1.827 |
| H6 |
1.393 |
0.671 |
0.836 |
| H7 |
0.003 |
-1.212 |
1.827 |
| H8 |
-1.393 |
-0.671 |
0.836 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Cl3 |
Cl4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5182 | 1.8044 | 2.7760 | 1.1028 | 1.1002 | 2.1457 | 2.1746 |
C2 | 1.5182 | | 2.7760 | 1.8044 | 2.1457 | 2.1746 | 1.1028 | 1.1002 | Cl3 | 1.8044 | 2.7760 | | 3.4794 | 2.3732 | 2.3774 | 3.7344 | 2.9342 | Cl4 | 2.7760 | 1.8044 | 3.4794 | | 3.7344 | 2.9342 | 2.3732 | 2.3774 | H5 | 1.1028 | 2.1457 | 2.3732 | 3.7344 | | 1.7952 | 2.4236 | 2.5415 | H6 | 1.1002 | 2.1746 | 2.3774 | 2.9342 | 1.7952 | | 2.5415 | 3.0917 | H7 | 2.1457 | 1.1028 | 3.7344 | 2.3732 | 2.4236 | 2.5415 | | 1.7952 | H8 | 2.1746 | 1.1002 | 2.9342 | 2.3774 | 2.5415 | 3.0917 | 1.7952 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
Cl4 |
113.053 |
|
C1 |
C2 |
H7 |
108.873 |
| C1 |
C2 |
H8 |
111.297 |
|
C2 |
C1 |
Cl3 |
113.053 |
| C2 |
C1 |
H5 |
108.873 |
|
C2 |
C1 |
H6 |
111.297 |
| Cl3 |
C1 |
H5 |
106.942 |
|
Cl3 |
C1 |
H6 |
107.375 |
| Cl4 |
C2 |
H7 |
106.942 |
|
Cl4 |
C2 |
H8 |
107.375 |
| H5 |
C1 |
H6 |
109.153 |
|
H7 |
C2 |
H8 |
109.153 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.401 |
|
|
|
| 2 |
C |
-0.401 |
|
|
|
| 3 |
Cl |
-0.043 |
|
|
|
| 4 |
Cl |
-0.043 |
|
|
|
| 5 |
H |
0.216 |
|
|
|
| 6 |
H |
0.228 |
|
|
|
| 7 |
H |
0.216 |
|
|
|
| 8 |
H |
0.228 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
2.856 |
2.856 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-37.259 |
1.202 |
0.000 |
| y |
1.202 |
-42.104 |
0.000 |
| z |
0.000 |
0.000 |
-35.111 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.348 |
1.202 |
0.000 |
| y |
1.202 |
-5.918 |
0.000 |
| z |
0.000 |
0.000 |
4.570 |
|
| Polar |
|---|
| 3z2-r2 | 9.140 |
|---|
| x2-y2 | 4.845 |
|---|
| xy | 1.202 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.765 |
-0.591 |
0.000 |
| y |
-0.591 |
6.968 |
0.000 |
| z |
0.000 |
0.000 |
6.426 |
<r2> (average value of r
2) Å
2
| <r2> |
166.720 |
| (<r2>)1/2 |
12.912 |
