return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for NH2CH2CH2CH3 (1-Propanamine)

using model chemistry: CID/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at CID/6-31G*
 hartrees
Energy at 0K-173.803209
Energy at 298.15K-173.813795
HF Energy-173.282143
Nuclear repulsion energy131.480063
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CID/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3602 3327 0.21      
2 A' 3230 2984 35.07      
3 A' 3154 2913 58.31      
4 A' 3148 2908 12.20      
5 A' 3133 2894 26.88      
6 A' 1772 1636 28.71      
7 A' 1603 1480 4.22      
8 A' 1589 1468 1.64      
9 A' 1581 1460 0.12      
10 A' 1510 1395 1.56      
11 A' 1493 1379 7.35      
12 A' 1402 1295 9.41      
13 A' 1209 1117 6.82      
14 A' 1162 1073 16.23      
15 A' 1099 1015 1.80      
16 A' 986 911 152.03      
17 A' 907 837 69.66      
18 A' 468 433 3.08      
19 A' 281 259 4.27      
20 A" 3690 3409 0.00      
21 A" 3224 2978 61.70      
22 A" 3199 2955 35.77      
23 A" 3166 2924 6.10      
24 A" 1592 1470 7.00      
25 A" 1462 1350 0.37      
26 A" 1395 1289 0.38      
27 A" 1317 1217 0.00      
28 A" 1086 1003 0.07      
29 A" 906 836 1.66      
30 A" 777 718 1.83      
31 A" 309 285 54.66      
32 A" 241 223 5.90      
33 A" 140 129 0.36      

Unscaled Zero Point Vibrational Energy (zpe) 27914.4 cm-1
Scaled (by 0.9237) Zero Point Vibrational Energy (zpe) 25784.6 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at CID/6-31G*
ABC
0.83222 0.12371 0.11659

See section I.F.4 to change rotational constant units
Geometric Data calculated at CID/6-31G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 1.425 1.287 0.000
C2 0.000 0.745 0.000
C3 -0.052 -0.781 0.000
N4 -1.386 -1.371 0.000
H5 1.437 2.377 0.000
H6 1.972 0.948 0.881
H7 1.972 0.948 -0.881
H8 -0.538 1.120 0.875
H9 -0.538 1.120 -0.875
H10 0.481 -1.163 -0.873
H11 0.481 -1.163 0.873
H12 -1.903 -1.051 -0.810
H13 -1.903 -1.051 0.810

Atom - Atom Distances (Å)
  C1 C2 C3 N4 H5 H6 H7 H8 H9 H10 H11 H12 H13
C11.52442.54083.86821.08961.09071.09072.15592.15592.76692.76694.14684.1468
C21.52441.52692.52942.17352.16922.16921.09351.09352.15292.15292.73932.7393
C32.54081.52691.45863.49032.80352.80352.14782.14781.09191.09192.03822.0382
N43.86822.52941.45864.69114.17454.17452.77242.77242.07112.07111.01221.0122
H51.08962.17353.49034.69111.76151.76152.49912.49913.76873.76874.85344.8534
H61.09072.16922.80354.17451.76151.76112.51603.06823.12342.58454.67624.3607
H71.09072.16922.80354.17451.76151.76113.06822.51602.58453.12344.36074.6762
H82.15591.09352.14782.77242.49912.51603.06821.75073.05042.49963.06812.5649
H92.15591.09352.14782.77242.49913.06822.51601.75072.49963.05042.56493.0681
H102.76692.15291.09192.07113.76873.12342.58453.05042.49961.74632.38692.9197
H112.76692.15291.09192.07113.76872.58453.12342.49963.05041.74632.91972.3869
H124.14682.73932.03821.01224.85344.67624.36073.06812.56492.38692.91971.6191
H134.14682.73932.03821.01224.85344.36074.67622.56493.06812.91972.38691.6191

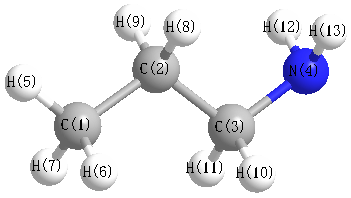
picture of 1-Propanamine state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 112.756 C1 C2 H8 109.781
C1 C2 H9 109.781 C2 C1 H5 111.424
C2 C1 H6 111.007 C2 C1 H7 111.007
C2 C3 N4 115.809 C2 C3 H10 109.474
C2 C3 H11 109.474 C3 C2 H8 108.980
C3 C2 H9 108.980 C3 N4 H12 109.839
C3 N4 H13 109.839 N4 C3 H10 107.724
N4 C3 H11 107.724 H5 C1 H6 107.783
H5 C1 H7 107.783 H6 C1 H7 107.669
H8 C2 H9 106.354 H10 C3 H11 106.196
H12 N4 H13 106.218
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability