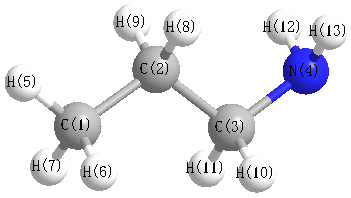
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3480 |
3330 |
1.70 |
|
|
|
| 2 |
A' |
3131 |
2996 |
35.07 |
|
|
|
| 3 |
A' |
3052 |
2920 |
47.62 |
|
|
|
| 4 |
A' |
3044 |
2913 |
42.31 |
|
|
|
| 5 |
A' |
3028 |
2897 |
18.85 |
|
|
|
| 6 |
A' |
1702 |
1628 |
23.37 |
|
|
|
| 7 |
A' |
1534 |
1467 |
4.87 |
|
|
|
| 8 |
A' |
1519 |
1453 |
0.96 |
|
|
|
| 9 |
A' |
1508 |
1443 |
0.27 |
|
|
|
| 10 |
A' |
1432 |
1370 |
3.10 |
|
|
|
| 11 |
A' |
1417 |
1356 |
5.79 |
|
|
|
| 12 |
A' |
1338 |
1280 |
9.47 |
|
|
|
| 13 |
A' |
1157 |
1107 |
3.69 |
|
|
|
| 14 |
A' |
1110 |
1062 |
14.03 |
|
|
|
| 15 |
A' |
1053 |
1007 |
1.32 |
|
|
|
| 16 |
A' |
932 |
892 |
141.10 |
|
|
|
| 17 |
A' |
859 |
822 |
82.47 |
|
|
|
| 18 |
A' |
446 |
427 |
3.46 |
|
|
|
| 19 |
A' |
262 |
251 |
4.09 |
|
|
|
| 20 |
A" |
3569 |
3415 |
0.39 |
|
|
|
| 21 |
A" |
3124 |
2989 |
57.72 |
|
|
|
| 22 |
A" |
3092 |
2958 |
43.17 |
|
|
|
| 23 |
A" |
3060 |
2928 |
4.25 |
|
|
|
| 24 |
A" |
1525 |
1459 |
7.65 |
|
|
|
| 25 |
A" |
1406 |
1346 |
0.42 |
|
|
|
| 26 |
A" |
1337 |
1279 |
0.48 |
|
|
|
| 27 |
A" |
1254 |
1200 |
0.02 |
|
|
|
| 28 |
A" |
1045 |
999 |
0.01 |
|
|
|
| 29 |
A" |
866 |
828 |
1.71 |
|
|
|
| 30 |
A" |
749 |
716 |
3.19 |
|
|
|
| 31 |
A" |
308 |
295 |
51.91 |
|
|
|
| 32 |
A" |
227 |
217 |
5.65 |
|
|
|
| 33 |
A" |
132 |
127 |
0.21 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 26849.2 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 25686.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.512 |
|
|
|
| 2 |
C |
-0.315 |
|
|
|
| 3 |
C |
-0.187 |
|
|
|
| 4 |
N |
-0.735 |
|
|
|
| 5 |
H |
0.169 |
|
|
|
| 6 |
H |
0.167 |
|
|
|
| 7 |
H |
0.167 |
|
|
|
| 8 |
H |
0.151 |
|
|
|
| 9 |
H |
0.151 |
|
|
|
| 10 |
H |
0.164 |
|
|
|
| 11 |
H |
0.164 |
|
|
|
| 12 |
H |
0.308 |
|
|
|
| 13 |
H |
0.308 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.227 |
1.491 |
0.000 |
1.508 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-25.445 |
-2.023 |
0.000 |
| y |
-2.023 |
-32.456 |
0.000 |
| z |
0.000 |
0.000 |
-25.288 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.428 |
-2.023 |
0.000 |
| y |
-2.023 |
-7.090 |
0.000 |
| z |
0.000 |
0.000 |
3.662 |
|
| Polar |
|---|
| 3z2-r2 | 7.324 |
|---|
| x2-y2 | 7.012 |
|---|
| xy | -2.023 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.399 |
0.546 |
0.000 |
| y |
0.546 |
5.852 |
0.000 |
| z |
0.000 |
0.000 |
5.668 |
<r2> (average value of r
2) Å
2
| <r2> |
112.782 |
| (<r2>)1/2 |
10.620 |