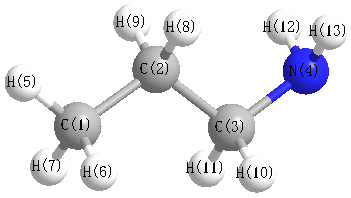
Vibrational Frequencies calculated at B2PLYP/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3483 |
3306 |
1.53 |
|
|
|
| 2 |
A' |
3154 |
2994 |
35.44 |
|
|
|
| 3 |
A' |
3078 |
2922 |
50.63 |
|
|
|
| 4 |
A' |
3072 |
2916 |
29.13 |
|
|
|
| 5 |
A' |
3054 |
2899 |
21.63 |
|
|
|
| 6 |
A' |
1719 |
1632 |
22.60 |
|
|
|
| 7 |
A' |
1560 |
1481 |
3.30 |
|
|
|
| 8 |
A' |
1545 |
1467 |
1.06 |
|
|
|
| 9 |
A' |
1534 |
1456 |
0.18 |
|
|
|
| 10 |
A' |
1458 |
1384 |
2.13 |
|
|
|
| 11 |
A' |
1437 |
1364 |
5.21 |
|
|
|
| 12 |
A' |
1359 |
1290 |
11.66 |
|
|
|
| 13 |
A' |
1170 |
1111 |
6.60 |
|
|
|
| 14 |
A' |
1112 |
1056 |
10.65 |
|
|
|
| 15 |
A' |
1057 |
1004 |
2.61 |
|
|
|
| 16 |
A' |
953 |
904 |
148.95 |
|
|
|
| 17 |
A' |
873 |
829 |
64.31 |
|
|
|
| 18 |
A' |
451 |
428 |
3.05 |
|
|
|
| 19 |
A' |
268 |
255 |
4.05 |
|
|
|
| 20 |
A" |
3572 |
3391 |
0.58 |
|
|
|
| 21 |
A" |
3148 |
2988 |
60.13 |
|
|
|
| 22 |
A" |
3119 |
2961 |
39.05 |
|
|
|
| 23 |
A" |
3087 |
2930 |
6.35 |
|
|
|
| 24 |
A" |
1551 |
1472 |
6.38 |
|
|
|
| 25 |
A" |
1422 |
1349 |
0.47 |
|
|
|
| 26 |
A" |
1355 |
1287 |
0.50 |
|
|
|
| 27 |
A" |
1276 |
1212 |
0.00 |
|
|
|
| 28 |
A" |
1057 |
1003 |
0.06 |
|
|
|
| 29 |
A" |
879 |
834 |
1.63 |
|
|
|
| 30 |
A" |
759 |
721 |
2.07 |
|
|
|
| 31 |
A" |
312 |
296 |
51.58 |
|
|
|
| 32 |
A" |
232 |
221 |
5.33 |
|
|
|
| 33 |
A" |
136 |
129 |
0.22 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 27120.9 cm
-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 25743.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.461 |
|
|
|
| 2 |
C |
-0.275 |
|
|
|
| 3 |
C |
-0.142 |
|
|
|
| 4 |
N |
-0.731 |
|
|
|
| 5 |
H |
0.152 |
|
|
|
| 6 |
H |
0.150 |
|
|
|
| 7 |
H |
0.150 |
|
|
|
| 8 |
H |
0.132 |
|
|
|
| 9 |
H |
0.132 |
|
|
|
| 10 |
H |
0.146 |
|
|
|
| 11 |
H |
0.146 |
|
|
|
| 12 |
H |
0.300 |
|
|
|
| 13 |
H |
0.300 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.190 |
1.513 |
0.000 |
1.525 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-25.919 |
2.115 |
0.000 |
| y |
2.115 |
-32.844 |
0.000 |
| z |
0.000 |
0.000 |
-25.632 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.319 |
2.115 |
0.000 |
| y |
2.115 |
-7.069 |
0.000 |
| z |
0.000 |
0.000 |
3.750 |
|
| Polar |
|---|
| 3z2-r2 | 7.499 |
|---|
| x2-y2 | 6.925 |
|---|
| xy | 2.115 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.288 |
0.516 |
0.000 |
| y |
0.516 |
5.735 |
0.000 |
| z |
0.000 |
0.000 |
5.611 |
<r2> (average value of r
2) Å
2
| <r2> |
113.041 |
| (<r2>)1/2 |
10.632 |