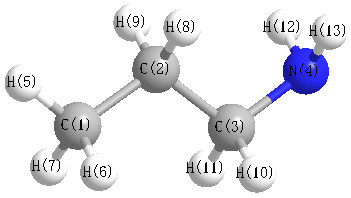
Vibrational Frequencies calculated at B3LYP/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3450 |
3313 |
2.68 |
|
|
|
| 2 |
A' |
3114 |
2990 |
39.00 |
|
|
|
| 3 |
A' |
3044 |
2923 |
48.92 |
|
|
|
| 4 |
A' |
3034 |
2914 |
39.14 |
|
|
|
| 5 |
A' |
3017 |
2897 |
21.63 |
|
|
|
| 6 |
A' |
1702 |
1634 |
20.64 |
|
|
|
| 7 |
A' |
1540 |
1479 |
2.99 |
|
|
|
| 8 |
A' |
1524 |
1464 |
0.94 |
|
|
|
| 9 |
A' |
1513 |
1453 |
0.19 |
|
|
|
| 10 |
A' |
1438 |
1381 |
2.17 |
|
|
|
| 11 |
A' |
1420 |
1363 |
5.15 |
|
|
|
| 12 |
A' |
1345 |
1291 |
12.93 |
|
|
|
| 13 |
A' |
1153 |
1107 |
5.23 |
|
|
|
| 14 |
A' |
1094 |
1051 |
11.08 |
|
|
|
| 15 |
A' |
1038 |
997 |
2.27 |
|
|
|
| 16 |
A' |
935 |
898 |
144.80 |
|
|
|
| 17 |
A' |
856 |
822 |
68.87 |
|
|
|
| 18 |
A' |
445 |
427 |
3.05 |
|
|
|
| 19 |
A' |
265 |
254 |
3.93 |
|
|
|
| 20 |
A" |
3534 |
3393 |
1.40 |
|
|
|
| 21 |
A" |
3106 |
2983 |
66.80 |
|
|
|
| 22 |
A" |
3077 |
2954 |
41.10 |
|
|
|
| 23 |
A" |
3043 |
2923 |
6.00 |
|
|
|
| 24 |
A" |
1530 |
1469 |
6.15 |
|
|
|
| 25 |
A" |
1408 |
1352 |
0.39 |
|
|
|
| 26 |
A" |
1339 |
1286 |
0.51 |
|
|
|
| 27 |
A" |
1258 |
1209 |
0.01 |
|
|
|
| 28 |
A" |
1047 |
1005 |
0.04 |
|
|
|
| 29 |
A" |
868 |
834 |
1.54 |
|
|
|
| 30 |
A" |
752 |
723 |
2.33 |
|
|
|
| 31 |
A" |
308 |
296 |
49.61 |
|
|
|
| 32 |
A" |
228 |
219 |
5.47 |
|
|
|
| 33 |
A" |
132 |
127 |
0.18 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 26777.1 cm
-1
Scaled (by 0.9603) Zero Point Vibrational Energy (zpe) 25714.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.444 |
|
|
|
| 2 |
C |
-0.255 |
|
|
|
| 3 |
C |
-0.135 |
|
|
|
| 4 |
N |
-0.704 |
|
|
|
| 5 |
H |
0.144 |
|
|
|
| 6 |
H |
0.144 |
|
|
|
| 7 |
H |
0.144 |
|
|
|
| 8 |
H |
0.124 |
|
|
|
| 9 |
H |
0.124 |
|
|
|
| 10 |
H |
0.139 |
|
|
|
| 11 |
H |
0.139 |
|
|
|
| 12 |
H |
0.290 |
|
|
|
| 13 |
H |
0.290 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.212 |
1.470 |
0.000 |
1.485 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-25.717 |
-2.016 |
0.000 |
| y |
-2.016 |
-32.581 |
0.000 |
| z |
0.000 |
0.000 |
-25.532 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.339 |
-2.016 |
0.000 |
| y |
-2.016 |
-6.956 |
0.000 |
| z |
0.000 |
0.000 |
3.617 |
|
| Polar |
|---|
| 3z2-r2 | 7.234 |
|---|
| x2-y2 | 6.864 |
|---|
| xy | -2.016 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.404 |
0.554 |
0.000 |
| y |
0.554 |
5.843 |
0.000 |
| z |
0.000 |
0.000 |
5.671 |
<r2> (average value of r
2) Å
2
| <r2> |
113.458 |
| (<r2>)1/2 |
10.652 |