return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C2H8N2 (Ethylenediamine)

using model chemistry: MP3/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2H 1Ag
Energy calculated at MP3/6-31G*
 hartrees
Energy at 0K-189.892223
Energy at 298.15K-189.902564
HF Energy-189.264970
Nuclear repulsion energy131.507993
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP3/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 Ag 3516 3300 0.00      
2 Ag 3084 2895 0.00      
3 Ag 1737 1631 0.00      
4 Ag 1548 1453 0.00      
5 Ag 1455 1366 0.00      
6 Ag 1166 1095 0.00      
7 Ag 1118 1050 0.00      
8 Ag 904 849 0.00      
9 Ag 474 445 0.00      
10 Au 3603 3381 0.14      
11 Au 3145 2952 68.05      
12 Au 1440 1352 2.13      
13 Au 1104 1036 0.45      
14 Au 776 728 0.28      
15 Au 285 267 130.48      
16 Au 155 145 0.12      
17 Bg 3603 3382 0.00      
18 Bg 3120 2928 0.00      
19 Bg 1414 1327 0.00      
20 Bg 1348 1265 0.00      
21 Bg 974 914 0.00      
22 Bg 318 298 0.00      
23 Bu 3516 3300 0.95      
24 Bu 3091 2901 76.43      
25 Bu 1735 1629 55.59      
26 Bu 1564 1468 1.10      
27 Bu 1369 1285 27.26      
28 Bu 1131 1062 6.22      
29 Bu 951 892 419.94      
30 Bu 283 266 21.25      

Unscaled Zero Point Vibrational Energy (zpe) 24962.8 cm-1
Scaled (by 0.9386) Zero Point Vibrational Energy (zpe) 23430.1 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP3/6-31G*
ABC
0.87460 0.12484 0.11809

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP3/6-31G*

Point Group is C2h

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.399 0.655 0.000
C2 -0.399 -0.655 0.000
N3 -0.399 1.883 0.000
N4 0.399 -1.883 0.000
H5 1.013 -1.888 0.813
H6 1.013 -1.888 -0.813
H7 -1.013 1.888 0.813
H8 -1.013 1.888 -0.813
H9 -1.056 -0.678 0.879
H10 -1.056 -0.678 -0.879
H11 1.056 0.678 0.879
H12 1.056 0.678 -0.879

Atom - Atom Distances (Å)
  C1 C2 N3 N4 H5 H6 H7 H8 H9 H10 H11 H12
C11.53361.46492.53802.73962.73962.04352.04352.16012.16011.09771.0977
C21.53362.53801.46492.04352.04352.73962.73961.09771.09772.16012.1601
N31.46492.53803.85004.10844.10841.01871.01872.78612.78612.08432.0843
N42.53801.46493.85001.01871.01874.10844.10842.08432.08432.78612.7861
H52.73962.04354.10841.01871.62644.28554.58382.39822.93432.56723.0739
H62.73962.04354.10841.01871.62644.58384.28552.93432.39823.07392.5672
H72.04352.73961.01874.10844.28554.58381.62642.56723.07392.39822.9343
H82.04352.73961.01874.10844.58384.28551.62643.07392.56722.93432.3982
H92.16011.09772.78612.08432.39822.93432.56723.07391.75752.51023.0643
H102.16011.09772.78612.08432.93432.39823.07392.56721.75753.06432.5102
H111.09772.16012.08432.78612.56723.07392.39822.93432.51023.06431.7575
H121.09772.16012.08432.78613.07392.56722.93432.39823.06432.51021.7575

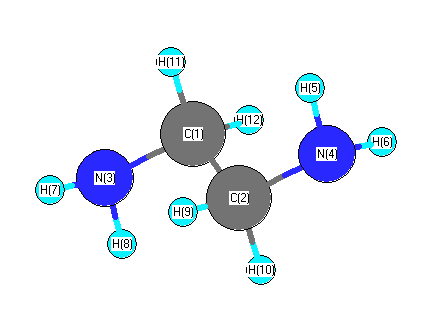
picture of Ethylenediamine state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 N4 115.628 C1 C2 H9 109.233
C1 C2 H10 109.233 C1 N3 H7 109.415
C1 N3 H8 109.415 C2 C1 N3 115.628
C2 C1 H11 109.233 C2 C1 H12 109.233
C2 N4 H5 109.415 C2 N4 H6 109.415
N3 C1 H11 107.991 N3 C1 H12 107.991
N4 C2 H9 107.991 N4 C2 H10 107.991
H5 N4 H6 105.927 H7 N3 H8 105.927
H9 C2 H10 106.365 H11 C1 H12 106.365
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability