Jump to
S1C2
Vibrational Frequencies calculated at B3PW91/6-31G*
Geometric Data calculated at B3PW91/6-31G*
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at B3PW91/6-31G*
| | hartrees |
|---|
| Energy at 0K | -207.867759 |
| Energy at 298.15K | -207.871129 |
| HF Energy | -207.867759 |
| Nuclear repulsion energy | 102.064432 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3783 |
3620 |
35.18 |
|
|
|
| 2 |
A |
3129 |
2993 |
5.02 |
|
|
|
| 3 |
A |
3032 |
2901 |
27.84 |
|
|
|
| 4 |
A |
2374 |
2271 |
0.28 |
|
|
|
| 5 |
A |
1506 |
1441 |
3.26 |
|
|
|
| 6 |
A |
1430 |
1368 |
47.82 |
|
|
|
| 7 |
A |
1390 |
1330 |
1.53 |
|
|
|
| 8 |
A |
1236 |
1182 |
18.81 |
|
|
|
| 9 |
A |
1108 |
1060 |
101.99 |
|
|
|
| 10 |
A |
995 |
951 |
24.05 |
|
|
|
| 11 |
A |
907 |
868 |
16.09 |
|
|
|
| 12 |
A |
586 |
561 |
2.36 |
|
|
|
| 13 |
A |
403 |
385 |
70.23 |
|
|
|
| 14 |
A |
314 |
300 |
92.01 |
|
|
|
| 15 |
A |
215 |
205 |
8.46 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 11203.1 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 10718.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3PW91/6-31G*
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.574 |
0.587 |
0.038 |
| C2 |
0.825 |
0.119 |
-0.007 |
| O3 |
-1.506 |
-0.455 |
-0.112 |
| H4 |
-0.715 |
1.155 |
0.970 |
| H5 |
-0.739 |
1.278 |
-0.794 |
| H6 |
-1.404 |
-1.063 |
0.633 |
| N7 |
1.914 |
-0.281 |
-0.014 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
O3 |
H4 |
H5 |
H6 |
N7 |
| C1 | | 1.4755 | 1.4059 | 1.1009 | 1.0943 | 1.9407 | 2.6356 |
C2 | 1.4755 | | 2.4027 | 2.0975 | 2.1002 | 2.6025 | 1.1606 | O3 | 1.4059 | 2.4027 | | 2.0954 | 2.0136 | 0.9682 | 3.4262 | H4 | 1.1009 | 2.0975 | 2.0954 | | 1.7684 | 2.3477 | 3.1537 | H5 | 1.0943 | 2.1002 | 2.0136 | 1.7684 | | 2.8216 | 3.1751 | H6 | 1.9407 | 2.6025 | 0.9682 | 2.3477 | 2.8216 | | 3.4699 | N7 | 2.6356 | 1.1606 | 3.4262 | 3.1537 | 3.1751 | 3.4699 | |
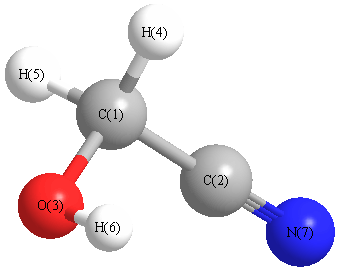 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N7 |
177.865 |
|
C1 |
O3 |
H6 |
108.246 |
| C2 |
C1 |
O3 |
112.976 |
|
C2 |
C1 |
H4 |
108.119 |
| C2 |
C1 |
H5 |
108.714 |
|
O3 |
C1 |
H4 |
112.848 |
| O3 |
C1 |
H5 |
106.632 |
|
H4 |
C1 |
H5 |
107.332 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.144 |
|
|
|
| 2 |
C |
0.348 |
|
|
|
| 3 |
O |
-0.593 |
|
|
|
| 4 |
H |
0.205 |
|
|
|
| 5 |
H |
0.233 |
|
|
|
| 6 |
H |
0.421 |
|
|
|
| 7 |
N |
-0.470 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-2.396 |
1.114 |
1.393 |
2.988 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-31.279 |
1.218 |
-2.014 |
| y |
1.218 |
-20.261 |
-1.816 |
| z |
-2.014 |
-1.816 |
-21.417 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-10.440 |
1.218 |
-2.014 |
| y |
1.218 |
6.087 |
-1.816 |
| z |
-2.014 |
-1.816 |
4.354 |
|
| Polar |
|---|
| 3z2-r2 | 8.707 |
|---|
| x2-y2 | -11.018 |
|---|
| xy | 1.218 |
|---|
| xz | -2.014 |
|---|
| yz | -1.816 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.302 |
-0.475 |
-0.037 |
| y |
-0.475 |
3.494 |
-0.137 |
| z |
-0.037 |
-0.137 |
2.989 |
<r2> (average value of r
2) Å
2
| <r2> |
78.615 |
| (<r2>)1/2 |
8.866 |
