Jump to
S1C2
Vibrational Frequencies calculated at MP3/6-31G*
Geometric Data calculated at MP3/6-31G*
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP3/6-31G*
| | hartrees |
|---|
| Energy at 0K | -207.359853 |
| Energy at 298.15K | -207.363227 |
| HF Energy | -206.763630 |
| Nuclear repulsion energy | 101.901515 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP3/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3813 |
3579 |
39.00 |
|
|
|
| 2 |
A |
3190 |
2994 |
4.75 |
|
|
|
| 3 |
A |
3105 |
2914 |
20.68 |
|
|
|
| 4 |
A |
2446 |
2295 |
0.69 |
|
|
|
| 5 |
A |
1555 |
1459 |
2.29 |
|
|
|
| 6 |
A |
1460 |
1370 |
52.96 |
|
|
|
| 7 |
A |
1421 |
1334 |
2.11 |
|
|
|
| 8 |
A |
1257 |
1179 |
20.11 |
|
|
|
| 9 |
A |
1133 |
1064 |
91.47 |
|
|
|
| 10 |
A |
1018 |
956 |
25.18 |
|
|
|
| 11 |
A |
930 |
873 |
22.79 |
|
|
|
| 12 |
A |
586 |
550 |
2.72 |
|
|
|
| 13 |
A |
396 |
371 |
85.62 |
|
|
|
| 14 |
A |
310 |
291 |
85.63 |
|
|
|
| 15 |
A |
211 |
199 |
8.92 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 11414.7 cm
-1
Scaled (by 0.9386) Zero Point Vibrational Energy (zpe) 10713.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP3/6-31G*
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.576 |
0.594 |
0.038 |
| C2 |
0.827 |
0.115 |
-0.006 |
| O3 |
-1.505 |
-0.459 |
-0.114 |
| H4 |
-0.721 |
1.153 |
0.971 |
| H5 |
-0.741 |
1.279 |
-0.795 |
| H6 |
-1.418 |
-1.055 |
0.645 |
| N7 |
1.916 |
-0.280 |
-0.014 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
O3 |
H4 |
H5 |
H6 |
N7 |
| C1 | | 1.4824 | 1.4123 | 1.0971 | 1.0916 | 1.9480 | 2.6413 |
C2 | 1.4824 | | 2.4035 | 2.1040 | 2.1060 | 2.6130 | 1.1592 | O3 | 1.4123 | 2.4035 | | 2.0948 | 2.0171 | 0.9694 | 3.4276 | H4 | 1.0971 | 2.1040 | 2.0948 | | 1.7707 | 2.3373 | 3.1590 | H5 | 1.0916 | 2.1060 | 2.0171 | 1.7707 | | 2.8249 | 3.1784 | H6 | 1.9480 | 2.6130 | 0.9694 | 2.3373 | 2.8249 | | 3.4858 | N7 | 2.6413 | 1.1592 | 3.4276 | 3.1590 | 3.1784 | 3.4858 | |
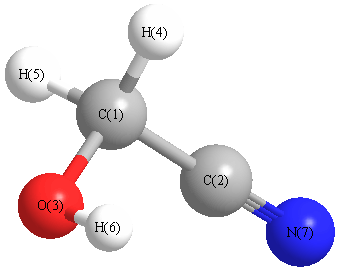 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N7 |
178.318 |
|
C1 |
O3 |
H6 |
108.317 |
| C2 |
C1 |
O3 |
112.238 |
|
C2 |
C1 |
H4 |
108.382 |
| C2 |
C1 |
H5 |
108.862 |
|
O3 |
C1 |
H4 |
112.581 |
| O3 |
C1 |
H5 |
106.627 |
|
H4 |
C1 |
H5 |
108.004 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
