return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C3H6O (2-Propen-1-ol)

using model chemistry: QCISD/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at QCISD/6-31G*
 hartrees
Energy at 0K-192.515537
Energy at 298.15K-192.521786
HF Energy-191.913219
Nuclear repulsion energy116.204982
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3766 3587 13.83      
2 A 3257 3102 18.44      
3 A 3193 3041 5.86      
4 A 3172 3021 11.74      
5 A 3049 2903 55.15      
6 A 3009 2865 55.58      
7 A 1749 1665 0.54      
8 A 1571 1496 1.42      
9 A 1514 1442 13.37      
10 A 1461 1391 5.52      
11 A 1337 1273 1.00      
12 A 1302 1240 46.78      
13 A 1270 1210 43.51      
14 A 1189 1132 19.66      
15 A 1102 1050 70.51      
16 A 1026 977 16.11      
17 A 998 950 7.28      
18 A 942 897 25.99      
19 A 940 895 12.42      
20 A 654 623 4.06      
21 A 454 432 4.28      
22 A 337 321 10.66      
23 A 275 262 136.66      
24 A 111 106 1.39      

Unscaled Zero Point Vibrational Energy (zpe) 18838.3 cm-1
Scaled (by 0.9523) Zero Point Vibrational Energy (zpe) 17939.7 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD/6-31G*
ABC
0.92562 0.14316 0.13699

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD/6-31G*

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.587 0.462 0.278
C2 -0.665 -0.364 0.251
C3 -1.841 0.075 -0.212
O4 1.629 -0.311 -0.323
H5 0.426 1.410 -0.261
H6 0.846 0.711 1.322
H7 -0.567 -1.372 0.654
H8 -2.733 -0.547 -0.191
H9 -1.956 1.074 -0.632
H10 2.461 0.175 -0.208

Atom - Atom Distances (Å)
  C1 C2 C3 O4 H5 H6 H7 H8 H9 H10
C11.50042.50761.43031.10231.10392.19963.50202.76941.9573
C21.50041.33792.36522.14422.14141.09082.12272.12403.2054
C32.50761.33793.49312.63133.15902.11411.08771.08954.3036
O41.43032.36523.49312.10132.08982.62694.37023.85540.9712
H51.10232.14422.63132.10131.78083.09253.71682.43362.3815
H61.10392.14143.15902.08981.78082.60424.08433.43502.2889
H72.19961.09082.11412.62693.09252.60422.46743.09313.5080
H83.50202.12271.08774.37023.71684.08432.46741.85125.2443
H92.76942.12401.08953.85542.43363.43503.09311.85124.5275
H101.95733.20544.30360.97122.38152.28893.50805.24434.5275

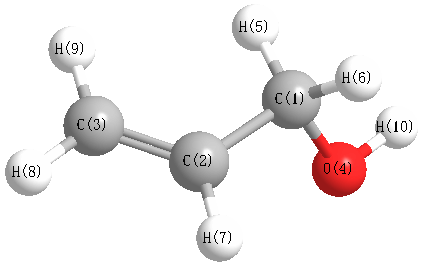
picture of 2-Propen-1-ol state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 124.033 C1 C2 H7 115.274
C1 O4 H10 107.647 C2 C1 O4 107.596
C2 C1 H5 110.000 C2 C1 H6 109.684
C2 C3 H8 121.772 C2 C3 H9 121.758
C3 C2 H7 120.689 O4 C1 H5 111.483
O4 C1 H6 110.445 H5 C1 H6 107.635
H8 C3 H9 116.469
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability