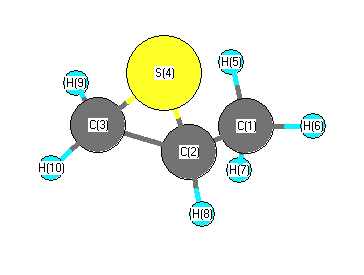
Vibrational Frequencies calculated at PBEPBEultrafine/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3146 |
3094 |
6.00 |
|
|
|
| 2 |
A |
3086 |
3035 |
21.54 |
|
|
|
| 3 |
A |
3070 |
3019 |
5.01 |
|
|
|
| 4 |
A |
3061 |
3011 |
14.91 |
|
|
|
| 5 |
A |
3052 |
3001 |
11.25 |
|
|
|
| 6 |
A |
2978 |
2928 |
25.73 |
|
|
|
| 7 |
A |
1478 |
1454 |
6.59 |
|
|
|
| 8 |
A |
1469 |
1445 |
5.45 |
|
|
|
| 9 |
A |
1458 |
1434 |
5.19 |
|
|
|
| 10 |
A |
1391 |
1368 |
4.93 |
|
|
|
| 11 |
A |
1353 |
1331 |
8.89 |
|
|
|
| 12 |
A |
1180 |
1160 |
3.76 |
|
|
|
| 13 |
A |
1161 |
1142 |
1.81 |
|
|
|
| 14 |
A |
1068 |
1050 |
20.56 |
|
|
|
| 15 |
A |
1038 |
1020 |
9.65 |
|
|
|
| 16 |
A |
998 |
982 |
4.28 |
|
|
|
| 17 |
A |
916 |
901 |
5.27 |
|
|
|
| 18 |
A |
908 |
893 |
4.19 |
|
|
|
| 19 |
A |
871 |
856 |
2.38 |
|
|
|
| 20 |
A |
646 |
635 |
9.79 |
|
|
|
| 21 |
A |
609 |
599 |
13.62 |
|
|
|
| 22 |
A |
378 |
371 |
0.33 |
|
|
|
| 23 |
A |
293 |
289 |
2.06 |
|
|
|
| 24 |
A |
234 |
230 |
0.08 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 17920.4 cm
-1
Scaled (by 0.9835) Zero Point Vibrational Energy (zpe) 17624.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.478 |
|
|
|
| 2 |
C |
-0.215 |
|
|
|
| 3 |
C |
-0.409 |
|
|
|
| 4 |
S |
-0.010 |
|
|
|
| 5 |
H |
0.178 |
|
|
|
| 6 |
H |
0.178 |
|
|
|
| 7 |
H |
0.168 |
|
|
|
| 8 |
H |
0.193 |
|
|
|
| 9 |
H |
0.196 |
|
|
|
| 10 |
H |
0.198 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.740 |
1.401 |
0.281 |
2.251 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-33.052 |
1.454 |
-0.467 |
| y |
1.454 |
-31.265 |
-0.458 |
| z |
-0.467 |
-0.458 |
-32.075 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.382 |
1.454 |
-0.467 |
| y |
1.454 |
1.299 |
-0.458 |
| z |
-0.467 |
-0.458 |
0.083 |
|
| Polar |
|---|
| 3z2-r2 | 0.166 |
|---|
| x2-y2 | -1.787 |
|---|
| xy | 1.454 |
|---|
| xz | -0.467 |
|---|
| yz | -0.458 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.109 |
-0.598 |
-0.270 |
| y |
-0.598 |
6.997 |
-0.041 |
| z |
-0.270 |
-0.041 |
5.626 |
<r2> (average value of r
2) Å
2
| <r2> |
104.848 |
| (<r2>)1/2 |
10.240 |