Jump to
S1C2
Vibrational Frequencies calculated at MP4=FULL/6-31G*
Geometric Data calculated at MP4=FULL/6-31G*
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP4=FULL/6-31G*
| | hartrees |
|---|
| Energy at 0K | -208.551861 |
| Energy at 298.15K | |
| HF Energy | -207.878179 |
| Nuclear repulsion energy | 118.970829 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP4=FULL/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3740 |
3596 |
|
|
|
|
| 2 |
A' |
3198 |
3075 |
|
|
|
|
| 3 |
A' |
3177 |
3055 |
|
|
|
|
| 4 |
A' |
3066 |
2948 |
|
|
|
|
| 5 |
A' |
1717 |
1651 |
|
|
|
|
| 6 |
A' |
1527 |
1468 |
|
|
|
|
| 7 |
A' |
1448 |
1393 |
|
|
|
|
| 8 |
A' |
1404 |
1350 |
|
|
|
|
| 9 |
A' |
1355 |
1303 |
|
|
|
|
| 10 |
A' |
1169 |
1124 |
|
|
|
|
| 11 |
A' |
944 |
908 |
|
|
|
|
| 12 |
A' |
906 |
871 |
|
|
|
|
| 13 |
A' |
670 |
645 |
|
|
|
|
| 14 |
A' |
317 |
305 |
|
|
|
|
| 15 |
A" |
3129 |
3009 |
|
|
|
|
| 16 |
A" |
1535 |
1476 |
|
|
|
|
| 17 |
A" |
1088 |
1046 |
|
|
|
|
| 18 |
A" |
859 |
826 |
|
|
|
|
| 19 |
A" |
488 |
470 |
|
|
|
|
| 20 |
A" |
380 |
365 |
|
|
|
|
| 21 |
A" |
82i |
79i |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16018.5 cm
-1
Scaled (by 0.9616) Zero Point Vibrational Energy (zpe) 15403.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP4=FULL/6-31G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.450 |
0.488 |
0.000 |
| C2 |
0.000 |
0.870 |
0.000 |
| N3 |
1.032 |
0.093 |
0.000 |
| O4 |
0.646 |
-1.278 |
0.000 |
| H5 |
1.515 |
-1.717 |
0.000 |
| H6 |
-1.571 |
-0.598 |
0.000 |
| H7 |
-1.948 |
0.905 |
0.885 |
| H8 |
-1.948 |
0.905 |
-0.885 |
| H9 |
0.266 |
1.929 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
O4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.4995 | 2.5129 | 2.7400 | 3.6948 | 1.0922 | 1.0982 | 1.0982 | 2.2410 |
C2 | 1.4995 | | 1.2914 | 2.2423 | 2.9977 | 2.1495 | 2.1403 | 2.1403 | 1.0920 | N3 | 2.5129 | 1.2914 | | 1.4241 | 1.8735 | 2.6924 | 3.2131 | 3.2131 | 1.9889 | O4 | 2.7400 | 2.2423 | 1.4241 | | 0.9741 | 2.3180 | 3.5038 | 3.5038 | 3.2288 | H5 | 3.6948 | 2.9977 | 1.8735 | 0.9741 | | 3.2822 | 4.4333 | 4.4333 | 3.8538 | H6 | 1.0922 | 2.1495 | 2.6924 | 2.3180 | 3.2822 | | 1.7847 | 1.7847 | 3.1237 | H7 | 1.0982 | 2.1403 | 3.2131 | 3.5038 | 4.4333 | 1.7847 | | 1.7701 | 2.5953 | H8 | 1.0982 | 2.1403 | 3.2131 | 3.5038 | 4.4333 | 1.7847 | 1.7701 | | 2.5953 | H9 | 2.2410 | 1.0920 | 1.9889 | 3.2288 | 3.8538 | 3.1237 | 2.5953 | 2.5953 | |
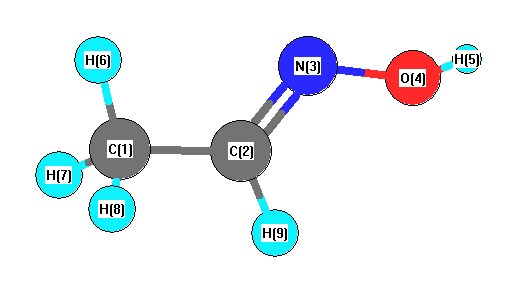 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N3 |
128.269 |
|
C1 |
C2 |
H9 |
118.872 |
| C2 |
C1 |
H6 |
111.098 |
|
C2 |
C1 |
H7 |
109.997 |
| C2 |
C1 |
H8 |
109.997 |
|
C2 |
N3 |
O4 |
111.237 |
| N3 |
C2 |
H9 |
112.859 |
|
N3 |
O4 |
H5 |
101.082 |
| H6 |
C1 |
H7 |
109.133 |
|
H6 |
C1 |
H8 |
109.133 |
| H7 |
C1 |
H8 |
107.399 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
