Jump to
S1C2
Energy calculated at B2PLYP/6-31G*
| | hartrees |
|---|
| Energy at 0K | -208.907812 |
| Energy at 298.15K | -208.913820 |
| HF Energy | -208.707296 |
| Nuclear repulsion energy | 116.950728 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3779 |
3587 |
69.02 |
|
|
|
| 2 |
A' |
3187 |
3025 |
13.56 |
|
|
|
| 3 |
A' |
3157 |
2996 |
11.31 |
|
|
|
| 4 |
A' |
3080 |
2923 |
18.07 |
|
|
|
| 5 |
A' |
1738 |
1649 |
1.09 |
|
|
|
| 6 |
A' |
1533 |
1455 |
7.67 |
|
|
|
| 7 |
A' |
1478 |
1403 |
7.10 |
|
|
|
| 8 |
A' |
1440 |
1367 |
25.68 |
|
|
|
| 9 |
A' |
1312 |
1245 |
56.16 |
|
|
|
| 10 |
A' |
1175 |
1116 |
4.89 |
|
|
|
| 11 |
A' |
1022 |
970 |
116.01 |
|
|
|
| 12 |
A' |
922 |
876 |
22.74 |
|
|
|
| 13 |
A' |
567 |
538 |
15.24 |
|
|
|
| 14 |
A' |
326 |
309 |
2.45 |
|
|
|
| 15 |
A" |
3138 |
2979 |
16.93 |
|
|
|
| 16 |
A" |
1529 |
1451 |
8.20 |
|
|
|
| 17 |
A" |
1102 |
1046 |
0.37 |
|
|
|
| 18 |
A" |
921 |
874 |
9.88 |
|
|
|
| 19 |
A" |
402 |
381 |
143.96 |
|
|
|
| 20 |
A" |
285 |
270 |
2.83 |
|
|
|
| 21 |
A" |
204 |
193 |
0.22 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16146.9 cm
-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 15326.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP/6-31G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.301 |
1.308 |
0.000 |
| C2 |
0.000 |
0.568 |
0.000 |
| N3 |
0.013 |
-0.709 |
0.000 |
| O4 |
1.313 |
-1.219 |
0.000 |
| H5 |
1.169 |
-2.176 |
0.000 |
| H6 |
-2.134 |
0.603 |
0.000 |
| H7 |
-1.379 |
1.952 |
0.882 |
| H8 |
-1.379 |
1.952 |
-0.882 |
| H9 |
0.935 |
1.134 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
O4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.4969 | 2.4069 | 3.6363 | 4.2702 | 1.0911 | 1.0949 | 1.0949 | 2.2433 |
C2 | 1.4969 | | 1.2768 | 2.2179 | 2.9821 | 2.1346 | 2.1438 | 2.1438 | 1.0933 | N3 | 2.4069 | 1.2768 | | 1.3974 | 1.8675 | 2.5161 | 3.1297 | 3.1297 | 2.0610 | O4 | 3.6363 | 2.2179 | 1.3974 | | 0.9674 | 3.8998 | 4.2526 | 4.2526 | 2.3834 | H5 | 4.2702 | 2.9821 | 1.8675 | 0.9674 | | 4.3164 | 4.9301 | 4.9301 | 3.3179 | H6 | 1.0911 | 2.1346 | 2.5161 | 3.8998 | 4.3164 | | 1.7795 | 1.7795 | 3.1152 | H7 | 1.0949 | 2.1438 | 3.1297 | 4.2526 | 4.9301 | 1.7795 | | 1.7640 | 2.6085 | H8 | 1.0949 | 2.1438 | 3.1297 | 4.2526 | 4.9301 | 1.7795 | 1.7640 | | 2.6085 | H9 | 2.2433 | 1.0933 | 2.0610 | 2.3834 | 3.3179 | 3.1152 | 2.6085 | 2.6085 | |
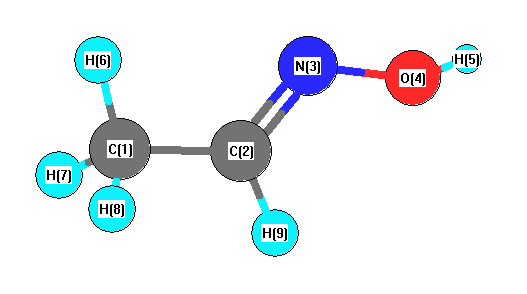 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N3 |
120.193 |
|
C1 |
C2 |
H9 |
119.188 |
| C2 |
C1 |
H6 |
110.150 |
|
C2 |
C1 |
H7 |
110.660 |
| C2 |
C1 |
H8 |
110.660 |
|
C2 |
N3 |
O4 |
111.984 |
| N3 |
C2 |
H9 |
120.619 |
|
N3 |
O4 |
H5 |
102.804 |
| H6 |
C1 |
H7 |
108.984 |
|
H6 |
C1 |
H8 |
108.984 |
| H7 |
C1 |
H8 |
107.333 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.511 |
|
|
|
| 2 |
C |
0.089 |
|
|
|
| 3 |
N |
-0.156 |
|
|
|
| 4 |
O |
-0.529 |
|
|
|
| 5 |
H |
0.425 |
|
|
|
| 6 |
H |
0.184 |
|
|
|
| 7 |
H |
0.171 |
|
|
|
| 8 |
H |
0.171 |
|
|
|
| 9 |
H |
0.157 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.401 |
0.495 |
0.000 |
0.637 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-25.232 |
-2.063 |
0.000 |
| y |
-2.063 |
-18.487 |
0.000 |
| z |
0.000 |
0.000 |
-24.927 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-3.525 |
-2.063 |
0.000 |
| y |
-2.063 |
6.593 |
0.000 |
| z |
0.000 |
0.000 |
-3.068 |
|
| Polar |
|---|
| 3z2-r2 | -6.136 |
|---|
| x2-y2 | -6.745 |
|---|
| xy | -2.063 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.976 |
-1.332 |
0.000 |
| y |
-1.332 |
6.598 |
0.000 |
| z |
0.000 |
0.000 |
2.936 |
<r2> (average value of r
2) Å
2
| <r2> |
91.910 |
| (<r2>)1/2 |
9.587 |
Jump to
S1C1
Energy calculated at B2PLYP/6-31G*
| | hartrees |
|---|
| Energy at 0K | -208.907206 |
| Energy at 298.15K | |
| HF Energy | -208.706388 |
| Nuclear repulsion energy | 119.466045 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3780 |
3588 |
70.48 |
|
|
|
| 2 |
A' |
3220 |
3056 |
8.79 |
|
|
|
| 3 |
A' |
3204 |
3041 |
7.48 |
|
|
|
| 4 |
A' |
3086 |
2929 |
12.56 |
|
|
|
| 5 |
A' |
1745 |
1656 |
5.19 |
|
|
|
| 6 |
A' |
1529 |
1452 |
12.87 |
|
|
|
| 7 |
A' |
1450 |
1376 |
14.04 |
|
|
|
| 8 |
A' |
1410 |
1339 |
31.71 |
|
|
|
| 9 |
A' |
1366 |
1297 |
48.44 |
|
|
|
| 10 |
A' |
1170 |
1111 |
10.14 |
|
|
|
| 11 |
A' |
947 |
899 |
88.93 |
|
|
|
| 12 |
A' |
923 |
876 |
41.40 |
|
|
|
| 13 |
A' |
679 |
645 |
13.17 |
|
|
|
| 14 |
A' |
315 |
299 |
1.47 |
|
|
|
| 15 |
A" |
3141 |
2981 |
14.25 |
|
|
|
| 16 |
A" |
1537 |
1459 |
9.85 |
|
|
|
| 17 |
A" |
1090 |
1035 |
0.22 |
|
|
|
| 18 |
A" |
872 |
827 |
12.03 |
|
|
|
| 19 |
A" |
499 |
474 |
37.31 |
|
|
|
| 20 |
A" |
382 |
362 |
97.25 |
|
|
|
| 21 |
A" |
51i |
48i |
0.19 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16146.2 cm
-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 15326.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP/6-31G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.449 |
0.487 |
0.000 |
| C2 |
0.000 |
0.865 |
0.000 |
| N3 |
1.023 |
0.092 |
0.000 |
| O4 |
0.652 |
-1.272 |
0.000 |
| H5 |
1.518 |
-1.707 |
0.000 |
| H6 |
-1.573 |
-0.595 |
0.000 |
| H7 |
-1.947 |
0.905 |
0.881 |
| H8 |
-1.947 |
0.905 |
-0.881 |
| H9 |
0.267 |
1.919 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
O4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.4976 | 2.5029 | 2.7400 | 3.6901 | 1.0882 | 1.0948 | 1.0948 | 2.2353 |
C2 | 1.4976 | | 1.2821 | 2.2342 | 2.9868 | 2.1461 | 2.1374 | 2.1374 | 1.0875 | N3 | 2.5029 | 1.2821 | | 1.4129 | 1.8658 | 2.6852 | 3.2024 | 3.2024 | 1.9776 | O4 | 2.7400 | 2.2342 | 1.4129 | | 0.9692 | 2.3266 | 3.5028 | 3.5028 | 3.2142 | H5 | 3.6901 | 2.9868 | 1.8658 | 0.9692 | | 3.2859 | 4.4278 | 4.4278 | 3.8361 | H6 | 1.0882 | 2.1461 | 2.6852 | 2.3266 | 3.2859 | | 1.7784 | 1.7784 | 3.1153 | H7 | 1.0948 | 2.1374 | 3.2024 | 3.5028 | 4.4278 | 1.7784 | | 1.7617 | 2.5897 | H8 | 1.0948 | 2.1374 | 3.2024 | 3.5028 | 4.4278 | 1.7784 | 1.7617 | | 2.5897 | H9 | 2.2353 | 1.0875 | 1.9776 | 3.2142 | 3.8361 | 3.1153 | 2.5897 | 2.5897 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N3 |
128.269 |
|
C1 |
C2 |
H9 |
118.842 |
| C2 |
C1 |
H6 |
111.203 |
|
C2 |
C1 |
H7 |
110.102 |
| C2 |
C1 |
H8 |
110.102 |
|
C2 |
N3 |
O4 |
111.904 |
| N3 |
C2 |
H9 |
112.890 |
|
N3 |
O4 |
H5 |
101.488 |
| H6 |
C1 |
H7 |
109.101 |
|
H6 |
C1 |
H8 |
109.101 |
| H7 |
C1 |
H8 |
107.133 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.522 |
|
|
|
| 2 |
C |
0.060 |
|
|
|
| 3 |
N |
-0.148 |
|
|
|
| 4 |
O |
-0.533 |
|
|
|
| 5 |
H |
0.428 |
|
|
|
| 6 |
H |
0.205 |
|
|
|
| 7 |
H |
0.173 |
|
|
|
| 8 |
H |
0.173 |
|
|
|
| 9 |
H |
0.165 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.575 |
0.214 |
0.000 |
0.614 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-22.720 |
-3.874 |
0.000 |
| y |
-3.874 |
-20.998 |
0.000 |
| z |
0.000 |
0.000 |
-24.948 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.253 |
-3.874 |
0.000 |
| y |
-3.874 |
2.836 |
0.000 |
| z |
0.000 |
0.000 |
-3.089 |
|
| Polar |
|---|
| 3z2-r2 | -6.178 |
|---|
| x2-y2 | -1.722 |
|---|
| xy | -3.874 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.430 |
-1.239 |
0.000 |
| y |
-1.239 |
5.734 |
0.000 |
| z |
0.000 |
0.000 |
2.929 |
<r2> (average value of r
2) Å
2
| <r2> |
79.100 |
| (<r2>)1/2 |
8.894 |
