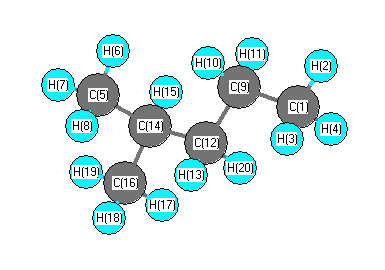
Vibrational Frequencies calculated at PBEPBEultrafine/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3058 |
3008 |
44.43 |
|
|
|
| 2 |
A |
3055 |
3005 |
29.33 |
|
|
|
| 3 |
A |
3051 |
3001 |
43.52 |
|
|
|
| 4 |
A |
3049 |
2999 |
55.12 |
|
|
|
| 5 |
A |
3045 |
2995 |
55.29 |
|
|
|
| 6 |
A |
3040 |
2990 |
6.15 |
|
|
|
| 7 |
A |
3007 |
2958 |
34.17 |
|
|
|
| 8 |
A |
2977 |
2928 |
34.64 |
|
|
|
| 9 |
A |
2977 |
2928 |
12.97 |
|
|
|
| 10 |
A |
2974 |
2925 |
29.06 |
|
|
|
| 11 |
A |
2970 |
2921 |
34.58 |
|
|
|
| 12 |
A |
2963 |
2914 |
33.40 |
|
|
|
| 13 |
A |
2942 |
2893 |
26.43 |
|
|
|
| 14 |
A |
2935 |
2887 |
5.50 |
|
|
|
| 15 |
A |
1494 |
1469 |
11.92 |
|
|
|
| 16 |
A |
1490 |
1465 |
3.10 |
|
|
|
| 17 |
A |
1487 |
1463 |
2.57 |
|
|
|
| 18 |
A |
1482 |
1457 |
9.37 |
|
|
|
| 19 |
A |
1477 |
1452 |
1.58 |
|
|
|
| 20 |
A |
1473 |
1449 |
0.92 |
|
|
|
| 21 |
A |
1470 |
1446 |
0.69 |
|
|
|
| 22 |
A |
1458 |
1434 |
0.11 |
|
|
|
| 23 |
A |
1396 |
1373 |
4.41 |
|
|
|
| 24 |
A |
1389 |
1367 |
2.21 |
|
|
|
| 25 |
A |
1377 |
1354 |
6.07 |
|
|
|
| 26 |
A |
1367 |
1344 |
2.18 |
|
|
|
| 27 |
A |
1340 |
1318 |
3.17 |
|
|
|
| 28 |
A |
1325 |
1304 |
0.73 |
|
|
|
| 29 |
A |
1301 |
1280 |
0.34 |
|
|
|
| 30 |
A |
1268 |
1247 |
1.97 |
|
|
|
| 31 |
A |
1243 |
1222 |
2.88 |
|
|
|
| 32 |
A |
1177 |
1157 |
0.55 |
|
|
|
| 33 |
A |
1171 |
1152 |
3.18 |
|
|
|
| 34 |
A |
1147 |
1129 |
2.38 |
|
|
|
| 35 |
A |
1069 |
1052 |
0.57 |
|
|
|
| 36 |
A |
1039 |
1022 |
0.54 |
|
|
|
| 37 |
A |
1006 |
990 |
0.16 |
|
|
|
| 38 |
A |
954 |
938 |
3.35 |
|
|
|
| 39 |
A |
948 |
933 |
1.17 |
|
|
|
| 40 |
A |
909 |
894 |
2.15 |
|
|
|
| 41 |
A |
889 |
875 |
0.67 |
|
|
|
| 42 |
A |
845 |
831 |
1.53 |
|
|
|
| 43 |
A |
807 |
794 |
0.74 |
|
|
|
| 44 |
A |
730 |
718 |
3.47 |
|
|
|
| 45 |
A |
436 |
429 |
0.07 |
|
|
|
| 46 |
A |
430 |
423 |
0.26 |
|
|
|
| 47 |
A |
366 |
360 |
0.14 |
|
|
|
| 48 |
A |
313 |
308 |
0.02 |
|
|
|
| 49 |
A |
255 |
250 |
0.00 |
|
|
|
| 50 |
A |
245 |
241 |
0.00 |
|
|
|
| 51 |
A |
225 |
222 |
0.02 |
|
|
|
| 52 |
A |
175 |
172 |
0.02 |
|
|
|
| 53 |
A |
109 |
108 |
0.02 |
|
|
|
| 54 |
A |
75 |
73 |
0.01 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 40601.2 cm
-1
Scaled (by 0.9835) Zero Point Vibrational Energy (zpe) 39931.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.483 |
|
|
|
| 2 |
H |
0.155 |
|
|
|
| 3 |
H |
0.156 |
|
|
|
| 4 |
H |
0.155 |
|
|
|
| 5 |
C |
-0.482 |
|
|
|
| 6 |
H |
0.158 |
|
|
|
| 7 |
H |
0.153 |
|
|
|
| 8 |
H |
0.151 |
|
|
|
| 9 |
C |
-0.285 |
|
|
|
| 10 |
H |
0.149 |
|
|
|
| 11 |
H |
0.144 |
|
|
|
| 12 |
C |
-0.264 |
|
|
|
| 13 |
H |
0.136 |
|
|
|
| 14 |
C |
-0.091 |
|
|
|
| 15 |
H |
0.135 |
|
|
|
| 16 |
C |
-0.480 |
|
|
|
| 17 |
H |
0.152 |
|
|
|
| 18 |
H |
0.151 |
|
|
|
| 19 |
H |
0.152 |
|
|
|
| 20 |
H |
0.138 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.015 |
0.031 |
-0.075 |
0.083 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-41.868 |
0.037 |
0.045 |
| y |
0.037 |
-41.583 |
0.147 |
| z |
0.045 |
0.147 |
-41.047 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.554 |
0.037 |
0.045 |
| y |
0.037 |
-0.125 |
0.147 |
| z |
0.045 |
0.147 |
0.679 |
|
| Polar |
|---|
| 3z2-r2 | 1.358 |
|---|
| x2-y2 | -0.286 |
|---|
| xy | 0.037 |
|---|
| xz | 0.045 |
|---|
| yz | 0.147 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
11.449 |
0.015 |
0.004 |
| y |
0.015 |
9.563 |
-0.003 |
| z |
0.004 |
-0.003 |
8.851 |
<r2> (average value of r
2) Å
2
| <r2> |
254.846 |
| (<r2>)1/2 |
15.964 |