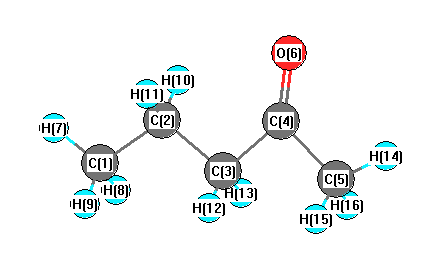
Vibrational Frequencies calculated at PBE1PBE/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3198 |
3039 |
8.13 |
|
|
|
| 2 |
A' |
3150 |
2993 |
32.63 |
|
|
|
| 3 |
A' |
3090 |
2936 |
17.28 |
|
|
|
| 4 |
A' |
3074 |
2921 |
4.50 |
|
|
|
| 5 |
A' |
3063 |
2911 |
26.83 |
|
|
|
| 6 |
A' |
3042 |
2891 |
13.08 |
|
|
|
| 7 |
A' |
1853 |
1760 |
136.78 |
|
|
|
| 8 |
A' |
1533 |
1457 |
5.60 |
|
|
|
| 9 |
A' |
1513 |
1438 |
3.29 |
|
|
|
| 10 |
A' |
1492 |
1417 |
18.68 |
|
|
|
| 11 |
A' |
1476 |
1403 |
2.79 |
|
|
|
| 12 |
A' |
1438 |
1366 |
9.04 |
|
|
|
| 13 |
A' |
1427 |
1356 |
50.97 |
|
|
|
| 14 |
A' |
1406 |
1336 |
24.72 |
|
|
|
| 15 |
A' |
1338 |
1271 |
18.21 |
|
|
|
| 16 |
A' |
1202 |
1142 |
60.73 |
|
|
|
| 17 |
A' |
1149 |
1092 |
0.13 |
|
|
|
| 18 |
A' |
1082 |
1029 |
0.14 |
|
|
|
| 19 |
A' |
973 |
924 |
2.00 |
|
|
|
| 20 |
A' |
928 |
882 |
9.15 |
|
|
|
| 21 |
A' |
841 |
799 |
1.17 |
|
|
|
| 22 |
A' |
601 |
571 |
12.09 |
|
|
|
| 23 |
A' |
393 |
374 |
1.19 |
|
|
|
| 24 |
A' |
342 |
325 |
2.98 |
|
|
|
| 25 |
A' |
172 |
164 |
4.69 |
|
|
|
| 26 |
A" |
3146 |
2990 |
59.60 |
|
|
|
| 27 |
A" |
3144 |
2987 |
0.94 |
|
|
|
| 28 |
A" |
3122 |
2967 |
0.23 |
|
|
|
| 29 |
A" |
3076 |
2923 |
7.80 |
|
|
|
| 30 |
A" |
1527 |
1451 |
7.53 |
|
|
|
| 31 |
A" |
1501 |
1426 |
12.98 |
|
|
|
| 32 |
A" |
1337 |
1271 |
0.09 |
|
|
|
| 33 |
A" |
1264 |
1201 |
0.00 |
|
|
|
| 34 |
A" |
1152 |
1095 |
1.71 |
|
|
|
| 35 |
A" |
984 |
936 |
1.34 |
|
|
|
| 36 |
A" |
839 |
798 |
0.75 |
|
|
|
| 37 |
A" |
729 |
692 |
4.77 |
|
|
|
| 38 |
A" |
476 |
453 |
0.00 |
|
|
|
| 39 |
A" |
248 |
236 |
0.05 |
|
|
|
| 40 |
A" |
96 |
92 |
0.10 |
|
|
|
| 41 |
A" |
82 |
78 |
0.54 |
|
|
|
| 42 |
A" |
34i |
33i |
0.51 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 31230.8 cm
-1
Scaled (by 0.9503) Zero Point Vibrational Energy (zpe) 29678.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBE1PBE/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.525 |
|
|
|
| 2 |
C |
-0.313 |
|
|
|
| 3 |
C |
-0.419 |
|
|
|
| 4 |
C |
0.455 |
|
|
|
| 5 |
C |
-0.611 |
|
|
|
| 6 |
O |
-0.435 |
|
|
|
| 7 |
H |
0.177 |
|
|
|
| 8 |
H |
0.168 |
|
|
|
| 9 |
H |
0.168 |
|
|
|
| 10 |
H |
0.184 |
|
|
|
| 11 |
H |
0.184 |
|
|
|
| 12 |
H |
0.185 |
|
|
|
| 13 |
H |
0.185 |
|
|
|
| 14 |
H |
0.209 |
|
|
|
| 15 |
H |
0.194 |
|
|
|
| 16 |
H |
0.194 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.730 |
2.551 |
0.000 |
2.653 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-34.975 |
1.434 |
0.000 |
| y |
1.434 |
-43.150 |
0.000 |
| z |
0.000 |
0.000 |
-36.638 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
4.919 |
1.434 |
0.000 |
| y |
1.434 |
-7.344 |
0.000 |
| z |
0.000 |
0.000 |
2.425 |
|
| Polar |
|---|
| 3z2-r2 | 4.849 |
|---|
| x2-y2 | 8.175 |
|---|
| xy | 1.434 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.288 |
-0.283 |
0.000 |
| y |
-0.283 |
8.379 |
0.000 |
| z |
0.000 |
0.000 |
6.655 |
<r2> (average value of r
2) Å
2
| <r2> |
227.118 |
| (<r2>)1/2 |
15.070 |